return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH2CH2NH2 (diaminomethane)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-150.815284
Energy at 298.15K-150.823404
HF Energy-150.272607
Counterpoise corrected energy-150.815284
CP Energy at 298.15K-150.823404
Counterpoise optimized geometry corrected energy 
CP opt. Energy at 298.15K 
Nuclear repulsion energy83.252397
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3546 3358 1.90      
2 A1 3118 2953 38.23      
3 A1 1748 1655 71.41      
4 A1 1523 1442 0.88      
5 A1 1152 1091 39.86      
6 A1 914 866 0.75      
7 A1 468 443 5.27      
8 A2 3648 3455 0.00      
9 A2 1436 1360 0.00      
10 A2 1107 1048 0.00      
11 A2 284 269 0.00      
12 B1 3646 3453 0.01      
13 B1 3177 3009 27.76      
14 B1 1417 1342 0.68      
15 B1 867 821 0.82      
16 B1 429 406 108.07      
17 B2 3547 3359 0.11      
18 B2 1736 1644 1.86      
19 B2 1423 1348 21.27      
20 B2 1125 1066 71.67      
21 B2 850 805 491.86      

Unscaled Zero Point Vibrational Energy (zpe) 18579.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17596.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.18900 0.30265 0.27466

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.551
N2 0.000 1.260 -0.184
N3 0.000 -1.260 -0.184
H4 0.879 0.000 1.198
H5 -0.879 0.000 1.198
H6 0.817 1.318 -0.780
H7 -0.817 1.318 -0.780
H8 -0.817 -1.318 -0.780
H9 0.817 -1.318 -0.780

Atom - Atom Distances (Å)
  C1 N2 N3 H4 H5 H6 H7 H8 H9
C11.45861.45861.09181.09182.04332.04332.04332.0433
N21.45862.51962.06682.06681.01291.01292.76882.7688
N31.45862.51962.06682.06682.76882.76881.01291.0129
H41.09182.06682.06681.75812.37772.92022.92022.3777
H51.09182.06682.06681.75812.92022.37772.37772.9202
H62.04331.01292.76882.37772.92021.63463.10132.6355
H72.04331.01292.76882.92022.37771.63462.63553.1013
H82.04332.76881.01292.92022.37773.10132.63551.6346
H92.04332.76881.01292.37772.92022.63553.10131.6346

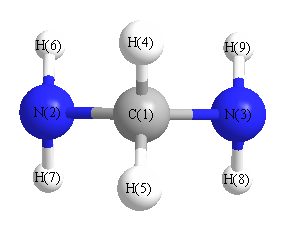
picture of diaminomethane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N2 H6 110.226 C1 N2 H7 110.226
C1 N3 H8 110.226 C1 N3 H9 110.226
N2 C1 N3 119.468 N2 C1 H4 107.393
N2 C1 H5 107.393 N3 C1 H4 107.393
N3 C1 H5 107.393 H4 C1 H5 107.247
H6 N2 H7 107.587 H8 N3 H9 107.587
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability