return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5 (phenyl)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-230.934946
Energy at 298.15K-230.940026
HF Energy-230.099323
Nuclear repulsion energy197.476234
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3253 3081 2.33      
2 A1 3248 3076 7.85      
3 A1 3219 3048 0.46      
4 A1 1827 1730 3.44      
5 A1 1575 1491 16.25      
6 A1 1238 1172 0.08      
7 A1 1127 1067 21.01      
8 A1 1085 1027 0.08      
9 A1 980 928 0.34      
10 A1 653 619 1.61      
11 A2 1094 1036 0.00      
12 A2 968 917 0.00      
13 A2 468 443 0.00      
14 B1 1065 1009 0.12      
15 B1 1017 964 0.93      
16 B1 810 767 129.59      
17 B1 705 668 2.52      
18 B1 491 465 7.86      
19 B2 3253 3081 7.20      
20 B2 3228 3057 9.30      
21 B2 1885 1785 9.30      
22 B2 1567 1484 9.04      
23 B2 1356 1285 0.41      
24 B2 1258 1192 0.14      
25 B2 1182 1119 5.98      
26 B2 1087 1030 5.59      
27 B2 629 596 0.81      

Unscaled Zero Point Vibrational Energy (zpe) 20132.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 19067.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.21706 0.19309 0.10219

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.372
C2 0.000 1.207 0.756
C3 0.000 -1.207 0.756
C4 0.000 1.190 -0.618
C5 0.000 -1.190 -0.618
C6 0.000 0.000 -1.301
H7 0.000 2.140 1.311
H8 0.000 -2.140 1.311
H9 0.000 2.129 -1.165
H10 0.000 -2.129 -1.165
H11 0.000 0.000 -2.386

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11
C11.35531.35532.31852.31852.67282.14062.14063.31183.31183.7581
C21.35532.41451.37432.76282.38501.08483.39252.13073.84963.3661
C31.35532.41452.76281.37432.38503.39251.08483.84962.13073.3661
C42.31851.37432.76282.37921.37162.15003.84761.08683.36312.1310
C52.31852.76281.37432.37921.37163.84762.15003.36311.08682.1310
C62.67282.38502.38501.37161.37163.37603.37602.13312.13311.0853
H72.14061.08483.39252.15003.84763.37604.27942.47554.93444.2713
H82.14063.39251.08483.84762.15003.37604.27944.93442.47554.2713
H93.31182.13073.84961.08683.36312.13312.47554.93444.25762.4542
H103.31183.84962.13073.36311.08682.13314.93442.47554.25762.4542
H113.75813.36613.36612.13102.13101.08534.27134.27132.45422.4542

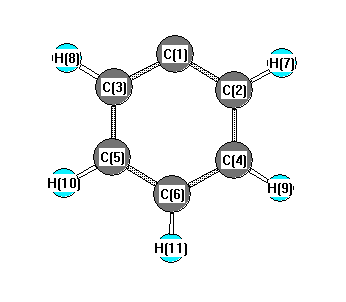
picture of phenyl state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 116.294 C1 C2 H7 122.234
C1 C3 C5 116.294 C1 C3 H8 122.234
C2 C1 C3 125.942 C2 C4 C6 120.588
C2 C4 H9 119.476 C3 C5 C6 120.588
C3 C5 H10 119.476 C4 C2 H7 121.472
C4 C6 C5 120.294 C4 C6 H11 119.853
C5 C3 H8 121.472 C5 C6 H11 119.853
C6 C4 H9 119.936 C6 C5 H10 119.936
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability