Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -189.067782 |
| Energy at 298.15K | -189.068897 |
| HF Energy | -189.067782 |
| Nuclear repulsion energy | 63.383641 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3573 |
3440 |
5.72 |
|
|
|
| 2 |
A' |
1902 |
1831 |
331.64 |
|
|
|
| 3 |
A' |
1307 |
1259 |
1.51 |
|
|
|
| 4 |
A' |
1106 |
1065 |
160.11 |
|
|
|
| 5 |
A' |
603 |
580 |
37.59 |
|
|
|
| 6 |
A" |
629 |
606 |
135.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4560.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 4389.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
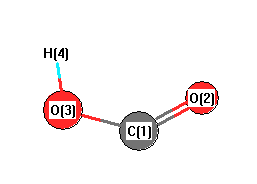
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.441 |
0.000 |
| O2 |
-1.057 |
-0.351 |
0.000 |
| O3 |
1.153 |
0.181 |
0.000 |
| H4 |
-0.772 |
-1.285 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
| C1 | | 1.3206 | 1.1821 | 1.8905 |
O2 | 1.3206 | | 2.2734 | 0.9764 | O3 | 1.1821 | 2.2734 | | 2.4201 | H4 | 1.8905 | 0.9764 | 2.4201 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
109.874 |
|
O2 |
C1 |
O3 |
130.488 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.307 |
|
|
|
| 2 |
O |
-0.445 |
|
|
|
| 3 |
O |
-0.254 |
|
|
|
| 4 |
H |
0.392 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.921 |
-1.847 |
0.000 |
2.064 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.272 |
0.744 |
0.000 |
| y |
0.744 |
-13.632 |
0.000 |
| z |
0.000 |
0.000 |
-16.010 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.451 |
0.744 |
0.000 |
| y |
0.744 |
4.509 |
0.000 |
| z |
0.000 |
0.000 |
0.942 |
|
| Polar |
|---|
| 3z2-r2 | 1.885 |
|---|
| x2-y2 | -6.640 |
|---|
| xy | 0.744 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.166 |
0.261 |
0.000 |
| y |
0.261 |
2.651 |
0.000 |
| z |
0.000 |
0.000 |
1.487 |
<r2> (average value of r
2) Å
2
| <r2> |
34.631 |
| (<r2>)1/2 |
5.885 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -189.069801 |
| Energy at 298.15K | -189.070889 |
| HF Energy | -189.069801 |
| Nuclear repulsion energy | 63.085934 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3806 |
3664 |
85.71 |
|
|
|
| 2 |
A' |
1935 |
1863 |
222.59 |
|
|
|
| 3 |
A' |
1265 |
1218 |
281.56 |
|
|
|
| 4 |
A' |
1104 |
1062 |
53.32 |
|
|
|
| 5 |
A' |
626 |
602 |
5.11 |
|
|
|
| 6 |
A" |
569 |
547 |
110.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4651.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 4478.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.410 |
0.000 |
| O2 |
-0.940 |
-0.544 |
0.000 |
| O3 |
1.166 |
0.252 |
0.000 |
| H4 |
-1.809 |
-0.126 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
| C1 | | 1.3388 | 1.1767 | 1.8871 |
O2 | 1.3388 | | 2.2513 | 0.9642 | O3 | 1.1767 | 2.2513 | | 2.9993 | H4 | 1.8871 | 0.9642 | 2.9993 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H4 |
108.963 |
|
O2 |
C1 |
O3 |
126.879 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.253 |
|
|
|
| 2 |
O |
-0.441 |
|
|
|
| 3 |
O |
-0.234 |
|
|
|
| 4 |
H |
0.423 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.094 |
0.421 |
0.000 |
3.122 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.488 |
-1.084 |
0.000 |
| y |
-1.084 |
-17.836 |
0.000 |
| z |
0.000 |
0.000 |
-16.055 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.457 |
-1.084 |
0.000 |
| y |
-1.084 |
-3.064 |
0.000 |
| z |
0.000 |
0.000 |
-0.393 |
|
| Polar |
|---|
| 3z2-r2 | -0.785 |
|---|
| x2-y2 | 4.348 |
|---|
| xy | -1.084 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.679 |
0.188 |
0.000 |
| y |
0.188 |
2.005 |
0.000 |
| z |
0.000 |
0.000 |
1.519 |
<r2> (average value of r
2) Å
2
| <r2> |
34.979 |
| (<r2>)1/2 |
5.914 |
