Jump to
S1C2
S2C1
S2C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.103940 |
| Energy at 298.15K | -131.103561 |
| HF Energy | -130.704156 |
| Nuclear repulsion energy | 47.512242 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3349 |
3172 |
13.83 |
|
|
|
| 2 |
A' |
2307 |
2185 |
0.60 |
|
|
|
| 3 |
A' |
1067 |
1010 |
30.65 |
|
|
|
| 4 |
A' |
846 |
801 |
9.39 |
|
|
|
| 5 |
A' |
455 |
431 |
5.01 |
|
|
|
| 6 |
A" |
536 |
508 |
7.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4280.1 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 4053.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.288 |
-1.225 |
0.000 |
| C2 |
0.000 |
0.122 |
0.000 |
| N3 |
0.204 |
1.249 |
0.000 |
| H4 |
0.298 |
-2.127 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3775 | 2.5216 | 1.0761 |
C2 | 1.3775 | | 1.1444 | 2.2692 | N3 | 2.5216 | 1.1444 | | 3.3768 | H4 | 1.0761 | 2.2692 | 3.3768 | |
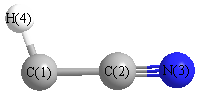 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
178.213 |
|
C2 |
C1 |
H4 |
134.931 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.098370 |
| Energy at 298.15K | |
| HF Energy | -130.703579 |
| Nuclear repulsion energy | 47.831787 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3483 |
3299 |
72.28 |
|
|
|
| 2 |
Σ |
1728 |
1636 |
32.08 |
|
|
|
| 3 |
Σ |
1179 |
1117 |
44.81 |
|
|
|
| 4 |
Π |
522 |
495 |
1.13 |
|
|
|
| 4 |
Π |
522 |
495 |
1.13 |
|
|
|
| 5 |
Π |
525i |
497i |
60.06 |
|
|
|
| 5 |
Π |
525i |
497i |
60.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3192.6 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 3023.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.209 |
| C2 |
0.000 |
0.000 |
0.103 |
| N3 |
0.000 |
0.000 |
1.273 |
| H4 |
0.000 |
0.000 |
-2.274 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3115 | 2.4820 | 1.0647 |
C2 | 1.3115 | | 1.1704 | 2.3763 | N3 | 2.4820 | 1.1704 | | 3.5467 | H4 | 1.0647 | 2.3763 | 3.5467 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.092 |
|
|
|
| 2 |
C |
0.037 |
|
|
|
| 3 |
N |
-0.259 |
|
|
|
| 4 |
H |
0.314 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.713 |
3.713 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.441 |
0.000 |
0.000 |
| y |
0.000 |
-17.441 |
0.000 |
| z |
0.000 |
0.000 |
-15.528 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.957 |
0.000 |
0.000 |
| y |
0.000 |
-0.957 |
0.000 |
| z |
0.000 |
0.000 |
1.914 |
|
| Polar |
|---|
| 3z2-r2 | 3.828 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.841 |
0.000 |
0.000 |
| y |
0.000 |
1.841 |
0.000 |
| z |
0.000 |
0.000 |
6.225 |
<r2> (average value of r
2) Å
2
| <r2> |
35.841 |
| (<r2>)1/2 |
5.987 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.097760 |
| Energy at 298.15K | -131.097219 |
| HF Energy | -130.644412 |
| Nuclear repulsion energy | 46.597922 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3105 |
2941 |
14.09 |
|
|
|
| 2 |
A' |
2240 |
2121 |
11.32 |
|
|
|
| 3 |
A' |
1074 |
1017 |
77.11 |
|
|
|
| 4 |
A' |
984 |
932 |
0.79 |
|
|
|
| 5 |
A' |
418 |
396 |
18.05 |
|
|
|
| 6 |
A" |
350 |
332 |
12.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4086.0 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 3869.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.064 |
-1.303 |
0.000 |
| C2 |
0.000 |
0.099 |
0.000 |
| N3 |
-0.215 |
1.262 |
0.000 |
| H4 |
1.121 |
-1.609 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.4038 | 2.5798 | 1.1005 |
C2 | 1.4038 | | 1.1820 | 2.0433 | N3 | 2.5798 | 1.1820 | | 3.1663 | H4 | 1.1005 | 2.0433 | 3.1663 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
S2C1
Energy calculated at MP2=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.097760 |
| Energy at 298.15K | -131.097219 |
| HF Energy | -130.644412 |
| Nuclear repulsion energy | 46.597922 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Geometric Data calculated at MP2=FULL/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
