Jump to
S1C2
S2C1
S2C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.305193 |
| Energy at 298.15K | -131.304523 |
| HF Energy | -131.305193 |
| Nuclear repulsion energy | 47.487148 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3395 |
3258 |
29.49 |
|
|
|
| 2 |
A' |
1818 |
1745 |
38.00 |
|
|
|
| 3 |
A' |
1233 |
1183 |
8.42 |
|
|
|
| 4 |
A' |
489 |
469 |
23.29 |
|
|
|
| 5 |
A' |
386 |
371 |
25.76 |
|
|
|
| 6 |
A" |
455 |
436 |
1.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3887.7 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 3730.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.048 |
-1.219 |
0.000 |
| C2 |
0.000 |
0.088 |
0.000 |
| N3 |
-0.125 |
1.275 |
0.000 |
| H4 |
0.591 |
-2.141 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3072 | 2.4996 | 1.0703 |
C2 | 1.3072 | | 1.1939 | 2.3055 | N3 | 2.4996 | 1.1939 | | 3.4900 | H4 | 1.0703 | 2.3055 | 3.4900 | |
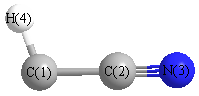 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
176.068 |
|
C2 |
C1 |
H4 |
151.594 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S2C1
S2C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.304567 |
| Energy at 298.15K | |
| HF Energy | -131.304567 |
| Nuclear repulsion energy | 47.549161 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3457 |
3317 |
56.41 |
|
|
|
| 2 |
Σ |
1748 |
1677 |
41.84 |
|
|
|
| 3 |
Σ |
1275 |
1224 |
6.09 |
|
|
|
| 4 |
Π |
455 |
437 |
0.41 |
|
|
|
| 4 |
Π |
455 |
437 |
0.41 |
|
|
|
| 5 |
Π |
287i |
275i |
51.48 |
|
|
|
| 5 |
Π |
287i |
275i |
51.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3407.8 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 3270.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
-1.206 |
| C2 |
0.000 |
0.000 |
0.083 |
| N3 |
0.000 |
0.000 |
1.286 |
| H4 |
0.000 |
0.000 |
-2.271 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.2890 | 2.4917 | 1.0653 |
C2 | 1.2890 | | 1.2027 | 2.3543 | N3 | 2.4917 | 1.2027 | | 3.5570 | H4 | 1.0653 | 2.3543 | 3.5570 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
180.000 |
|
C2 |
C1 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.063 |
|
|
|
| 2 |
C |
0.023 |
|
|
|
| 3 |
N |
-0.251 |
|
|
|
| 4 |
H |
0.291 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.290 |
3.290 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.241 |
0.000 |
0.000 |
| y |
0.000 |
-17.241 |
0.000 |
| z |
0.000 |
0.000 |
-15.293 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.974 |
0.000 |
0.000 |
| y |
0.000 |
-0.974 |
0.000 |
| z |
0.000 |
0.000 |
1.948 |
|
| Polar |
|---|
| 3z2-r2 | 3.895 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.900 |
0.000 |
0.000 |
| y |
0.000 |
1.900 |
0.000 |
| z |
0.000 |
0.000 |
6.297 |
<r2> (average value of r
2) Å
2
| <r2> |
35.861 |
| (<r2>)1/2 |
5.988 |
Jump to
S1C1
S1C2
S2C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.273426 |
| Energy at 298.15K | -131.272888 |
| HF Energy | -131.273426 |
| Nuclear repulsion energy | 47.175112 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3096 |
2971 |
9.62 |
|
|
|
| 2 |
A' |
2124 |
2038 |
21.64 |
|
|
|
| 3 |
A' |
1099 |
1054 |
37.45 |
|
|
|
| 4 |
A' |
948 |
909 |
48.71 |
|
|
|
| 5 |
A' |
462 |
443 |
25.29 |
|
|
|
| 6 |
A" |
323 |
310 |
10.56 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4025.4 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 3862.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.069 |
-1.277 |
0.000 |
| C2 |
0.000 |
0.093 |
0.000 |
| N3 |
-0.218 |
1.247 |
0.000 |
| H4 |
1.113 |
-1.621 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
| C1 | | 1.3713 | 2.5401 | 1.0994 |
C2 | 1.3713 | | 1.1747 | 2.0435 | N3 | 2.5401 | 1.1747 | | 3.1619 | H4 | 1.0994 | 2.0435 | 3.1619 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.141 |
|
|
|
| 2 |
C |
0.112 |
|
|
|
| 3 |
N |
-0.227 |
|
|
|
| 4 |
H |
0.256 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.816 |
-1.383 |
0.000 |
2.282 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.916 |
-2.532 |
0.000 |
| y |
-2.532 |
-21.049 |
0.000 |
| z |
0.000 |
0.000 |
-15.851 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.534 |
-2.532 |
0.000 |
| y |
-2.532 |
-4.666 |
0.000 |
| z |
0.000 |
0.000 |
3.132 |
|
| Polar |
|---|
| 3z2-r2 | 6.264 |
|---|
| x2-y2 | 4.133 |
|---|
| xy | -2.532 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.578 |
-0.664 |
0.000 |
| y |
-0.664 |
6.308 |
0.000 |
| z |
0.000 |
0.000 |
2.412 |
<r2> (average value of r
2) Å
2
| <r2> |
36.151 |
| (<r2>)1/2 |
6.013 |
Jump to
S1C1
S1C2
S2C1
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -131.273426 |
| Energy at 298.15K | -131.272888 |
| HF Energy | -131.273426 |
| Nuclear repulsion energy | 47.175112 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
