return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H7N ((E)-2-Pentenenitrile)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-248.835561
Energy at 298.15K-248.842385
HF Energy-247.886378
Nuclear repulsion energy195.935647
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3238 3067 1.75      
2 A' 3192 3023 7.11      
3 A' 3178 3010 21.22      
4 A' 3086 2923 16.32      
5 A' 3070 2908 13.78      
6 A' 2194 2078 0.03      
7 A' 1706 1615 8.39      
8 A' 1548 1466 7.50      
9 A' 1509 1429 11.08      
10 A' 1455 1378 9.25      
11 A' 1408 1334 4.45      
12 A' 1351 1279 0.60      
13 A' 1320 1250 3.23      
14 A' 1139 1079 2.52      
15 A' 1084 1027 2.79      
16 A' 995 943 2.49      
17 A' 921 872 4.24      
18 A' 579 549 0.60      
19 A' 505 478 0.02      
20 A' 265 251 0.99      
21 A' 151 143 3.92      
22 A" 3174 3006 23.05      
23 A" 3107 2942 7.86      
24 A" 1543 1462 11.20      
25 A" 1318 1248 0.19      
26 A" 1128 1068 1.45      
27 A" 993 941 41.63      
28 A" 850 805 7.47      
29 A" 740 701 0.40      
30 A" 474 449 4.52      
31 A" 293 277 0.00      
32 A" 175 166 0.01      
33 A" 115 109 3.38      

Unscaled Zero Point Vibrational Energy (zpe) 23900.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 22636.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.48473 0.05163 0.04747

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -1.541 2.763 0.000
C2 -0.849 1.811 0.000
H3 1.068 0.850 0.000
C4 0.000 0.661 0.000
H5 -1.585 -0.706 0.000
C6 -0.502 -0.587 0.000
H7 0.005 -2.443 0.871
H8 0.005 -2.443 -0.871
C9 0.308 -1.848 0.000
H10 2.322 -2.629 0.000
H11 2.153 -1.114 -0.885
H12 2.153 -1.114 0.885
C13 1.820 -1.660 0.000

Atom - Atom Distances (Å)
  N1 C2 H3 C4 H5 C6 H7 H8 C9 H10 H11 H12 C13
N11.17593.23532.60593.46863.50745.49975.49974.96776.63255.42755.42755.5544
C21.17592.14551.43002.62242.42394.42594.42593.83835.45674.28464.28464.3788
H33.23532.14551.08503.07552.12853.56773.56772.80283.69762.41172.41172.6195
C42.60591.43001.08502.09271.34553.22373.22372.52804.02692.92772.92772.9492
H53.46862.62243.07552.09271.08952.51112.51112.21114.35463.86343.86343.5358
C63.50742.42392.12851.34551.08952.11162.11161.49853.48452.84812.84812.5573
H75.49974.42593.56773.22372.51112.11161.74191.09722.48173.07652.52592.1595
H85.49974.42593.56773.22372.51112.11161.74191.09722.48172.52593.07652.1595
C94.96773.83832.80282.52802.21111.49851.09721.09722.15982.17442.17441.5233
H106.63255.45673.69764.02694.35463.48452.48172.48172.15981.76301.76301.0916
H115.42754.28462.41172.92773.86342.84813.07652.52592.17441.76301.77091.0925
H125.42754.28462.41172.92773.86342.84812.52593.07652.17441.76301.77091.0925
C135.55444.37882.61952.94923.53582.55732.15952.15951.52331.09161.09251.0925

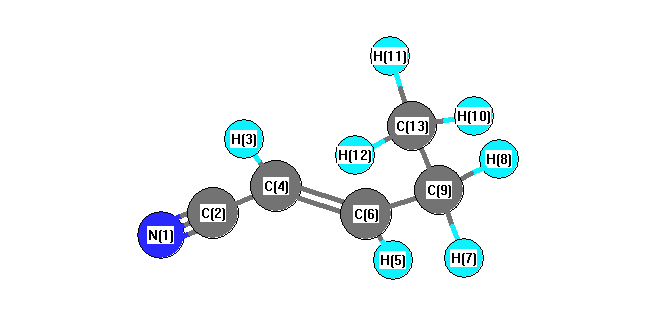
picture of (E)-2-Pentenenitrile state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 179.574 C2 C4 H3 116.425
C2 C4 C6 121.666 H3 C4 C6 121.909
C4 C6 H5 118.128 C4 C6 C9 125.382
H5 C6 C9 116.490 C6 C9 H7 107.872
C6 C9 H8 107.872 C6 C9 C13 115.618
H7 C9 H8 105.083 H7 C9 C13 109.924
H8 C9 C13 109.924 C9 C13 H10 110.287
C9 C13 H11 111.400 C9 C13 H12 111.400
H10 C13 H11 107.653 H10 C13 H12 107.653
H11 C13 H12 108.293
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability