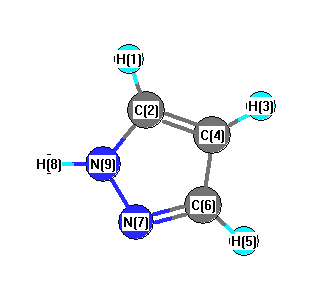
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3952 |
3574 |
90.92 |
|
|
|
| 2 |
A' |
3437 |
3109 |
3.13 |
|
|
|
| 3 |
A' |
3418 |
3091 |
10.02 |
|
|
|
| 4 |
A' |
3404 |
3079 |
9.19 |
|
|
|
| 5 |
A' |
1743 |
1576 |
10.40 |
|
|
|
| 6 |
A' |
1637 |
1480 |
12.70 |
|
|
|
| 7 |
A' |
1542 |
1394 |
25.53 |
|
|
|
| 8 |
A' |
1540 |
1393 |
10.58 |
|
|
|
| 9 |
A' |
1404 |
1269 |
3.36 |
|
|
|
| 10 |
A' |
1281 |
1159 |
3.10 |
|
|
|
| 11 |
A' |
1226 |
1109 |
27.82 |
|
|
|
| 12 |
A' |
1139 |
1030 |
55.77 |
|
|
|
| 13 |
A' |
1123 |
1016 |
0.27 |
|
|
|
| 14 |
A' |
1019 |
921 |
6.44 |
|
|
|
| 15 |
A' |
1001 |
906 |
13.94 |
|
|
|
| 16 |
A" |
1014 |
917 |
3.84 |
|
|
|
| 17 |
A" |
977 |
883 |
3.33 |
|
|
|
| 18 |
A" |
845 |
764 |
108.28 |
|
|
|
| 19 |
A" |
748 |
677 |
23.34 |
|
|
|
| 20 |
A" |
689 |
623 |
0.16 |
|
|
|
| 21 |
A" |
555 |
502 |
91.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16846.0 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 15235.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.246 |
0.143 |
|
0.176 |
| 2 |
C |
0.082 |
-0.069 |
|
-0.087 |
| 3 |
H |
0.220 |
0.152 |
|
0.209 |
| 4 |
C |
-0.498 |
-0.313 |
|
-0.396 |
| 5 |
H |
0.225 |
0.081 |
|
0.127 |
| 6 |
C |
0.023 |
0.224 |
|
0.168 |
| 7 |
N |
-0.231 |
-0.496 |
|
-0.466 |
| 8 |
H |
0.386 |
0.289 |
|
0.324 |
| 9 |
N |
-0.453 |
-0.011 |
|
-0.056 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.243 |
0.866 |
0.000 |
2.405 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
2.241 |
0.855 |
0.000 |
2.399 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.518 |
2.836 |
0.000 |
| y |
2.836 |
-21.996 |
0.000 |
| z |
0.000 |
0.000 |
-32.349 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.345 |
2.836 |
0.000 |
| y |
2.836 |
8.938 |
0.000 |
| z |
0.000 |
0.000 |
-6.592 |
|
| Polar |
|---|
| 3z2-r2 | -13.185 |
|---|
| x2-y2 | -7.522 |
|---|
| xy | 2.836 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.535 |
0.260 |
0.000 |
| y |
0.260 |
6.994 |
0.000 |
| z |
0.000 |
0.000 |
3.143 |
<r2> (average value of r
2) Å
2
| <r2> |
78.706 |
| (<r2>)1/2 |
8.872 |