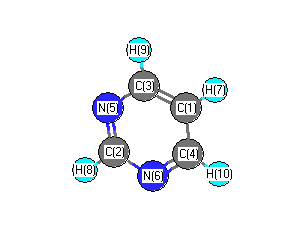
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3386 |
3062 |
11.66 |
|
|
|
| 2 |
A1 |
3371 |
3049 |
34.98 |
|
|
|
| 3 |
A1 |
3344 |
3024 |
12.23 |
|
|
|
| 4 |
A1 |
1787 |
1616 |
95.27 |
|
|
|
| 5 |
A1 |
1562 |
1413 |
90.00 |
|
|
|
| 6 |
A1 |
1253 |
1133 |
1.04 |
|
|
|
| 7 |
A1 |
1159 |
1048 |
2.69 |
|
|
|
| 8 |
A1 |
1100 |
995 |
5.52 |
|
|
|
| 9 |
A1 |
751 |
680 |
3.86 |
|
|
|
| 10 |
A2 |
1125 |
1018 |
0.00 |
|
|
|
| 11 |
A2 |
461 |
417 |
0.00 |
|
|
|
| 12 |
B1 |
1150 |
1041 |
0.02 |
|
|
|
| 13 |
B1 |
1092 |
988 |
1.22 |
|
|
|
| 14 |
B1 |
908 |
821 |
6.31 |
|
|
|
| 15 |
B1 |
786 |
711 |
58.62 |
|
|
|
| 16 |
B1 |
415 |
375 |
2.99 |
|
|
|
| 17 |
B2 |
3349 |
3029 |
40.16 |
|
|
|
| 18 |
B2 |
1790 |
1619 |
139.08 |
|
|
|
| 19 |
B2 |
1632 |
1476 |
22.89 |
|
|
|
| 20 |
B2 |
1513 |
1368 |
1.47 |
|
|
|
| 21 |
B2 |
1352 |
1222 |
12.81 |
|
|
|
| 22 |
B2 |
1201 |
1086 |
8.23 |
|
|
|
| 23 |
B2 |
1143 |
1034 |
4.56 |
|
|
|
| 24 |
B2 |
684 |
619 |
13.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18155.9 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 16420.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.388 |
-0.733 |
|
-0.837 |
| 2 |
C |
0.126 |
0.887 |
|
0.882 |
| 3 |
C |
0.059 |
0.681 |
|
0.688 |
| 4 |
C |
0.059 |
0.681 |
|
0.688 |
| 5 |
N |
-0.394 |
-0.849 |
|
-0.856 |
| 6 |
N |
-0.394 |
-0.849 |
|
-0.856 |
| 7 |
H |
0.230 |
0.206 |
|
0.255 |
| 8 |
H |
0.232 |
-0.036 |
|
-0.017 |
| 9 |
H |
0.236 |
0.006 |
|
0.026 |
| 10 |
H |
0.236 |
0.006 |
|
0.026 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.377 |
2.377 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.000 |
0.000 |
2.404 |
2.404 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.592 |
0.000 |
0.000 |
| y |
0.000 |
8.202 |
0.000 |
| z |
0.000 |
0.000 |
8.700 |
<r2> (average value of r
2) Å
2
| <r2> |
112.927 |
| (<r2>)1/2 |
10.627 |