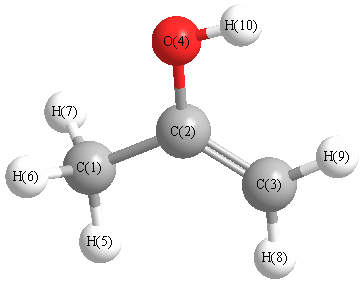
Vibrational Frequencies calculated at PBE1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3835 |
3676 |
18.77 |
|
|
|
| 2 |
A' |
3264 |
3128 |
11.13 |
|
|
|
| 3 |
A' |
3170 |
3039 |
15.67 |
|
|
|
| 4 |
A' |
3163 |
3031 |
7.53 |
|
|
|
| 5 |
A' |
3062 |
2935 |
19.15 |
|
|
|
| 6 |
A' |
1750 |
1677 |
164.22 |
|
|
|
| 7 |
A' |
1505 |
1443 |
10.17 |
|
|
|
| 8 |
A' |
1470 |
1409 |
1.11 |
|
|
|
| 9 |
A' |
1429 |
1370 |
52.19 |
|
|
|
| 10 |
A' |
1386 |
1328 |
5.39 |
|
|
|
| 11 |
A' |
1236 |
1184 |
167.38 |
|
|
|
| 12 |
A' |
1033 |
990 |
29.39 |
|
|
|
| 13 |
A' |
990 |
949 |
7.33 |
|
|
|
| 14 |
A' |
884 |
847 |
4.32 |
|
|
|
| 15 |
A' |
485 |
465 |
19.94 |
|
|
|
| 16 |
A' |
411 |
394 |
1.74 |
|
|
|
| 17 |
A" |
3123 |
2993 |
15.84 |
|
|
|
| 18 |
A" |
1485 |
1424 |
11.20 |
|
|
|
| 19 |
A" |
1080 |
1035 |
0.96 |
|
|
|
| 20 |
A" |
798 |
765 |
90.60 |
|
|
|
| 21 |
A" |
733 |
702 |
2.43 |
|
|
|
| 22 |
A" |
510 |
489 |
0.26 |
|
|
|
| 23 |
A" |
463 |
444 |
127.87 |
|
|
|
| 24 |
A" |
182 |
175 |
1.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18722.6 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 17945.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBE1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.689 |
|
|
|
| 2 |
C |
0.263 |
|
|
|
| 3 |
C |
-0.577 |
|
|
|
| 4 |
O |
-0.548 |
|
|
|
| 5 |
H |
0.235 |
|
|
|
| 6 |
H |
0.244 |
|
|
|
| 7 |
H |
0.244 |
|
|
|
| 8 |
H |
0.220 |
|
|
|
| 9 |
H |
0.207 |
|
|
|
| 10 |
H |
0.402 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.144 |
0.631 |
0.000 |
0.647 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.147 |
-3.478 |
0.000 |
| y |
-3.478 |
-24.340 |
0.000 |
| z |
0.000 |
0.000 |
-27.101 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.574 |
-3.478 |
0.000 |
| y |
-3.478 |
0.285 |
0.000 |
| z |
0.000 |
0.000 |
-3.858 |
|
| Polar |
|---|
| 3z2-r2 | -7.717 |
|---|
| x2-y2 | 2.193 |
|---|
| xy | -3.478 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.635 |
0.305 |
0.000 |
| y |
0.305 |
7.294 |
0.000 |
| z |
0.000 |
0.000 |
3.583 |
<r2> (average value of r
2) Å
2
| <r2> |
80.010 |
| (<r2>)1/2 |
8.945 |