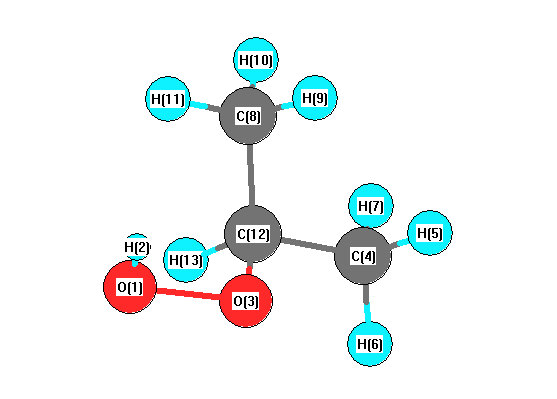
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3763 |
3622 |
21.48 |
|
|
|
| 2 |
A |
3136 |
3019 |
23.24 |
|
|
|
| 3 |
A |
3130 |
3013 |
23.17 |
|
|
|
| 4 |
A |
3126 |
3010 |
45.94 |
|
|
|
| 5 |
A |
3113 |
2997 |
16.40 |
|
|
|
| 6 |
A |
3051 |
2937 |
18.20 |
|
|
|
| 7 |
A |
3042 |
2928 |
23.90 |
|
|
|
| 8 |
A |
3037 |
2924 |
17.99 |
|
|
|
| 9 |
A |
1527 |
1470 |
10.42 |
|
|
|
| 10 |
A |
1507 |
1451 |
4.44 |
|
|
|
| 11 |
A |
1498 |
1442 |
2.91 |
|
|
|
| 12 |
A |
1492 |
1436 |
2.90 |
|
|
|
| 13 |
A |
1425 |
1372 |
1.67 |
|
|
|
| 14 |
A |
1413 |
1360 |
37.32 |
|
|
|
| 15 |
A |
1408 |
1356 |
39.89 |
|
|
|
| 16 |
A |
1384 |
1332 |
12.09 |
|
|
|
| 17 |
A |
1354 |
1304 |
16.83 |
|
|
|
| 18 |
A |
1200 |
1156 |
4.19 |
|
|
|
| 19 |
A |
1184 |
1140 |
33.17 |
|
|
|
| 20 |
A |
1151 |
1108 |
13.27 |
|
|
|
| 21 |
A |
975 |
939 |
2.86 |
|
|
|
| 22 |
A |
955 |
919 |
2.35 |
|
|
|
| 23 |
A |
933 |
898 |
1.98 |
|
|
|
| 24 |
A |
915 |
881 |
10.10 |
|
|
|
| 25 |
A |
832 |
801 |
2.98 |
|
|
|
| 26 |
A |
507 |
488 |
6.05 |
|
|
|
| 27 |
A |
470 |
453 |
4.53 |
|
|
|
| 28 |
A |
351 |
338 |
0.42 |
|
|
|
| 29 |
A |
289 |
278 |
4.36 |
|
|
|
| 30 |
A |
264 |
254 |
44.14 |
|
|
|
| 31 |
A |
220 |
212 |
78.87 |
|
|
|
| 32 |
A |
201 |
194 |
31.12 |
|
|
|
| 33 |
A |
138 |
133 |
6.49 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24495.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 23581.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.415 |
|
|
|
| 2 |
H |
0.411 |
|
|
|
| 3 |
O |
-0.233 |
|
|
|
| 4 |
C |
-0.665 |
|
|
|
| 5 |
H |
0.226 |
|
|
|
| 6 |
H |
0.241 |
|
|
|
| 7 |
H |
0.225 |
|
|
|
| 8 |
C |
-0.666 |
|
|
|
| 9 |
H |
0.217 |
|
|
|
| 10 |
H |
0.222 |
|
|
|
| 11 |
H |
0.230 |
|
|
|
| 12 |
C |
-0.005 |
|
|
|
| 13 |
H |
0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.745 |
1.514 |
0.677 |
1.818 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.228 |
-2.575 |
-3.497 |
| y |
-2.575 |
-32.773 |
0.614 |
| z |
-3.497 |
0.614 |
-31.027 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.672 |
-2.575 |
-3.497 |
| y |
-2.575 |
-2.646 |
0.614 |
| z |
-3.497 |
0.614 |
-0.026 |
|
| Polar |
|---|
| 3z2-r2 | -0.052 |
|---|
| x2-y2 | 3.545 |
|---|
| xy | -2.575 |
|---|
| xz | -3.497 |
|---|
| yz | 0.614 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
132.137 |
| (<r2>)1/2 |
11.495 |