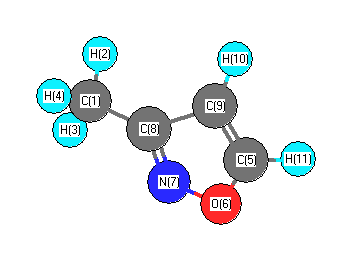
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3450 |
3121 |
3.99 |
|
|
|
| 2 |
A |
3426 |
3098 |
0.65 |
|
|
|
| 3 |
A |
3291 |
2976 |
15.84 |
|
|
|
| 4 |
A |
3249 |
2939 |
28.01 |
|
|
|
| 5 |
A |
3197 |
2891 |
23.56 |
|
|
|
| 6 |
A |
1813 |
1639 |
41.34 |
|
|
|
| 7 |
A |
1651 |
1493 |
22.52 |
|
|
|
| 8 |
A |
1620 |
1465 |
7.39 |
|
|
|
| 9 |
A |
1616 |
1462 |
7.93 |
|
|
|
| 10 |
A |
1580 |
1429 |
62.92 |
|
|
|
| 11 |
A |
1540 |
1392 |
7.51 |
|
|
|
| 12 |
A |
1406 |
1272 |
6.81 |
|
|
|
| 13 |
A |
1268 |
1147 |
24.20 |
|
|
|
| 14 |
A |
1193 |
1079 |
8.07 |
|
|
|
| 15 |
A |
1169 |
1057 |
1.73 |
|
|
|
| 16 |
A |
1132 |
1024 |
3.00 |
|
|
|
| 17 |
A |
1066 |
964 |
49.66 |
|
|
|
| 18 |
A |
1051 |
951 |
3.31 |
|
|
|
| 19 |
A |
1032 |
933 |
0.79 |
|
|
|
| 20 |
A |
999 |
903 |
8.03 |
|
|
|
| 21 |
A |
874 |
790 |
52.52 |
|
|
|
| 22 |
A |
729 |
659 |
0.75 |
|
|
|
| 23 |
A |
703 |
636 |
8.82 |
|
|
|
| 24 |
A |
653 |
591 |
8.76 |
|
|
|
| 25 |
A |
360 |
325 |
7.39 |
|
|
|
| 26 |
A |
300 |
271 |
5.61 |
|
|
|
| 27 |
A |
139 |
126 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20252.9 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 18316.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.667 |
-0.257 |
|
-0.609 |
| 2 |
H |
0.231 |
0.071 |
|
0.162 |
| 3 |
H |
0.252 |
0.079 |
|
0.163 |
| 4 |
H |
0.231 |
0.071 |
|
0.162 |
| 5 |
C |
0.160 |
0.227 |
|
0.263 |
| 6 |
O |
-0.281 |
-0.122 |
|
-0.136 |
| 7 |
N |
-0.189 |
-0.469 |
|
-0.479 |
| 8 |
C |
0.323 |
0.617 |
|
0.785 |
| 9 |
C |
-0.543 |
-0.517 |
|
-0.682 |
| 10 |
H |
0.228 |
0.193 |
|
0.253 |
| 11 |
H |
0.254 |
0.108 |
|
0.118 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.543 |
3.217 |
0.000 |
3.263 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.521 |
3.195 |
0.001 |
3.237 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.469 |
2.344 |
0.000 |
| y |
2.344 |
-36.957 |
-0.000 |
| z |
0.000 |
-0.000 |
-36.538 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.279 |
2.344 |
0.000 |
| y |
2.344 |
-3.953 |
-0.000 |
| z |
0.000 |
-0.000 |
-3.326 |
|
| Polar |
|---|
| 3z2-r2 | -6.651 |
|---|
| x2-y2 | 7.488 |
|---|
| xy | 2.344 |
|---|
| xz | 0.000 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.644 |
-0.118 |
0.000 |
| y |
-0.118 |
6.963 |
0.000 |
| z |
0.000 |
0.000 |
4.268 |
<r2> (average value of r
2) Å
2
| <r2> |
134.761 |
| (<r2>)1/2 |
11.609 |