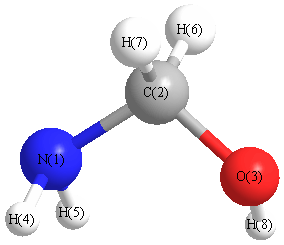
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3860 |
3860 |
11.78 |
|
|
|
| 2 |
A |
3624 |
3624 |
1.50 |
|
|
|
| 3 |
A |
3539 |
3539 |
0.41 |
|
|
|
| 4 |
A |
3155 |
3155 |
28.43 |
|
|
|
| 5 |
A |
3069 |
3069 |
58.22 |
|
|
|
| 6 |
A |
1724 |
1724 |
40.05 |
|
|
|
| 7 |
A |
1525 |
1525 |
0.16 |
|
|
|
| 8 |
A |
1455 |
1455 |
52.85 |
|
|
|
| 9 |
A |
1411 |
1411 |
0.55 |
|
|
|
| 10 |
A |
1398 |
1398 |
4.34 |
|
|
|
| 11 |
A |
1188 |
1188 |
35.48 |
|
|
|
| 12 |
A |
1129 |
1129 |
36.91 |
|
|
|
| 13 |
A |
1068 |
1068 |
237.53 |
|
|
|
| 14 |
A |
924 |
924 |
3.02 |
|
|
|
| 15 |
A |
858 |
858 |
174.20 |
|
|
|
| 16 |
A |
488 |
488 |
55.04 |
|
|
|
| 17 |
A |
431 |
431 |
130.57 |
|
|
|
| 18 |
A |
322 |
322 |
82.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15584.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15584.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.759 |
|
|
|
| 2 |
C |
-0.124 |
|
|
|
| 3 |
O |
-0.573 |
|
|
|
| 4 |
H |
0.326 |
|
|
|
| 5 |
H |
0.312 |
|
|
|
| 6 |
H |
0.211 |
|
|
|
| 7 |
H |
0.234 |
|
|
|
| 8 |
H |
0.372 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.284 |
-1.246 |
1.406 |
1.900 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.018 |
-1.467 |
-2.072 |
| y |
-1.467 |
-17.105 |
-1.471 |
| z |
-2.072 |
-1.471 |
-16.631 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.150 |
-1.467 |
-2.072 |
| y |
-1.467 |
3.219 |
-1.471 |
| z |
-2.072 |
-1.471 |
3.931 |
|
| Polar |
|---|
| 3z2-r2 | 7.862 |
|---|
| x2-y2 | -6.913 |
|---|
| xy | -1.467 |
|---|
| xz | -2.072 |
|---|
| yz | -1.471 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.564 |
-0.069 |
-0.080 |
| y |
-0.069 |
3.387 |
-0.185 |
| z |
-0.080 |
-0.185 |
3.381 |
<r2> (average value of r
2) Å
2
| <r2> |
49.492 |
| (<r2>)1/2 |
7.035 |