return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SO2NH2 (methanesulfonamide)

using model chemistry: QCISD(T)/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at QCISD(T)/6-311G*
 hartrees
Energy at 0K-643.372347
Energy at 298.15K-643.380183
HF Energy-642.463448
Nuclear repulsion energy277.295913
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3536 3404        
2 A' 3165 3048        
3 A' 3062 2948        
4 A' 1680 1617        
5 A' 1477 1422        
6 A' 1400 1348        
7 A' 1141 1098        
8 A' 1020 982        
9 A' 921 886        
10 A' 748 720        
11 A' 712 686        
12 A' 501 482        
13 A' 474 456        
14 A' 300 289        
15 A" 3641 3506        
16 A" 3176 3058        
17 A" 1476 1421        
18 A" 1370 1319        
19 A" 1143 1101        
20 A" 1002 965        
21 A" 404 389        
22 A" 332 320        
23 A" 231 223        
24 A" 176 170        

Unscaled Zero Point Vibrational Energy (zpe) 16543.1 cm-1
Scaled (by 0.9628) Zero Point Vibrational Energy (zpe) 15927.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/6-311G*
ABC
0.15604 0.14530 0.14276

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.672 -0.053 0.000
S2 0.110 -0.139 0.000
N3 0.530 1.491 0.000
O4 0.530 -0.699 1.275
O5 0.530 -0.699 -1.275
H6 -2.041 -1.083 0.000
H7 -1.998 0.472 0.901
H8 -1.998 0.472 -0.901
H9 1.058 1.713 0.837
H10 1.058 1.713 -0.837

Atom - Atom Distances (Å)
  C1 S2 N3 O4 O5 H6 H7 H8 H9 H10
C11.78452.68902.62492.62491.09391.09231.09233.35733.3573
S21.78451.68251.45441.45442.34962.37252.37252.24232.2423
N32.68901.68252.53392.53393.63762.87012.87011.01441.0144
O42.62491.45442.53392.54992.89522.81063.53462.50813.2496
O52.62491.45442.53392.54992.89523.53462.81063.24962.5081
H61.09392.34963.63762.89522.89521.79731.79734.25694.2569
H71.09232.37252.87012.81063.53461.79731.80153.29853.7278
H81.09232.37252.87013.53462.81061.79731.80153.72783.2985
H93.35732.24231.01442.50813.24964.25693.29853.72781.6744
H103.35732.24231.01443.24962.50814.25693.72783.29851.6744

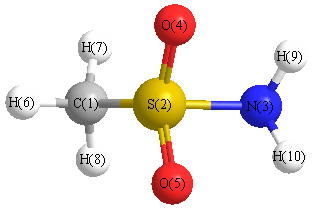
picture of methanesulfonamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 N3 101.680 C1 S2 O4 107.845
C1 S2 O5 107.845 S2 C1 H6 106.973
S2 C1 H7 108.723 S2 C1 H8 108.723
S2 N3 H9 110.013 S2 N3 H10 110.013
N3 S2 O4 107.538 N3 S2 O5 107.538
O4 S2 O5 122.468 H6 C1 H7 110.595
H6 C1 H8 110.595 H7 C1 H8 111.104
H9 N3 H10 111.232
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability