return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H5OO (ethylperoxy radical)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 2A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-229.006724
Energy at 298.15K-229.012754
HF Energy-228.297055
Nuclear repulsion energy124.715380
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3193 3024 13.44      
2 A' 3126 2961 14.43      
3 A' 3100 2936 9.25      
4 A' 1552 1470 7.13      
5 A' 1539 1457 2.82      
6 A' 1461 1384 21.81      
7 A' 1421 1345 1.60      
8 A' 1293 1224 125.21      
9 A' 1174 1112 8.81      
10 A' 1070 1014 15.63      
11 A' 885 838 2.72      
12 A' 527 499 11.41      
13 A' 319 302 1.69      
14 A" 3207 3037 29.14      
15 A" 3186 3018 0.34      
16 A" 1526 1445 9.34      
17 A" 1318 1248 0.29      
18 A" 1191 1128 5.36      
19 A" 821 777 1.22      
20 A" 241 228 0.41      
21 A" 75 71 0.23      

Unscaled Zero Point Vibrational Energy (zpe) 16111.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15259.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
1.11964 0.14972 0.13899

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 1.494 0.892 0.000
C2 0.000 0.677 0.000
O3 -0.219 -0.760 0.000
O4 -1.481 -1.043 0.000
H5 1.713 1.961 0.000
H6 1.948 0.447 0.886
H7 1.948 0.447 -0.886
H8 -0.486 1.077 -0.891
H9 -0.486 1.077 0.891

Atom - Atom Distances (Å)
  C1 C2 O3 O4 H5 H6 H7 H8 H9
C11.50942.38023.54891.09161.09071.09072.17892.1789
C21.50941.45392.26952.14132.15272.15271.09071.0907
O32.38021.45391.29343.33802.63462.63462.05922.0592
O43.54892.26951.29344.38503.84283.84282.50532.5053
H51.09162.14133.33804.38501.77021.77022.53222.5322
H61.09072.15272.63463.84281.77021.77213.07892.5144
H71.09072.15272.63463.84281.77021.77212.51443.0789
H82.17891.09072.05922.50532.53223.07892.51441.7819
H92.17891.09072.05922.50532.53222.51443.07891.7819

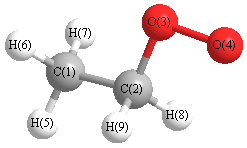
picture of ethylperoxy radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 106.860 C1 C2 H8 112.868
C1 C2 H9 112.868 C2 C1 H5 109.781
C2 C1 H6 110.746 C2 C1 H7 110.746
C2 O3 O4 111.265 O3 C2 H8 107.177
O3 C2 H9 107.177 H5 C1 H6 108.421
H5 C1 H7 108.421 H6 C1 H7 108.659
H8 C2 H9 109.535
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability