return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3SSSCH3 (dimethyl trisulfide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-1272.900458
Energy at 298.15K-1272.906797
Nuclear repulsion energy365.115566
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3204 3034 5.19      
2 A 3192 3023 0.02      
3 A 3090 2927 3.27      
4 A 1515 1435 13.20      
5 A 1498 1419 9.23      
6 A 1420 1345 0.03      
7 A 1029 975 6.12      
8 A 1024 970 3.46      
9 A 749 709 0.15      
10 A 488 462 0.44      
11 A 275 260 0.28      
12 A 208 197 1.02      
13 A 165 156 2.43      
14 A 65 61 2.35      
15 B 3204 3034 3.03      
16 B 3192 3023 5.13      
17 B 3090 2927 29.51      
18 B 1513 1433 14.26      
19 B 1498 1419 14.85      
20 B 1422 1347 1.68      
21 B 1029 975 2.10      
22 B 1019 965 23.78      
23 B 748 708 1.64      
24 B 482 457 14.80      
25 B 283 268 1.19      
26 B 205 194 0.92      
27 B 94 89 3.59      

Unscaled Zero Point Vibrational Energy (zpe) 17849.9 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 16905.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14995 0.04626 0.04587

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 1.078
S2 0.000 1.658 -0.161
S3 0.000 -1.658 -0.161
C4 1.741 1.717 -0.636
C5 -1.741 -1.717 -0.636
H6 1.870 2.593 -1.274
H7 -1.870 -2.593 -1.274
H8 2.378 1.814 0.242
H9 2.014 0.825 -1.198
H10 -2.378 -1.814 0.242
H11 -2.014 -0.825 -1.198

Atom - Atom Distances (Å)
  S1 S2 S3 C4 C5 H6 H7 H8 H9 H10 H11
S12.07022.07022.98662.98663.96973.96973.10523.14983.10523.1498
S22.07023.31591.80543.82742.36894.77592.41662.41384.22723.3612
S32.07023.31593.82741.80544.77592.36894.22723.36122.41662.4138
C42.98661.80543.82744.89091.09145.65941.08901.08975.49574.5696
C52.98663.82741.80544.89095.65941.09145.49574.56961.08901.0897
H63.96972.36894.77591.09145.65946.39451.77901.77566.30605.1749
H73.96974.77592.36895.65941.09146.39456.30605.17491.77901.7756
H83.10522.41664.22721.08905.49571.77906.30601.78455.98105.3223
H93.14982.41383.36121.08974.56961.77565.17491.78455.32234.3533
H103.10524.22722.41665.49571.08906.30601.77905.98105.32231.7845
H113.14983.36122.41384.56961.08975.17491.77565.32234.35331.7845

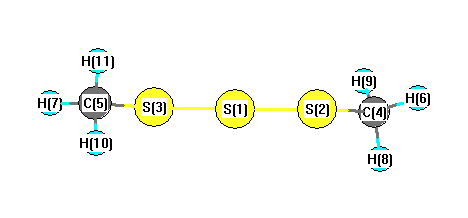
picture of dimethyl trisulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 S2 C4 100.595 S1 S3 C5 100.595
S2 S1 S3 106.424 S2 C4 H6 107.133
S2 C4 H8 110.779 S2 C4 H9 110.525
S3 C5 H7 107.133 S3 C5 H10 110.779
S3 C5 H11 110.525 H6 C4 H8 109.352
H6 C4 H9 108.995 H7 C5 H10 109.352
H7 C5 H11 108.995 H8 C4 H9 109.990
H10 C5 H11 109.990
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability