return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4F2 (orthodifluorobenzene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-429.837382
Energy at 298.15K-429.841946
Nuclear repulsion energy346.992446
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3247 3075 2.74      
2 A1 3233 3062 6.72      
3 A1 1667 1579 28.16      
4 A1 1563 1480 193.20      
5 A1 1461 1383 4.84      
6 A1 1317 1247 102.20      
7 A1 1188 1126 0.83      
8 A1 1054 998 7.64      
9 A1 777 736 32.28      
10 A1 579 548 3.58      
11 A1 288 272 0.35      
12 A2 864 819 0.00      
13 A2 816 773 0.00      
14 A2 600 569 0.00      
15 A2 447 424 0.00      
16 A2 184 174 0.00      
17 B1 875 829 11.01      
18 B1 755 715 98.11      
19 B1 458 434 3.56      
20 B1 295 279 0.03      
21 B2 3241 3070 2.68      
22 B2 3222 3052 0.84      
23 B2 1669 1581 8.18      
24 B2 1498 1419 14.15      
25 B2 1295 1227 4.88      
26 B2 1237 1172 33.71      
27 B2 1127 1068 24.80      
28 B2 861 815 16.61      
29 B2 552 523 3.31      
30 B2 439 416 0.28      

Unscaled Zero Point Vibrational Energy (zpe) 18405.0 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17431.4 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.10861 0.07398 0.04401

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.696 -0.538
C2 0.000 -0.696 -0.538
C3 0.000 -1.397 0.660
C4 0.000 -0.699 1.868
C5 0.000 0.699 1.868
C6 0.000 1.397 0.660
F7 0.000 1.351 -1.708
F8 0.000 -1.351 -1.708
H9 0.000 -2.482 0.626
H10 0.000 -1.247 2.804
H11 0.000 1.247 2.804
H12 0.000 2.482 0.626

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 H9 H10 H11 H12
C11.39112.41152.78052.40581.38821.34082.35713.38373.86553.38732.1318
C21.39111.38822.40582.78052.41152.35711.34082.13183.38733.86553.3837
C32.41151.38821.39582.41922.79503.62742.36801.08492.14983.40443.8795
C42.78052.40581.39581.39702.41924.12123.63462.17331.08502.15873.4143
C52.40582.78052.41921.39701.39583.63464.12123.41432.15871.08502.1733
C61.38822.41152.79502.41921.39582.36803.62743.87953.40442.14981.0849
F71.34082.35713.62744.12123.63462.36802.70154.48705.20624.51322.5931
F82.35711.34082.36803.63464.12123.62742.70152.59314.51325.20624.4870
H93.38372.13181.08492.17333.41433.87954.48702.59312.50454.31824.9636
H103.86553.38732.14981.08502.15873.40445.20624.51322.50452.49304.3182
H113.38733.86553.40442.15871.08502.14984.51325.20624.31822.49302.5045
H122.13183.38373.87953.41432.17331.08492.59314.48704.96364.31822.5045

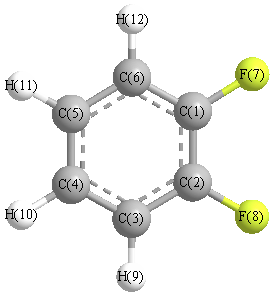
picture of orthodifluorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.373 C1 C2 F8 119.252
C1 C6 C5 119.575 C1 C6 H12 118.574
C2 C1 C6 120.373 C2 C1 F7 119.252
C2 C3 C4 119.575 C2 C3 H9 118.574
C3 C2 F8 120.375 C3 C4 C5 120.052
C3 C4 H10 119.611 C4 C3 H9 121.851
C4 C5 C6 120.052 C4 C5 H11 120.338
C5 C4 H10 120.338 C5 C6 H12 121.851
C6 C1 F7 120.375 C6 C5 H11 119.611
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability