return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2FCH2OH (2-fluoroethanol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-253.735628
Energy at 298.15K 
HF Energy-252.988849
Nuclear repulsion energy131.272763
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3844 3640 29.75 47.60 0.27 0.42
2 A 3189 3020 35.93 49.07 0.74 0.85
3 A 3168 3000 26.07 77.44 0.44 0.61
4 A 3114 2949 29.12 106.38 0.13 0.23
5 A 3068 2906 44.25 111.98 0.17 0.30
6 A 1547 1466 4.27 5.09 0.74 0.85
7 A 1537 1456 3.04 8.95 0.75 0.86
8 A 1474 1396 33.97 4.01 0.41 0.59
9 A 1446 1370 24.50 3.91 0.74 0.85
10 A 1421 1346 2.93 8.33 0.73 0.85
11 A 1304 1235 8.65 9.27 0.70 0.83
12 A 1257 1191 19.56 5.67 0.74 0.85
13 A 1162 1101 6.77 2.31 0.41 0.58
14 A 1139 1079 85.47 3.95 0.74 0.85
15 A 1085 1028 61.82 2.24 0.70 0.82
16 A 922 874 14.41 5.36 0.34 0.50
17 A 890 843 37.65 4.13 0.41 0.58
18 A 530 502 15.19 0.96 0.74 0.85
19 A 448 425 147.31 2.64 0.74 0.85
20 A 330 312 19.64 0.39 0.59 0.74
21 A 171 162 12.18 0.04 0.30 0.46

Unscaled Zero Point Vibrational Energy (zpe) 16522.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15648.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.52517 0.18520 0.15382

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.682 0.579 0.284
C2 -0.713 0.563 -0.283
O3 1.439 -0.517 -0.188
F4 -1.346 -0.602 0.160
H5 1.204 1.483 -0.035
H6 0.623 0.581 1.381
H7 -1.305 1.418 0.053
H8 -0.684 0.523 -1.374
H9 0.947 -1.310 0.039

Atom - Atom Distances (Å)
  C1 C2 O3 F4 H5 H6 H7 H8 H9
C11.50671.41352.35081.09171.09782.16972.14921.9236
C21.50672.41021.39772.14132.13391.09271.09142.5235
O31.41352.41022.80872.01982.08193.36662.64480.9613
F42.35081.39772.80873.30012.60132.02252.01372.4030
H51.09172.14132.01983.30011.77672.51192.50572.8065
H61.09782.13392.08192.60131.77672.48593.04902.3414
H72.16971.09273.36662.02252.51192.48591.79493.5373
H82.14921.09142.64482.01372.50573.04901.79492.8310
H91.92362.52350.96132.40302.80652.34143.53732.8310

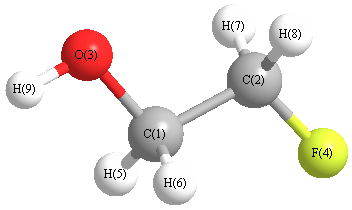
picture of 2-fluoroethanol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 F4 108.008 C1 C2 H7 112.183
C1 C2 H8 110.606 C1 O3 H9 106.642
C2 C1 O3 111.206 C2 C1 H5 109.956
C2 C1 H6 109.016 O3 C1 H5 106.756
O3 C1 H6 111.373
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability