return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C8H6 (benzocyclobutadiene)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-307.590736
Energy at 298.15K-307.595676
HF Energy-306.384645
Nuclear repulsion energy311.683135
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3266 3093 10.96      
2 A1 3231 3060 15.82      
3 A1 3215 3045 7.72      
4 A1 1703 1612 2.09      
5 A1 1540 1459 1.16      
6 A1 1521 1441 2.20      
7 A1 1449 1373 12.08      
8 A1 1197 1134 0.05      
9 A1 1131 1071 0.03      
10 A1 1071 1014 1.24      
11 A1 1003 950 3.62      
12 A1 817 774 0.45      
13 A1 546 517 0.04      
14 A2 843 798 0.00      
15 A2 836 792 0.00      
16 A2 762 722 0.00      
17 A2 503 477 0.00      
18 A2 301 285 0.00      
19 A2 256 243 0.00      
20 B1 816 773 7.12      
21 B1 717 679 102.67      
22 B1 657 623 30.64      
23 B1 327 310 0.89      
24 B1 207 196 14.83      
25 B2 3236 3065 2.23      
26 B2 3223 3052 27.56      
27 B2 3204 3035 0.17      
28 B2 1650 1563 0.65      
29 B2 1473 1395 4.81      
30 B2 1309 1239 4.41      
31 B2 1251 1185 16.87      
32 B2 1117 1058 1.36      
33 B2 1050 994 1.38      
34 B2 866 821 0.57      
35 B2 651 616 1.44      
36 B2 417 395 3.82      

Unscaled Zero Point Vibrational Energy (zpe) 23680.8 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 22428.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.16015 0.07299 0.05014

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 1.447 -0.623
C2 0.000 0.690 -1.847
C3 0.000 -0.690 -1.847
C4 0.000 -1.447 -0.623
C5 0.000 -0.680 2.047
C6 0.000 0.680 2.047
C7 0.000 0.718 0.529
C8 0.000 -0.718 0.529
H9 0.000 2.533 -0.654
H10 0.000 1.226 -2.791
H11 0.000 -1.226 -2.791
H12 0.000 -2.533 -0.654
H13 0.000 -1.449 2.811
H14 0.000 1.449 2.811

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 H9 H10 H11 H12 H13 H14
C11.43852.46182.89323.41342.77831.36382.45191.08712.17903.44163.97994.49263.4349
C21.43851.37962.46184.12743.89362.37602.76142.19571.08642.13643.43665.12574.7196
C32.46181.37961.43853.89364.12742.76142.37603.43662.13641.08642.19574.71965.1257
C42.89322.46181.43852.77833.41342.45191.36383.97993.44162.17901.08713.43494.4926
C53.41344.12743.89362.77831.35932.06291.51824.19735.20014.86893.27571.08472.2618
C62.77833.89364.12743.41341.35931.51822.06293.27574.86895.20014.19732.26181.0847
C71.36382.37602.76142.45192.06291.51821.43512.16713.35923.84763.45943.14692.3967
C82.45192.76142.37601.36381.51822.06291.43513.45943.84763.35922.16712.39673.1469
H91.08712.19573.43663.97994.19733.27572.16713.45942.50514.32475.06655.27903.6311
H102.17901.08642.13643.44165.20014.86893.35923.84762.50512.45304.32476.20885.6071
H113.44162.13641.08642.17904.86895.20013.84763.35924.32472.45302.50515.60716.2088
H123.97993.43662.19571.08713.27574.19733.45942.16715.06654.32472.50513.63115.2790
H134.49265.12574.71963.43491.08472.26183.14692.39675.27906.20885.60713.63112.8981
H143.43494.71965.12574.49262.26181.08472.39673.14693.63115.60716.20885.27902.8981

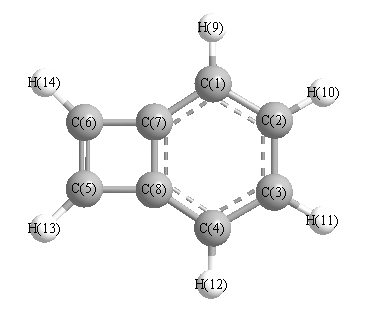
picture of benzocyclobutadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.744 C1 C2 H10 118.650
C1 C7 C6 149.114 C1 C7 N8 122.316
C2 C1 C7 115.941 C2 C1 H9 120.133
C2 C3 C4 121.744 C2 C3 H11 119.606
C3 C2 H10 119.606 C3 C4 N8 115.941
C3 C4 H12 120.133 C4 C3 H11 118.650
C4 N8 C5 149.114 C4 N8 C7 122.316
C5 C6 C7 91.429 C5 C6 H14 135.180
C5 N8 C7 88.571 C6 C5 N8 91.429
C6 C5 H13 135.180 C6 C7 N8 88.571
C7 C1 H9 123.926 C7 C6 H14 133.391
N8 C4 H12 123.926 N8 C5 H13 133.391
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability