return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H8Cl2 (Butane, 2,3-dichloro-, (r*,s*)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CI 1Ag
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-1076.193363
Energy at 298.15K-1076.202565
HF Energy-1075.180738
Nuclear repulsion energy370.898316
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3203 3034 0.00      
2 Ag 3190 3022 0.00      
3 Ag 3131 2966 0.00      
4 Ag 3098 2934 0.00      
5 Ag 1535 1454 0.00      
6 Ag 1526 1446 0.00      
7 Ag 1453 1376 0.00      
8 Ag 1417 1342 0.00      
9 Ag 1338 1268 0.00      
10 Ag 1215 1151 0.00      
11 Ag 1176 1114 0.00      
12 Ag 1068 1012 0.00      
13 Ag 890 843 0.00      
14 Ag 751 711 0.00      
15 Ag 493 467 0.00      
16 Ag 366 346 0.00      
17 Ag 300 284 0.00      
18 Ag 232 220 0.00      
19 Au 3204 3034 13.54      
20 Au 3191 3022 24.70      
21 Au 3146 2979 11.11      
22 Au 3097 2933 15.61      
23 Au 1536 1455 25.19      
24 Au 1522 1442 10.56      
25 Au 1453 1376 27.57      
26 Au 1344 1273 6.15      
27 Au 1284 1216 33.10      
28 Au 1127 1067 9.65      
29 Au 1057 1001 33.75      
30 Au 1000 947 11.84      
31 Au 716 678 58.02      
32 Au 377 357 2.05      
33 Au 343 325 1.93      
34 Au 263 249 2.61      
35 Au 205 194 1.75      
36 Au 62 59 3.20      

Unscaled Zero Point Vibrational Energy (zpe) 25654.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24297.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.12359 0.04814 0.03617

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Ci

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.951 1.175 -1.543
Cl2 0.951 -1.175 1.543
C3 -1.869 -0.438 0.408
C4 1.869 0.438 -0.408
C5 -0.652 0.394 0.051
C6 0.652 -0.394 -0.051
H7 -2.765 0.183 0.384
H8 2.765 -0.183 -0.384
H9 1.767 0.862 -1.406
H10 -1.767 -0.862 1.406
H11 -1.997 -1.251 -0.311
H12 1.997 1.251 0.311
H13 0.514 -1.209 -0.766
H14 -0.514 1.209 0.766

Atom - Atom Distances (Å)
  Cl1 Cl2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
Cl14.31982.69243.12831.79972.69382.82634.12312.73903.67562.91473.48322.90422.3494
Cl24.31983.12832.69242.69381.79974.12312.82633.67562.73903.48322.91472.34942.9042
C32.69243.12833.92551.51712.56281.09064.70884.26601.08981.09224.22022.76622.1627
C43.12832.69243.92552.56281.51714.70881.09061.08984.26604.22021.09222.16272.7662
C51.79972.69381.51712.56281.52642.15023.49272.86182.15792.15512.79612.14351.0929
C62.69381.79972.56281.51711.52643.49272.15022.15792.86182.79612.15511.09292.1435
H72.82634.12311.09064.70882.15023.49275.59604.91981.77061.76884.88123.74372.5033
H84.12312.82634.70881.09063.49272.15025.59601.77064.91984.88121.76882.50333.7437
H92.73903.67564.26601.08982.86182.15794.91981.77064.83364.45271.77572.50333.1685
H103.67562.73901.08984.26602.15792.86181.77064.91984.83361.77574.45273.16852.5033
H112.91473.48321.09224.22022.15512.79611.76884.88124.45271.77574.75352.55243.0671
H123.48322.91474.22021.09222.79612.15514.88121.76881.77574.45274.75353.06712.5524
H132.90422.34942.76622.16272.14351.09293.74372.50332.50333.16852.55243.06713.0410
H142.34942.90422.16272.76621.09292.14352.50333.74373.16852.50333.06712.55243.0410

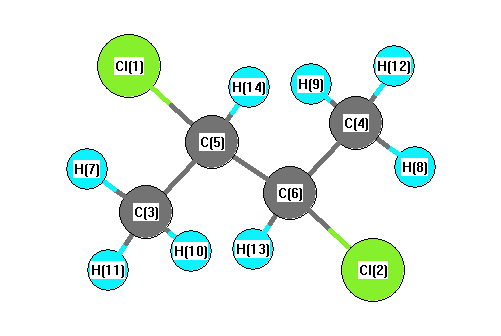
picture of Butane, 2,3-dichloro-, (r*,s*)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C5 C3 108.229 Cl1 C5 C6 107.889
Cl1 C5 H14 106.035 Cl2 C6 C4 108.229
Cl2 C6 C5 107.889 Cl2 C6 H13 106.035
C3 C5 C6 114.715 C3 C5 H14 110.863
C4 C6 C5 114.715 C4 C6 H13 110.863
C5 C3 H7 110.005 C5 C3 H10 110.673
C5 C3 H11 110.307 C5 C6 H13 108.712
C6 C4 H8 110.005 C6 C4 H9 110.673
C6 C4 H12 110.307 C6 C5 H14 108.712
H7 C3 H10 108.594 H7 C3 H11 108.264
H8 C4 H9 108.594 H8 C4 H12 108.264
H9 C4 H12 108.941 H10 C3 H11 108.941
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability