return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4N4 (1H-Tetrazole, 5-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-296.930638
Energy at 298.15K-296.938110
HF Energy-295.855221
Nuclear repulsion energy228.508363
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3694 3499 88.50      
2 A 3211 3041 0.94      
3 A 3094 2931 10.69      
4 A 1605 1520 41.58      
5 A 1542 1460 0.64      
6 A 1458 1381 17.61      
7 A 1451 1374 12.96      
8 A 1367 1295 9.23      
9 A 1211 1147 7.56      
10 A 1154 1093 1.34      
11 A 1130 1070 21.71      
12 A 1085 1027 14.68      
13 A 1023 969 0.68      
14 A 1017 963 0.83      
15 A 693 657 3.79      
16 A 337 319 4.72      
17 A 3173 3005 7.09      
18 A 1522 1442 12.76      
19 A 1086 1029 1.04      
20 A 737 698 25.48      
21 A 715 677 30.15      
22 A 632 599 45.49      
23 A 265 251 0.08      
24 A 86 81 0.59      

Unscaled Zero Point Vibrational Energy (zpe) 16643.6 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 15763.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.32524 0.12215 0.09030

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 -1.148 -0.062 0.000
C2 0.000 0.600 0.000
C3 0.181 2.076 0.000
H4 0.724 2.412 0.886
H5 0.724 2.412 -0.886
H6 -0.804 2.540 0.000
N7 0.988 -0.320 0.000
N8 0.466 -1.554 0.000
N9 -0.837 -1.381 0.000
H10 1.990 -0.203 0.000

Atom - Atom Distances (Å)
  N1 C2 C3 H4 H5 H6 N7 N8 N9 H10
N11.32552.51713.22633.22632.62532.15182.19761.35483.1413
C21.32551.48642.14232.14232.10021.35032.20412.15062.1460
C32.51711.48641.09231.09231.08892.52793.64093.60312.9095
H43.22632.14231.09231.77201.77122.88354.07154.19563.0367
H53.22632.14231.09231.77201.77122.88354.07154.19563.0367
H62.62532.10021.08891.77121.77123.37554.28693.92133.9157
N72.15181.35032.52792.88352.88353.37551.34042.11121.0086
N82.19762.20413.64094.07154.07154.28691.34041.31412.0372
N91.35482.15063.60314.19564.19563.92132.11121.31413.0627
H103.14132.14602.90953.03673.03673.91571.00862.03723.0627

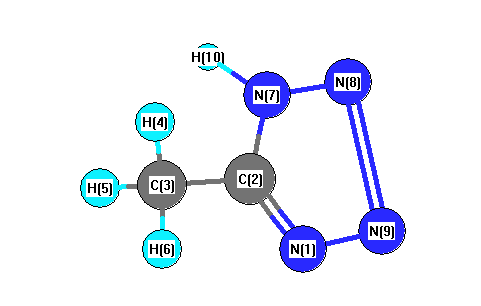
picture of 1H-Tetrazole, 5-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C3 126.964 N1 C2 N7 107.060
N1 N9 N8 110.853 C2 N1 N9 106.711
C2 C3 H4 111.445 C2 C3 H5 111.445
C2 C3 H6 108.290 C2 N7 N8 110.002
C2 N7 H10 130.381 C3 C2 N7 125.975
H4 C3 H5 108.413 H4 C3 H6 108.585
H5 C3 H6 108.585 N7 N8 N9 105.373
N8 N7 H10 119.617
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability