return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (Ethyl propyl sulfide)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-594.939687
Energy at 298.15K-594.952556
HF Energy-593.896396
Nuclear repulsion energy300.539770
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3174 3006 20.52      
2 A' 3169 3002 29.21      
3 A' 3091 2928 23.21      
4 A' 3087 2923 22.16      
5 A' 3085 2921 29.50      
6 A' 3078 2915 30.46      
7 A' 3070 2908 20.41      
8 A' 1547 1466 8.13      
9 A' 1543 1462 3.63      
10 A' 1532 1451 1.09      
11 A' 1527 1447 1.13      
12 A' 1517 1437 8.17      
13 A' 1452 1375 4.61      
14 A' 1450 1373 7.34      
15 A' 1412 1337 4.66      
16 A' 1355 1283 19.61      
17 A' 1308 1239 18.39      
18 A' 1149 1089 3.96      
19 A' 1098 1040 5.50      
20 A' 1089 1031 0.49      
21 A' 1029 974 4.42      
22 A' 936 887 2.61      
23 A' 808 765 0.68      
24 A' 730 692 0.28      
25 A' 397 376 0.24      
26 A' 313 296 0.78      
27 A' 282 267 0.72      
28 A' 111 105 0.27      
29 A" 3184 3015 21.54      
30 A" 3169 3001 48.97      
31 A" 3145 2979 2.16      
32 A" 3140 2974 14.79      
33 A" 3124 2959 5.61      
34 A" 1540 1459 10.75      
35 A" 1534 1452 11.32      
36 A" 1356 1284 0.17      
37 A" 1301 1232 0.19      
38 A" 1281 1213 0.15      
39 A" 1115 1056 3.79      
40 A" 1085 1028 0.01      
41 A" 898 850 0.60      
42 A" 815 772 7.47      
43 A" 769 728 4.08      
44 A" 266 252 0.08      
45 A" 253 239 0.02      
46 A" 123 117 1.01      
47 A" 67 63 0.39      
48 A" 50 48 0.34      

Unscaled Zero Point Vibrational Energy (zpe) 36276.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 34357.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.38309 0.03441 0.03253

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.376 2.249 0.000
H2 3.468 2.221 0.000
H3 2.051 2.801 0.884
H4 2.051 2.801 -0.884
C5 1.812 0.834 0.000
H6 2.153 0.292 -0.886
H7 2.153 0.292 0.886
S8 0.000 0.891 0.000
C9 -0.335 -0.888 0.000
H10 0.119 -1.341 -0.888
H11 0.119 -1.341 0.888
C12 -1.837 -1.147 0.000
H13 -2.285 -0.673 0.879
H14 -2.285 -0.673 -0.879
C15 -2.154 -2.640 0.000
H16 -1.738 -3.131 -0.884
H17 -1.738 -3.131 0.884
H18 -3.233 -2.814 0.000

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 S8 C9 H10 H11 C12 H13 H14 C15 H16 H17 H18
C11.09251.09171.09171.52332.16042.16042.73674.14684.33324.33325.41185.57105.57106.66556.82986.82987.5559
H21.09251.76791.76792.16012.49752.49753.71434.91294.96984.96986.28446.49976.49977.43267.51827.51828.3817
H31.09171.76791.76852.16973.07282.51142.93894.48224.90254.57115.61175.55625.82936.93367.25717.03857.7607
H41.09171.76791.76852.16972.51143.07282.93894.48224.57114.90255.61175.82935.55626.93367.03857.25717.7607
C51.52332.16012.16972.16971.09371.09371.81292.75302.89642.89644.15274.45314.45315.27295.39465.39466.2256
H62.16042.49753.07282.51141.09371.77262.40422.89332.60903.15494.33364.87284.54185.28545.18185.47586.2800
H72.16042.49752.51143.07281.09371.77262.40422.89333.15492.60904.33364.54184.87285.28545.47585.18186.2800
S82.73673.71432.93892.93891.81292.40422.40421.81072.40532.40532.74422.90522.90524.13654.46924.46924.9170
C94.14684.91294.48224.48222.75302.89332.89331.81071.09511.09511.52412.14942.14942.52532.78832.78833.4788
H104.33324.96984.90254.57112.89642.60903.15492.40531.09511.77522.15673.05692.49482.76412.57823.12803.7666
H114.33324.96984.57114.90252.89643.15492.60902.40531.09511.77522.15672.49483.05692.76413.12802.57823.7666
C125.41186.28445.61175.61174.15274.33364.33362.74421.52412.15672.15671.09451.09451.52632.17372.17372.1736
H135.57106.49975.55625.82934.45314.87284.54182.90522.14943.05692.49481.09451.75812.15873.07362.51792.5009
H145.57106.49975.82935.55624.45314.54184.87282.90522.14942.49483.05691.09451.75812.15872.51793.07362.5009
C156.66557.43266.93366.93365.27295.28545.28544.13652.52532.76412.76411.52632.15872.15871.09311.09311.0924
H166.82987.51827.25717.03855.39465.18185.47584.46922.78832.57823.12802.17373.07362.51791.09311.76751.7654
H176.82987.51827.03857.25715.39465.47585.18184.46922.78833.12802.57822.17372.51793.07361.09311.76751.7654
H187.55598.38177.76077.76076.22566.28006.28004.91703.47883.76663.76662.17362.50092.50091.09241.76541.7654

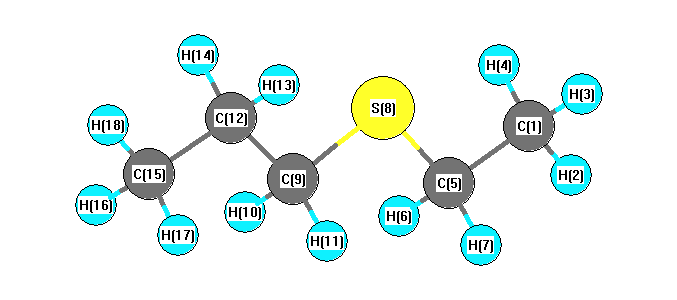
picture of Ethyl propyl sulfide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 110.205 C1 C5 H7 110.205
C1 C5 S8 109.929 H2 C1 H3 108.078
H2 C1 H4 108.078 H2 C1 C5 110.251
H3 C1 H4 108.191 H3 C1 C5 111.066
H4 C1 C5 111.066 C5 S8 C9 98.884
H6 C5 H7 108.262 H6 C5 S8 109.102
H7 C5 S8 109.102 S8 C9 H10 109.249
S8 C9 H11 109.249 S8 C9 C12 110.455
C9 C12 H13 109.233 C9 C12 H14 109.233
C9 C12 C15 111.757 H10 C9 H11 108.299
H10 C9 C12 109.775 H11 C9 C12 109.775
C12 C15 H16 111.083 C12 C15 H17 111.083
C12 C15 H18 111.119 H13 C12 H14 106.863
H13 C12 C15 109.814 H14 C12 C15 109.814
H16 C15 H17 107.888 H16 C15 H18 107.756
H17 C15 H18 107.756
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability