return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3NNCH3 ((Z)-1,2-Dimethyldiazene)

using model chemistry: B2PLYP=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at B2PLYP=FULL/6-311G*
 hartrees
Energy at 0K-189.113927
Energy at 298.15K 
HF Energy-188.885255
Nuclear repulsion energy122.125577
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A1 3182 3182 2.22      
2 A1 3047 3047 17.64      
3 A1 1627 1627 10.87      
4 A1 1504 1504 0.26      
5 A1 1436 1436 25.83      
6 A1 1113 1113 4.81      
7 A1 875 875 0.12      
8 A1 372 372 0.73      
9 A2 3117 3117 0.00      
10 A2 1515 1515 0.00      
11 A2 1103 1103 0.00      
12 A2 474 474 0.00      
13 A2 73i 73i 0.00      
14 B1 3114 3114 43.48      
15 B1 1541 1541 26.98      
16 B1 965 965 3.01      
17 B1 202 202 0.21      
18 B2 3181 3181 34.50      
19 B2 3045 3045 3.03      
20 B2 1495 1495 17.47      
21 B2 1418 1418 1.07      
22 B2 1194 1194 15.46      
23 B2 959 959 15.82      
24 B2 625 625 0.24      

Unscaled Zero Point Vibrational Energy (zpe) 18515.0 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 18515.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULL/6-311G*
ABC
0.54507 0.22430 0.16900

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULL/6-311G*

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.622 -0.782
N2 0.000 -0.622 -0.782
C3 0.000 1.358 0.502
C4 0.000 -1.358 0.502
H5 0.000 2.419 0.266
H6 0.000 -2.419 0.266
H7 -0.886 1.122 1.097
H8 0.886 1.122 1.097
H9 0.886 -1.122 1.097
H10 -0.886 -1.122 1.097

Atom - Atom Distances (Å)
  N1 N2 C3 C4 H5 H6 H7 H8 H9 H10
N11.24371.48032.36022.08013.21612.13732.13732.71282.7128
N21.24372.36021.48033.21612.08012.71282.71282.13732.1373
C31.48032.36022.71701.08623.78451.09371.09372.70072.7007
C42.36021.48032.71703.78451.08622.70072.70071.09371.0937
H52.08013.21611.08623.78454.83741.77711.77713.74363.7436
H63.21612.08013.78451.08624.83743.74363.74361.77711.7771
H72.13732.71281.09372.70071.77713.74361.77302.86022.2444
H82.13732.71281.09372.70071.77713.74361.77302.24442.8602
H92.71282.13732.70071.09373.74361.77712.86022.24441.7730
H102.71282.13732.70071.09373.74361.77712.24442.86021.7730

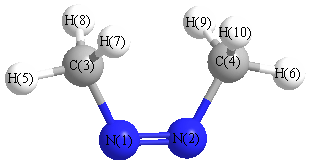
picture of (Z)-1,2-Dimethyldiazene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 N2 C4 119.842 N1 C3 H5 107.293
N1 C3 H7 111.386 N1 C3 H8 111.386
N2 N1 C3 119.842 N2 C4 H6 107.293
N2 C4 H9 111.386 N2 C4 H10 111.386
H5 C3 H7 109.218 H5 C3 H8 109.218
H6 C4 H9 109.218 H6 C4 H10 109.218
H7 C3 H8 108.302 H9 C4 H10 108.302
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability