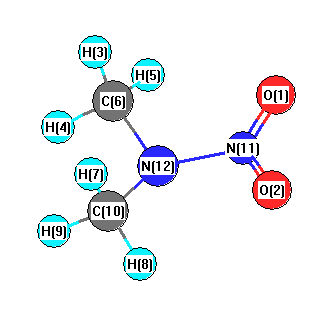
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3347 |
3027 |
12.07 |
|
|
|
| 2 |
A |
3343 |
3024 |
7.92 |
|
|
|
| 3 |
A |
3299 |
2983 |
59.35 |
|
|
|
| 4 |
A |
3287 |
2973 |
0.27 |
|
|
|
| 5 |
A |
3201 |
2895 |
60.31 |
|
|
|
| 6 |
A |
3195 |
2889 |
27.14 |
|
|
|
| 7 |
A |
1817 |
1644 |
604.76 |
|
|
|
| 8 |
A |
1659 |
1500 |
102.66 |
|
|
|
| 9 |
A |
1648 |
1491 |
40.20 |
|
|
|
| 10 |
A |
1637 |
1480 |
1.98 |
|
|
|
| 11 |
A |
1628 |
1472 |
28.61 |
|
|
|
| 12 |
A |
1622 |
1467 |
9.32 |
|
|
|
| 13 |
A |
1582 |
1431 |
3.53 |
|
|
|
| 14 |
A |
1541 |
1393 |
332.82 |
|
|
|
| 15 |
A |
1436 |
1299 |
0.00 |
|
|
|
| 16 |
A |
1416 |
1281 |
12.51 |
|
|
|
| 17 |
A |
1274 |
1152 |
39.07 |
|
|
|
| 18 |
A |
1238 |
1119 |
3.08 |
|
|
|
| 19 |
A |
1153 |
1043 |
18.93 |
|
|
|
| 20 |
A |
1144 |
1034 |
90.81 |
|
|
|
| 21 |
A |
955 |
864 |
11.65 |
|
|
|
| 22 |
A |
911 |
824 |
31.05 |
|
|
|
| 23 |
A |
697 |
630 |
4.82 |
|
|
|
| 24 |
A |
674 |
609 |
19.75 |
|
|
|
| 25 |
A |
461 |
417 |
4.41 |
|
|
|
| 26 |
A |
394 |
356 |
2.56 |
|
|
|
| 27 |
A |
249 |
225 |
8.42 |
|
|
|
| 28 |
A |
175 |
158 |
0.67 |
|
|
|
| 29 |
A |
165 |
149 |
4.42 |
|
|
|
| 30 |
A |
128 |
116 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 22637.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 20473.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.415 |
-0.526 |
|
-0.516 |
| 2 |
O |
-0.415 |
-0.526 |
|
-0.516 |
| 3 |
H |
0.227 |
0.102 |
|
0.171 |
| 4 |
H |
0.224 |
0.106 |
|
0.170 |
| 5 |
H |
0.263 |
0.102 |
|
0.166 |
| 6 |
C |
-0.449 |
-0.139 |
|
-0.395 |
| 7 |
H |
0.227 |
0.102 |
|
0.171 |
| 8 |
H |
0.263 |
0.102 |
|
0.166 |
| 9 |
H |
0.224 |
0.107 |
|
0.170 |
| 10 |
C |
-0.449 |
-0.140 |
|
-0.395 |
| 11 |
N |
0.638 |
0.991 |
|
0.954 |
| 12 |
N |
-0.339 |
-0.280 |
|
-0.146 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.801 |
0.001 |
0.211 |
4.806 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-4.834 |
0.001 |
0.234 |
4.839 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.853 |
0.001 |
-0.568 |
| y |
0.001 |
-36.114 |
0.001 |
| z |
-0.568 |
0.001 |
-35.054 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.269 |
0.001 |
-0.568 |
| y |
0.001 |
0.340 |
0.001 |
| z |
-0.568 |
0.001 |
1.929 |
|
| Polar |
|---|
| 3z2-r2 | 3.858 |
|---|
| x2-y2 | -1.739 |
|---|
| xy | 0.001 |
|---|
| xz | -0.568 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.661 |
0.000 |
0.064 |
| y |
0.000 |
7.160 |
-0.000 |
| z |
0.064 |
-0.000 |
4.331 |
<r2> (average value of r
2) Å
2
| <r2> |
145.537 |
| (<r2>)1/2 |
12.064 |