Jump to
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -147.945463 |
| Energy at 298.15K | -147.947909 |
| Nuclear repulsion energy | 60.141601 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3803 |
3440 |
43.61 |
|
|
|
| 2 |
A' |
2591 |
2344 |
131.49 |
|
|
|
| 3 |
A' |
1824 |
1650 |
56.35 |
|
|
|
| 4 |
A' |
1151 |
1041 |
5.77 |
|
|
|
| 5 |
A' |
701 |
634 |
249.81 |
|
|
|
| 6 |
A' |
547 |
495 |
69.60 |
|
|
|
| 7 |
A" |
3900 |
3527 |
56.90 |
|
|
|
| 8 |
A" |
1336 |
1208 |
0.18 |
|
|
|
| 9 |
A" |
472 |
426 |
1.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8162.7 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 7382.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.227 |
0.000 |
| N2 |
-0.065 |
1.357 |
0.000 |
| N3 |
0.123 |
-1.108 |
0.000 |
| H4 |
-0.203 |
-1.551 |
0.830 |
| H5 |
-0.203 |
-1.551 |
-0.830 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1316 | 1.3407 | 1.9721 | 1.9721 |
N2 | 1.1316 | | 2.4719 | 3.0264 | 3.0264 | N3 | 1.3407 | 2.4719 | | 0.9949 | 0.9949 | H4 | 1.9721 | 3.0264 | 0.9949 | | 1.6593 | H5 | 1.9721 | 3.0264 | 0.9949 | 1.6593 | |
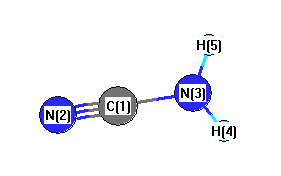 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
114.389 |
|
C1 |
N3 |
H5 |
114.389 |
| N2 |
C1 |
N3 |
178.034 |
|
H4 |
N3 |
H5 |
113.005 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.443 |
|
|
|
| 2 |
N |
-0.374 |
|
|
|
| 3 |
N |
-0.853 |
|
|
|
| 4 |
H |
0.392 |
|
|
|
| 5 |
H |
0.392 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.033 |
-4.514 |
0.000 |
4.631 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.532 |
2.212 |
0.000 |
| y |
2.212 |
-18.840 |
0.000 |
| z |
0.000 |
0.000 |
-14.982 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.621 |
2.212 |
0.000 |
| y |
2.212 |
-2.083 |
0.000 |
| z |
0.000 |
0.000 |
3.704 |
|
| Polar |
|---|
| 3z2-r2 | 7.408 |
|---|
| x2-y2 | 0.308 |
|---|
| xy | 2.212 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.762 |
-0.068 |
0.000 |
| y |
-0.068 |
4.457 |
0.000 |
| z |
0.000 |
0.000 |
2.139 |
<r2> (average value of r
2) Å
2
| <r2> |
39.090 |
| (<r2>)1/2 |
6.252 |
Jump to
S1C1
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -147.943666 |
| Energy at 298.15K | |
| HF Energy | -147.943666 |
| Nuclear repulsion energy | 60.315202 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3865 |
3496 |
72.77 |
|
|
|
| 2 |
A1 |
2578 |
2332 |
173.91 |
|
|
|
| 3 |
A1 |
1792 |
1621 |
61.68 |
|
|
|
| 4 |
A1 |
1193 |
1079 |
13.09 |
|
|
|
| 5 |
B1 |
606 |
548 |
5.55 |
|
|
|
| 6 |
B1 |
485i |
439i |
372.11 |
|
|
|
| 7 |
B2 |
3992 |
3611 |
95.17 |
|
|
|
| 8 |
B2 |
1260 |
1139 |
3.93 |
|
|
|
| 9 |
B2 |
466 |
421 |
0.06 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7633.4 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 6903.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.226 |
| N2 |
0.000 |
0.000 |
1.360 |
| N3 |
0.000 |
0.000 |
-1.097 |
| H4 |
0.000 |
0.854 |
-1.597 |
| H5 |
0.000 |
-0.854 |
-1.597 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1335 | 1.3235 | 2.0131 | 2.0131 |
N2 | 1.1335 | | 2.4569 | 3.0773 | 3.0773 | N3 | 1.3235 | 2.4569 | | 0.9894 | 0.9894 | H4 | 2.0131 | 3.0773 | 0.9894 | | 1.7080 | H5 | 2.0131 | 3.0773 | 0.9894 | 1.7080 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
120.322 |
|
C1 |
N3 |
H5 |
120.322 |
| N2 |
C1 |
N3 |
180.000 |
|
H4 |
N3 |
H5 |
119.356 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.466 |
|
|
|
| 2 |
N |
-0.381 |
|
|
|
| 3 |
N |
-0.900 |
|
|
|
| 4 |
H |
0.408 |
|
|
|
| 5 |
H |
0.408 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.913 |
4.913 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-18.680 |
0.000 |
0.000 |
| y |
0.000 |
-14.702 |
0.000 |
| z |
0.000 |
0.000 |
-18.068 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.295 |
0.000 |
0.000 |
| y |
0.000 |
3.672 |
0.000 |
| z |
0.000 |
0.000 |
-1.377 |
|
| Polar |
|---|
| 3z2-r2 | -2.754 |
|---|
| x2-y2 | -3.978 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.695 |
0.000 |
0.000 |
| y |
0.000 |
2.068 |
0.000 |
| z |
0.000 |
0.000 |
4.506 |
<r2> (average value of r
2) Å
2
| <r2> |
38.946 |
| (<r2>)1/2 |
6.241 |
