return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-217.863944
Energy at 298.15K-217.871885
HF Energy-217.169208
Nuclear repulsion energy132.830128
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3189 3021 28.50      
2 A' 3186 3017 39.76      
3 A' 3108 2943 30.69      
4 A' 3090 2927 6.69      
5 A' 1551 1469 11.44      
6 A' 1532 1451 6.79      
7 A' 1456 1379 26.31      
8 A' 1418 1343 12.23      
9 A' 1233 1168 10.79      
10 A' 1189 1126 48.60      
11 A' 988 936 36.15      
12 A' 849 804 10.01      
13 A' 488 462 3.38      
14 A' 362 343 0.87      
15 A' 276 261 0.18      
16 A" 3187 3019 14.41      
17 A" 3179 3011 0.00      
18 A" 3087 2924 12.66      
19 A" 1525 1444 0.04      
20 A" 1520 1439 0.21      
21 A" 1452 1375 41.68      
22 A" 1409 1334 1.00      
23 A" 1199 1136 12.30      
24 A" 965 914 0.00      
25 A" 951 901 0.75      
26 A" 420 397 7.16      
27 A" 225 214 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 21516.3 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 20378.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.29170 0.27131 0.16061

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.282 0.244 0.000
F2 -0.874 1.035 0.000
H3 1.126 0.941 0.000
C4 0.282 -0.582 1.267
C5 0.282 -0.582 -1.267
H6 1.189 -1.187 1.334
H7 1.189 -1.187 -1.334
H8 0.230 0.067 2.143
H9 0.230 0.067 -2.143
H10 -0.584 -1.248 1.278
H11 -0.584 -1.248 -1.278

Atom - Atom Distances (Å)
  C1 F2 H3 C4 C5 H6 H7 H8 H9 H10 H11
C11.40091.09511.51221.51222.15592.15592.15042.15042.14652.1465
F21.40092.00252.35732.35733.31253.31252.59772.59772.63232.6323
H31.09512.00252.15342.15342.51162.51162.48132.48133.05753.0575
C41.51222.35732.15342.53371.09242.81991.09123.47101.09222.7696
C51.51222.35732.15342.53372.81991.09243.47101.09122.76961.0922
H62.15593.31252.51161.09242.81992.66751.77343.81771.77543.1578
H72.15593.31252.51162.81991.09242.66753.81771.77343.15781.7754
H82.15042.59772.48131.09123.47101.77343.81774.28511.77163.7541
H92.15042.59772.48133.47101.09123.81771.77344.28513.75411.7716
H102.14652.63233.05751.09222.76961.77543.15781.77163.75412.5565
H112.14652.63233.05752.76961.09223.15781.77543.75411.77162.5565

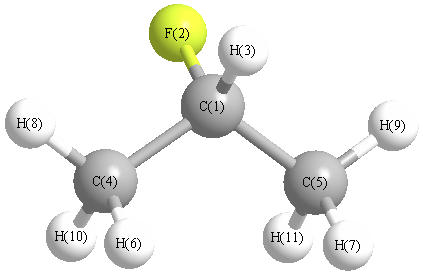
picture of 2-Fluoropropane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C4 H8 110.337 C1 C4 H10 109.962
C1 C5 H7 110.696 C1 C5 H9 110.337
C1 C5 H11 109.962 F2 C1 H3 106.052
F2 C1 C4 107.975 F2 C1 C5 107.975
H3 C1 C4 110.338 H3 C1 C5 110.338
C4 C1 C5 113.811 H7 C5 H9 108.607
H7 C5 H11 108.720 H8 C4 H10 108.463
H9 C5 H11 108.463
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability