Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -154.337951 |
| Energy at 298.15K | |
| HF Energy | -154.337951 |
| Nuclear repulsion energy | 74.793164 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3791 |
3650 |
3.23 |
|
|
|
| 2 |
A' |
3150 |
3033 |
12.74 |
|
|
|
| 3 |
A' |
2983 |
2872 |
75.28 |
|
|
|
| 4 |
A' |
1531 |
1474 |
0.37 |
|
|
|
| 5 |
A' |
1489 |
1433 |
6.45 |
|
|
|
| 6 |
A' |
1443 |
1389 |
2.67 |
|
|
|
| 7 |
A' |
1247 |
1201 |
64.77 |
|
|
|
| 8 |
A' |
1062 |
1022 |
81.21 |
|
|
|
| 9 |
A' |
1001 |
963 |
43.61 |
|
|
|
| 10 |
A' |
622 |
599 |
20.92 |
|
|
|
| 11 |
A' |
376 |
362 |
36.19 |
|
|
|
| 12 |
A" |
3256 |
3135 |
17.23 |
|
|
|
| 13 |
A" |
3015 |
2902 |
67.84 |
|
|
|
| 14 |
A" |
1297 |
1249 |
0.28 |
|
|
|
| 15 |
A" |
1174 |
1130 |
1.12 |
|
|
|
| 16 |
A" |
814 |
784 |
0.22 |
|
|
|
| 17 |
A" |
265 |
255 |
146.11 |
|
|
|
| 18 |
A" |
148i |
143i |
1.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14183.6 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 13654.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.111 |
-0.369 |
0.000 |
| C2 |
0.000 |
0.534 |
0.000 |
| C3 |
1.259 |
-0.250 |
0.000 |
| H4 |
-1.916 |
0.160 |
0.000 |
| H5 |
-0.036 |
1.182 |
0.888 |
| H6 |
-0.036 |
1.182 |
-0.888 |
| H7 |
1.662 |
-0.640 |
-0.928 |
| H8 |
1.662 |
-0.640 |
0.928 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4317 | 2.3728 | 0.9630 | 2.0862 | 2.0862 | 2.9367 | 2.9367 |
C2 | 1.4317 | | 1.4829 | 1.9518 | 1.1003 | 1.1003 | 2.2362 | 2.2362 | C3 | 2.3728 | 1.4829 | | 3.2008 | 2.1251 | 2.1251 | 1.0842 | 1.0842 | H4 | 0.9630 | 1.9518 | 3.2008 | | 2.3169 | 2.3169 | 3.7816 | 3.7816 | H5 | 2.0862 | 1.1003 | 2.1251 | 2.3169 | | 1.7763 | 3.0822 | 2.4907 | H6 | 2.0862 | 1.1003 | 2.1251 | 2.3169 | 1.7763 | | 2.4907 | 3.0822 | H7 | 2.9367 | 2.2362 | 1.0842 | 3.7816 | 3.0822 | 2.4907 | | 1.8558 | H8 | 2.9367 | 2.2362 | 1.0842 | 3.7816 | 2.4907 | 3.0822 | 1.8558 | |
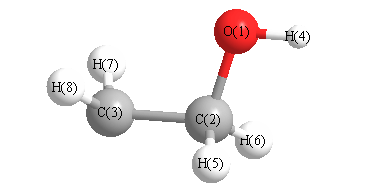 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
108.987 |
|
O1 |
C2 |
H5 |
110.276 |
| O1 |
C2 |
H6 |
110.276 |
|
C2 |
O1 |
H4 |
107.565 |
| C2 |
C3 |
H7 |
120.379 |
|
C2 |
C3 |
H8 |
120.379 |
| C3 |
C2 |
H5 |
109.819 |
|
C3 |
C2 |
H6 |
109.819 |
| H5 |
C2 |
H6 |
107.653 |
|
H7 |
C3 |
H8 |
117.708 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.568 |
|
|
|
| 2 |
C |
-0.233 |
|
|
|
| 3 |
C |
-0.420 |
|
|
|
| 4 |
H |
0.383 |
|
|
|
| 5 |
H |
0.195 |
|
|
|
| 6 |
H |
0.195 |
|
|
|
| 7 |
H |
0.224 |
|
|
|
| 8 |
H |
0.224 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.262 |
1.599 |
0.000 |
1.621 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.628 |
-2.966 |
0.000 |
| y |
-2.966 |
-20.168 |
0.000 |
| z |
0.000 |
0.000 |
-18.733 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.822 |
-2.966 |
0.000 |
| y |
-2.966 |
-2.487 |
0.000 |
| z |
0.000 |
0.000 |
-0.335 |
|
| Polar |
|---|
| 3z2-r2 | -0.670 |
|---|
| x2-y2 | 3.539 |
|---|
| xy | -2.966 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.561 |
-0.474 |
0.000 |
| y |
-0.474 |
3.497 |
0.000 |
| z |
0.000 |
0.000 |
3.519 |
<r2> (average value of r
2) Å
2
| <r2> |
50.255 |
| (<r2>)1/2 |
7.089 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -154.339636 |
| Energy at 298.15K | -154.344311 |
| HF Energy | -154.339636 |
| Nuclear repulsion energy | 75.130677 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3826 |
3683 |
10.39 |
|
|
|
| 2 |
A |
3275 |
3153 |
9.52 |
|
|
|
| 3 |
A |
3162 |
3044 |
7.72 |
|
|
|
| 4 |
A |
2953 |
2843 |
74.63 |
|
|
|
| 5 |
A |
2887 |
2779 |
78.65 |
|
|
|
| 6 |
A |
1510 |
1454 |
6.44 |
|
|
|
| 7 |
A |
1476 |
1421 |
4.82 |
|
|
|
| 8 |
A |
1427 |
1374 |
0.50 |
|
|
|
| 9 |
A |
1287 |
1239 |
75.03 |
|
|
|
| 10 |
A |
1234 |
1188 |
10.03 |
|
|
|
| 11 |
A |
1131 |
1089 |
48.13 |
|
|
|
| 12 |
A |
1078 |
1038 |
29.84 |
|
|
|
| 13 |
A |
970 |
933 |
12.28 |
|
|
|
| 14 |
A |
869 |
837 |
15.09 |
|
|
|
| 15 |
A |
456 |
439 |
29.71 |
|
|
|
| 16 |
A |
402 |
387 |
46.90 |
|
|
|
| 17 |
A |
299 |
288 |
117.17 |
|
|
|
| 18 |
A |
148 |
142 |
21.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14193.7 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 13664.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-1.107 |
-0.377 |
-0.057 |
| C2 |
-0.011 |
0.517 |
0.037 |
| C3 |
1.248 |
-0.252 |
-0.022 |
| H4 |
-1.916 |
0.119 |
0.094 |
| H5 |
-0.062 |
1.090 |
0.984 |
| H6 |
-0.034 |
1.267 |
-0.773 |
| H7 |
2.189 |
0.247 |
-0.222 |
| H8 |
1.253 |
-1.290 |
0.287 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| O1 | | 1.4171 | 2.3587 | 0.9611 | 2.0810 | 2.0898 | 3.3583 | 2.5534 |
C2 | 1.4171 | | 1.4771 | 1.9461 | 1.1090 | 1.1040 | 2.2319 | 2.2195 | C3 | 2.3587 | 1.4771 | | 3.1878 | 2.1288 | 2.1254 | 1.0834 | 1.0830 | H4 | 0.9611 | 1.9461 | 3.1878 | | 2.2741 | 2.3685 | 4.1187 | 3.4732 | H5 | 2.0810 | 1.1090 | 2.1288 | 2.2741 | | 1.7663 | 2.6898 | 2.8074 | H6 | 2.0898 | 1.1040 | 2.1254 | 2.3685 | 1.7663 | | 2.5073 | 3.0529 | H7 | 3.3583 | 2.2319 | 1.0834 | 4.1187 | 2.6898 | 2.5073 | | 1.8702 | H8 | 2.5534 | 2.2195 | 1.0830 | 3.4732 | 2.8074 | 3.0529 | 1.8702 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
109.158 |
|
O1 |
C2 |
H5 |
110.343 |
| O1 |
C2 |
H6 |
111.374 |
|
C2 |
O1 |
H4 |
108.309 |
| C2 |
C3 |
H7 |
120.535 |
|
C2 |
C3 |
H8 |
119.419 |
| C3 |
C2 |
H5 |
109.995 |
|
C3 |
C2 |
H6 |
110.027 |
| H5 |
C2 |
H6 |
105.908 |
|
H7 |
C3 |
H8 |
119.377 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.574 |
|
|
|
| 2 |
C |
-0.212 |
|
|
|
| 3 |
C |
-0.454 |
|
|
|
| 4 |
H |
0.389 |
|
|
|
| 5 |
H |
0.198 |
|
|
|
| 6 |
H |
0.201 |
|
|
|
| 7 |
H |
0.217 |
|
|
|
| 8 |
H |
0.235 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.361 |
1.788 |
0.489 |
1.888 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.773 |
-1.883 |
-0.655 |
| y |
-1.883 |
-19.289 |
-0.502 |
| z |
-0.655 |
-0.502 |
-20.328 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.036 |
-1.883 |
-0.655 |
| y |
-1.883 |
-1.239 |
-0.502 |
| z |
-0.655 |
-0.502 |
-2.797 |
|
| Polar |
|---|
| 3z2-r2 | -5.593 |
|---|
| x2-y2 | 3.516 |
|---|
| xy | -1.883 |
|---|
| xz | -0.655 |
|---|
| yz | -0.502 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.613 |
-0.265 |
-0.155 |
| y |
-0.265 |
3.940 |
-0.084 |
| z |
-0.155 |
-0.084 |
3.002 |
<r2> (average value of r
2) Å
2
| <r2> |
50.120 |
| (<r2>)1/2 |
7.080 |
