return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H12N2 (1,2-Butanediamine)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-268.412558
Energy at 298.15K-268.427471
HF Energy-267.392711
Nuclear repulsion energy264.063018
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3648 3455 3.49      
2 A 3641 3448 0.51      
3 A 3549 3361 3.28      
4 A 3543 3355 0.90      
5 A 3174 3006 32.43      
6 A 3163 2995 31.31      
7 A 3149 2983 11.34      
8 A 3112 2947 40.62      
9 A 3093 2929 14.53      
10 A 3072 2910 27.10      
11 A 2993 2835 124.29      
12 A 2978 2820 51.14      
13 A 1737 1645 54.09      
14 A 1722 1631 62.96      
15 A 1553 1471 10.64      
16 A 1552 1470 6.10      
17 A 1538 1456 7.19      
18 A 1519 1438 2.92      
19 A 1465 1388 1.13      
20 A 1456 1379 7.76      
21 A 1448 1371 4.55      
22 A 1421 1346 9.98      
23 A 1371 1298 0.38      
24 A 1334 1264 0.89      
25 A 1319 1249 2.39      
26 A 1286 1218 1.46      
27 A 1221 1156 7.45      
28 A 1185 1123 8.28      
29 A 1134 1074 3.39      
30 A 1100 1042 5.11      
31 A 1097 1039 3.15      
32 A 1040 985 0.69      
33 A 1012 958 36.63      
34 A 963 912 50.09      
35 A 952 902 52.81      
36 A 900 853 131.01      
37 A 845 800 18.19      
38 A 780 739 0.72      
39 A 636 603 20.60      
40 A 456 432 5.59      
41 A 395 374 3.42      
42 A 346 328 38.45      
43 A 287 272 54.31      
44 A 282 267 8.40      
45 A 229 217 20.10      
46 A 219 207 2.32      
47 A 155 147 6.25      
48 A 72 68 1.94      

Unscaled Zero Point Vibrational Energy (zpe) 37569.2 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 35581.8 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.18796 0.07439 0.06433

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 1.927 -0.627 -0.655
H2 1.806 0.155 -1.288
H3 2.915 -0.842 -0.603
C4 -2.394 -0.332 -0.116
H5 -3.033 -1.170 -0.402
H6 -2.741 0.033 0.856
H7 -2.561 0.456 -0.854
N8 0.025 1.485 -0.335
H9 0.505 2.239 0.146
H10 -0.917 1.808 -0.516
C11 -0.929 -0.761 -0.056
H12 -0.839 -1.659 0.566
H13 -0.564 -1.032 -1.049
C14 1.431 -0.212 0.653
H15 2.036 0.582 1.128
H16 1.445 -1.071 1.333
C17 -0.001 0.298 0.529
H18 -0.356 0.533 1.549

Atom - Atom Distances (Å)
  N1 H2 H3 C4 H5 H6 H7 N8 H9 H10 C11 H12 H13 C14 H15 H16 C17 H18
N11.01351.01234.36394.99574.95074.62072.86013.29763.74642.92113.19472.55381.45922.15702.09292.44443.3782
H21.01351.64144.38695.09435.02864.39822.41822.84403.27733.13633.70452.66092.01072.46422.91572.56653.5864
H31.01231.64145.35485.95975.90615.63283.71993.98224.65973.88314.01533.51192.04322.40772.44103.32924.1492
C44.36394.38695.35481.09261.09451.09323.03283.88262.63001.52742.15452.17003.90334.69034.16852.55622.7690
H54.99575.09435.95971.09261.76511.75304.05024.94313.65452.17092.44732.55604.68645.57684.80313.49463.7242
H64.95075.02865.90611.09451.76511.77133.34363.98822.89142.17882.56193.08274.18494.81604.35542.77212.5335
H74.62074.39825.63281.09321.75301.77132.83083.68432.15432.18693.07512.49804.31945.00724.81252.91363.2620
N82.86012.41823.71993.03284.05023.34362.83081.01461.01232.45653.38262.68142.41552.64563.36601.46852.1450
H93.29762.84403.98223.88264.94313.98823.68431.01461.62693.33114.14363.64192.66822.45993.63902.04162.3704
H103.74643.27734.65972.63003.65452.89142.15431.01231.62692.61003.63232.91053.31103.59534.15752.05202.4911
C112.92113.13633.88311.52742.17092.17882.18692.45653.33112.61001.09571.09162.52533.46352.76771.52492.1396
H123.19473.70454.01532.15452.44732.56193.07513.38264.14363.63231.09571.75422.69373.68772.47972.12872.4492
H132.55382.66093.51192.17002.55603.08272.49802.68143.64192.91051.09161.75422.74773.75493.11562.13853.0391
C141.45922.01072.04323.90334.68644.18494.31942.41552.66823.31102.52532.69372.74771.10481.09531.52592.1333
H152.15702.46422.40774.69035.57684.81605.00722.64562.45993.59533.46353.68773.75491.10481.76712.14222.4289
H162.09292.91572.44104.16854.80314.35544.81253.36603.63904.15752.76772.47973.11561.09531.76712.14742.4208
C172.44442.56653.32922.55623.49462.77212.91361.46852.04162.05201.52492.12872.13851.52592.14222.14741.1048
H183.37823.58644.14922.76903.72422.53353.26202.14502.37042.49112.13962.44923.03912.13332.42892.42081.1048

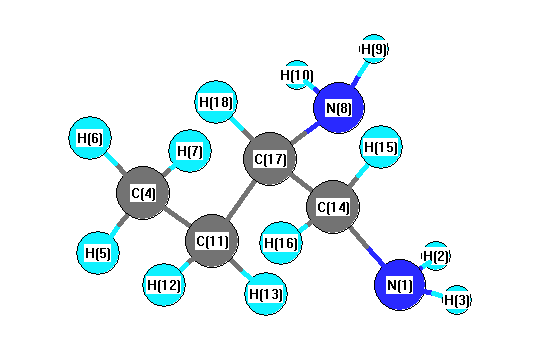
picture of 1,2-Butanediamine state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C14 H15 113.838 N1 C14 H16 109.203
N1 C14 C17 109.925 H2 N1 H3 108.240
H2 N1 C14 107.446 H3 N1 C14 110.214
C4 C11 H12 109.338 C4 C11 H13 110.805
C4 C11 C17 113.753 H5 C4 H6 107.621
H5 C4 H7 106.640 H5 C4 C11 110.816
H6 C4 H7 108.129 H6 C4 C11 111.340
H7 C4 C11 112.068 N8 C17 C11 110.288
N8 C17 C14 107.528 N8 C17 H18 112.163
H9 N8 H10 106.773 H9 N8 C17 109.249
H10 N8 C17 110.263 C11 C17 C14 111.741
C11 C17 H18 107.835 H12 C11 H13 106.638
H12 C11 C17 107.512 H13 C11 C17 108.503
C14 C17 H18 107.291 H15 C14 H16 106.867
H15 C14 C17 107.965 H16 C14 C17 108.905
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability