Jump to
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -243.884457 |
| Energy at 298.15K | |
| HF Energy | -243.884457 |
| Nuclear repulsion energy | 125.561022 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4158 |
3761 |
105.88 |
|
|
|
| 2 |
A' |
4016 |
3632 |
70.52 |
|
|
|
| 3 |
A' |
3879 |
3509 |
60.58 |
|
|
|
| 4 |
A' |
2002 |
1811 |
739.62 |
|
|
|
| 5 |
A' |
1799 |
1627 |
147.69 |
|
|
|
| 6 |
A' |
1571 |
1421 |
186.16 |
|
|
|
| 7 |
A' |
1376 |
1244 |
251.79 |
|
|
|
| 8 |
A' |
1194 |
1080 |
54.61 |
|
|
|
| 9 |
A' |
1059 |
958 |
21.84 |
|
|
|
| 10 |
A' |
650 |
588 |
49.54 |
|
|
|
| 11 |
A' |
534 |
483 |
6.47 |
|
|
|
| 12 |
A" |
889 |
804 |
78.85 |
|
|
|
| 13 |
A" |
639 |
578 |
84.49 |
|
|
|
| 14 |
A" |
549 |
496 |
81.25 |
|
|
|
| 15 |
A" |
144i |
130i |
310.76 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12086.0 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 10930.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
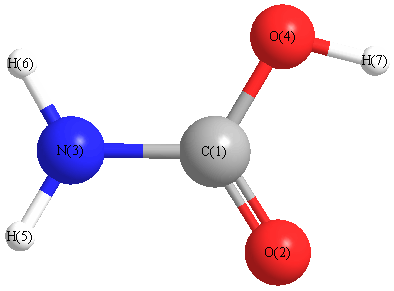
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.128 |
0.000 |
| O2 |
-0.044 |
1.314 |
0.000 |
| N3 |
1.119 |
-0.617 |
0.000 |
| O4 |
-1.088 |
-0.640 |
0.000 |
| H5 |
1.994 |
-0.157 |
0.000 |
| H6 |
1.077 |
-1.605 |
0.000 |
| H7 |
-1.844 |
-0.078 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.1878 | 1.3439 | 1.3317 | 2.0144 | 2.0396 | 1.8552 |
O2 | 1.1878 | | 2.2546 | 2.2160 | 2.5142 | 3.1270 | 2.2751 | N3 | 1.3439 | 2.2546 | | 2.2070 | 0.9888 | 0.9887 | 3.0113 | O4 | 1.3317 | 2.2160 | 2.2070 | | 3.1199 | 2.3698 | 0.9422 | H5 | 2.0144 | 2.5142 | 0.9888 | 3.1199 | | 1.7140 | 3.8388 | H6 | 2.0396 | 3.1270 | 0.9887 | 2.3698 | 1.7140 | | 3.2956 | H7 | 1.8552 | 2.2751 | 3.0113 | 0.9422 | 3.8388 | 3.2956 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.646 |
|
C1 |
N3 |
H6 |
121.192 |
| C1 |
O4 |
H7 |
108.119 |
|
O2 |
C1 |
N3 |
125.778 |
| O2 |
C1 |
O4 |
123.073 |
|
N3 |
C1 |
O4 |
111.149 |
| H5 |
N3 |
H6 |
120.162 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.785 |
|
|
|
| 2 |
O |
-0.505 |
|
|
|
| 3 |
N |
-0.883 |
|
|
|
| 4 |
O |
-0.635 |
|
|
|
| 5 |
H |
0.394 |
|
|
|
| 6 |
H |
0.392 |
|
|
|
| 7 |
H |
0.450 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.634 |
-2.453 |
0.000 |
2.533 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.999 |
-2.662 |
0.000 |
| y |
-2.662 |
-26.508 |
0.000 |
| z |
0.000 |
0.000 |
-23.708 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.109 |
-2.662 |
0.000 |
| y |
-2.662 |
-7.154 |
0.000 |
| z |
0.000 |
0.000 |
-2.954 |
|
| Polar |
|---|
| 3z2-r2 | -5.909 |
|---|
| x2-y2 | 11.509 |
|---|
| xy | -2.662 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.521 |
-0.262 |
0.000 |
| y |
-0.262 |
3.622 |
0.000 |
| z |
0.000 |
0.000 |
1.849 |
<r2> (average value of r
2) Å
2
| <r2> |
62.831 |
| (<r2>)1/2 |
7.927 |
Jump to
S1C1
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -243.884479 |
| Energy at 298.15K | -243.889343 |
| HF Energy | -243.884479 |
| Nuclear repulsion energy | 125.540990 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4158 |
3760 |
105.93 |
|
|
|
| 2 |
A |
4005 |
3622 |
64.71 |
|
|
|
| 3 |
A |
3873 |
3502 |
56.16 |
|
|
|
| 4 |
A |
2004 |
1813 |
727.57 |
|
|
|
| 5 |
A |
1804 |
1631 |
141.55 |
|
|
|
| 6 |
A |
1571 |
1420 |
185.89 |
|
|
|
| 7 |
A |
1378 |
1246 |
238.70 |
|
|
|
| 8 |
A |
1200 |
1086 |
66.16 |
|
|
|
| 9 |
A |
1059 |
957 |
21.47 |
|
|
|
| 10 |
A |
889 |
804 |
85.27 |
|
|
|
| 11 |
A |
651 |
589 |
55.36 |
|
|
|
| 12 |
A |
633 |
572 |
92.54 |
|
|
|
| 13 |
A |
551 |
499 |
55.34 |
|
|
|
| 14 |
A |
528 |
477 |
19.08 |
|
|
|
| 15 |
A |
213 |
192 |
316.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12257.1 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 11085.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
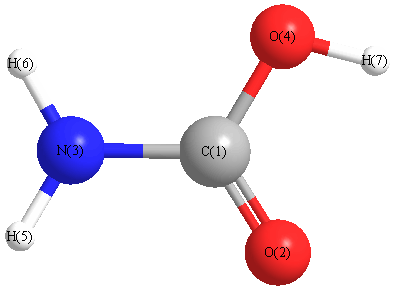
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.041 |
0.121 |
-0.002 |
| O2 |
-0.456 |
1.233 |
0.004 |
| N3 |
1.258 |
-0.233 |
-0.032 |
| O4 |
-0.830 |
-0.951 |
0.002 |
| H5 |
1.934 |
0.480 |
0.086 |
| H6 |
1.522 |
-1.177 |
0.100 |
| H7 |
-1.725 |
-0.656 |
0.004 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.1874 | 1.3464 | 1.3313 | 2.0089 | 2.0341 | 1.8552 |
O2 | 1.1874 | | 2.2562 | 2.2156 | 2.5076 | 3.1198 | 2.2756 | N3 | 1.3464 | 2.2562 | | 2.2088 | 0.9894 | 0.9894 | 3.0135 | O4 | 1.3313 | 2.2156 | 2.2088 | | 3.1138 | 2.3655 | 0.9423 | H5 | 2.0089 | 2.5076 | 0.9894 | 3.1138 | | 1.7075 | 3.8324 | H6 | 2.0341 | 3.1198 | 0.9894 | 2.3655 | 1.7075 | | 3.2905 | H7 | 1.8552 | 2.2756 | 3.0135 | 0.9423 | 3.8324 | 3.2905 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.826 |
|
C1 |
N3 |
H6 |
120.345 |
| C1 |
O4 |
H7 |
108.152 |
|
O2 |
C1 |
N3 |
125.744 |
| O2 |
C1 |
O4 |
123.104 |
|
N3 |
C1 |
O4 |
111.146 |
| H5 |
N3 |
H6 |
119.285 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.778 |
|
|
|
| 2 |
O |
-0.502 |
|
|
|
| 3 |
N |
-0.873 |
|
|
|
| 4 |
O |
-0.632 |
|
|
|
| 5 |
H |
0.390 |
|
|
|
| 6 |
H |
0.388 |
|
|
|
| 7 |
H |
0.450 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.303 |
-2.101 |
0.451 |
2.513 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.715 |
1.349 |
0.883 |
| y |
1.349 |
-26.979 |
-0.229 |
| z |
0.883 |
-0.229 |
-23.669 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.609 |
1.349 |
0.883 |
| y |
1.349 |
-7.787 |
-0.229 |
| z |
0.883 |
-0.229 |
-2.822 |
|
| Polar |
|---|
| 3z2-r2 | -5.644 |
|---|
| x2-y2 | 12.264 |
|---|
| xy | 1.349 |
|---|
| xz | 0.883 |
|---|
| yz | -0.229 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.686 |
-0.235 |
0.008 |
| y |
-0.235 |
3.473 |
0.001 |
| z |
0.008 |
0.001 |
1.859 |
<r2> (average value of r
2) Å
2
| <r2> |
62.851 |
| (<r2>)1/2 |
7.928 |
