Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -245.109030 |
| Energy at 298.15K | |
| HF Energy | -245.109030 |
| Nuclear repulsion energy | 123.781716 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3810 |
3668 |
59.68 |
|
|
|
| 2 |
A' |
3790 |
3649 |
53.47 |
|
|
|
| 3 |
A' |
3656 |
3520 |
47.26 |
|
|
|
| 4 |
A' |
1870 |
1800 |
536.16 |
|
|
|
| 5 |
A' |
1651 |
1590 |
133.07 |
|
|
|
| 6 |
A' |
1451 |
1397 |
131.77 |
|
|
|
| 7 |
A' |
1254 |
1207 |
226.98 |
|
|
|
| 8 |
A' |
1091 |
1050 |
41.63 |
|
|
|
| 9 |
A' |
967 |
931 |
40.23 |
|
|
|
| 10 |
A' |
591 |
569 |
37.10 |
|
|
|
| 11 |
A' |
489 |
471 |
7.29 |
|
|
|
| 12 |
A" |
794 |
764 |
37.43 |
|
|
|
| 13 |
A" |
608 |
585 |
73.54 |
|
|
|
| 14 |
A" |
494 |
475 |
74.22 |
|
|
|
| 15 |
A" |
96i |
92i |
272.84 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11209.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 10791.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.131 |
0.000 |
| O2 |
-0.074 |
1.336 |
0.000 |
| N3 |
1.141 |
-0.597 |
0.000 |
| O4 |
-1.085 |
-0.685 |
0.000 |
| H5 |
2.018 |
-0.112 |
0.000 |
| H6 |
1.122 |
-1.601 |
0.000 |
| H7 |
-1.852 |
-0.101 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2078 | 1.3534 | 1.3578 | 2.0329 | 2.0633 | 1.8668 |
O2 | 1.2078 | | 2.2836 | 2.2605 | 2.5448 | 3.1713 | 2.2865 | N3 | 1.3534 | 2.2836 | | 2.2277 | 1.0027 | 1.0034 | 3.0340 | O4 | 1.3578 | 2.2605 | 2.2277 | | 3.1559 | 2.3898 | 0.9644 | H5 | 2.0329 | 2.5448 | 1.0027 | 3.1559 | | 1.7372 | 3.8706 | H6 | 2.0633 | 3.1713 | 1.0034 | 2.3898 | 1.7372 | | 3.3314 | H7 | 1.8668 | 2.2865 | 3.0340 | 0.9644 | 3.8706 | 3.3314 | |
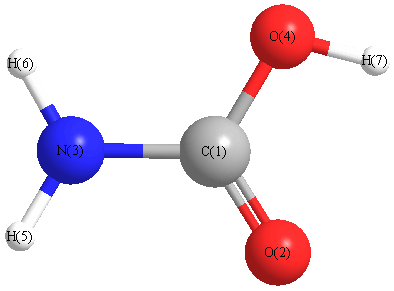 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.514 |
|
C1 |
N3 |
H6 |
121.500 |
| C1 |
O4 |
H7 |
105.752 |
|
O2 |
C1 |
N3 |
126.061 |
| O2 |
C1 |
O4 |
123.435 |
|
N3 |
C1 |
O4 |
110.503 |
| H5 |
N3 |
H6 |
119.986 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.580 |
|
|
|
| 2 |
O |
-0.397 |
|
|
|
| 3 |
N |
-0.798 |
|
|
|
| 4 |
O |
-0.546 |
|
|
|
| 5 |
H |
0.375 |
|
|
|
| 6 |
H |
0.373 |
|
|
|
| 7 |
H |
0.413 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.827 |
-2.256 |
0.000 |
2.403 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.852 |
-2.481 |
0.000 |
| y |
-2.481 |
-26.002 |
0.000 |
| z |
0.000 |
0.000 |
-23.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.042 |
-2.481 |
0.000 |
| y |
-2.481 |
-6.683 |
0.000 |
| z |
0.000 |
0.000 |
-3.359 |
|
| Polar |
|---|
| 3z2-r2 | -6.718 |
|---|
| x2-y2 | 11.149 |
|---|
| xy | -2.481 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.175 |
-0.317 |
0.000 |
| y |
-0.317 |
4.045 |
0.000 |
| z |
0.000 |
0.000 |
1.960 |
<r2> (average value of r
2) Å
2
| <r2> |
64.026 |
| (<r2>)1/2 |
8.002 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -245.109032 |
| Energy at 298.15K | -245.113690 |
| HF Energy | -245.109032 |
| Nuclear repulsion energy | 123.768126 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3811 |
3669 |
59.54 |
|
|
|
| 2 |
A |
3780 |
3639 |
49.63 |
|
|
|
| 3 |
A |
3649 |
3513 |
43.71 |
|
|
|
| 4 |
A |
1870 |
1800 |
528.34 |
|
|
|
| 5 |
A |
1654 |
1593 |
128.22 |
|
|
|
| 6 |
A |
1451 |
1397 |
132.90 |
|
|
|
| 7 |
A |
1255 |
1209 |
216.08 |
|
|
|
| 8 |
A |
1095 |
1054 |
52.41 |
|
|
|
| 9 |
A |
969 |
933 |
39.20 |
|
|
|
| 10 |
A |
793 |
764 |
41.56 |
|
|
|
| 11 |
A |
605 |
583 |
83.31 |
|
|
|
| 12 |
A |
591 |
569 |
39.04 |
|
|
|
| 13 |
A |
500 |
482 |
40.84 |
|
|
|
| 14 |
A |
480 |
462 |
28.98 |
|
|
|
| 15 |
A |
203 |
196 |
279.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11353.5 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 10930.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.040 |
0.125 |
-0.001 |
| O2 |
-0.472 |
1.252 |
0.004 |
| N3 |
1.269 |
-0.227 |
-0.032 |
| O4 |
-0.829 |
-0.979 |
0.002 |
| H5 |
1.951 |
0.499 |
0.085 |
| H6 |
1.545 |
-1.184 |
0.097 |
| H7 |
-1.737 |
-0.654 |
0.003 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2076 | 1.3557 | 1.3572 | 2.0273 | 2.0579 | 1.8674 |
O2 | 1.2076 | | 2.2850 | 2.2601 | 2.5382 | 3.1644 | 2.2881 | N3 | 1.3557 | 2.2850 | | 2.2291 | 1.0033 | 1.0040 | 3.0364 | O4 | 1.3572 | 2.2601 | 2.2291 | | 3.1498 | 2.3852 | 0.9644 | H5 | 2.0273 | 2.5382 | 1.0033 | 3.1498 | | 1.7315 | 3.8648 | H6 | 2.0579 | 3.1644 | 1.0040 | 2.3852 | 1.7315 | | 3.3262 | H7 | 1.8674 | 2.2881 | 3.0364 | 0.9644 | 3.8648 | 3.3262 | |
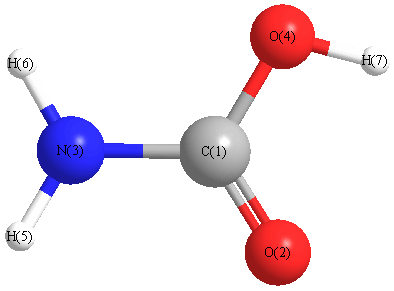 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.721 |
|
C1 |
N3 |
H6 |
120.678 |
| C1 |
O4 |
H7 |
105.853 |
|
O2 |
C1 |
N3 |
126.018 |
| O2 |
C1 |
O4 |
123.471 |
|
N3 |
C1 |
O4 |
110.503 |
| H5 |
N3 |
H6 |
119.212 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.573 |
|
|
|
| 2 |
O |
-0.395 |
|
|
|
| 3 |
N |
-0.790 |
|
|
|
| 4 |
O |
-0.543 |
|
|
|
| 5 |
H |
0.372 |
|
|
|
| 6 |
H |
0.370 |
|
|
|
| 7 |
H |
0.413 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.399 |
-1.885 |
0.415 |
2.384 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.584 |
1.202 |
0.824 |
| y |
1.202 |
-26.436 |
-0.201 |
| z |
0.824 |
-0.201 |
-23.752 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.510 |
1.202 |
0.824 |
| y |
1.202 |
-7.268 |
-0.201 |
| z |
0.824 |
-0.201 |
-3.242 |
|
| Polar |
|---|
| 3z2-r2 | -6.484 |
|---|
| x2-y2 | 11.852 |
|---|
| xy | 1.202 |
|---|
| xz | 0.824 |
|---|
| yz | -0.201 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.344 |
-0.217 |
0.004 |
| y |
-0.217 |
3.890 |
0.002 |
| z |
0.004 |
0.002 |
1.973 |
<r2> (average value of r
2) Å
2
| <r2> |
64.037 |
| (<r2>)1/2 |
8.002 |
