Jump to
S1C2
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -244.958909 |
| Energy at 298.15K | |
| HF Energy | -244.958909 |
| Nuclear repulsion energy | 124.033309 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3835 |
3680 |
63.21 |
|
|
|
| 2 |
A' |
3811 |
3657 |
55.88 |
|
|
|
| 3 |
A' |
3675 |
3526 |
49.12 |
|
|
|
| 4 |
A' |
1886 |
1810 |
545.72 |
|
|
|
| 5 |
A' |
1658 |
1591 |
142.56 |
|
|
|
| 6 |
A' |
1464 |
1405 |
132.60 |
|
|
|
| 7 |
A' |
1263 |
1212 |
227.77 |
|
|
|
| 8 |
A' |
1098 |
1054 |
38.24 |
|
|
|
| 9 |
A' |
980 |
940 |
37.56 |
|
|
|
| 10 |
A' |
595 |
571 |
38.66 |
|
|
|
| 11 |
A' |
491 |
472 |
7.02 |
|
|
|
| 12 |
A" |
800 |
768 |
38.68 |
|
|
|
| 13 |
A" |
612 |
587 |
73.94 |
|
|
|
| 14 |
A" |
498 |
477 |
73.41 |
|
|
|
| 15 |
A" |
107i |
103i |
273.57 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11279.4 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 10823.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.130 |
0.000 |
| O2 |
-0.070 |
1.334 |
0.000 |
| N3 |
1.136 |
-0.600 |
0.000 |
| O4 |
-1.085 |
-0.680 |
0.000 |
| H5 |
2.014 |
-0.118 |
0.000 |
| H6 |
1.114 |
-1.602 |
0.000 |
| H7 |
-1.848 |
-0.094 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2060 | 1.3509 | 1.3537 | 2.0294 | 2.0596 | 1.8619 |
O2 | 1.2060 | | 2.2797 | 2.2553 | 2.5399 | 3.1661 | 2.2810 | N3 | 1.3509 | 2.2797 | | 2.2224 | 1.0016 | 1.0024 | 3.0275 | O4 | 1.3537 | 2.2553 | 2.2224 | | 3.1493 | 2.3840 | 0.9628 | H5 | 2.0294 | 2.5399 | 1.0016 | 3.1493 | | 1.7363 | 3.8627 | H6 | 2.0596 | 3.1661 | 1.0024 | 2.3840 | 1.7363 | | 3.3242 | H7 | 1.8619 | 2.2810 | 3.0275 | 0.9628 | 3.8627 | 3.3242 | |
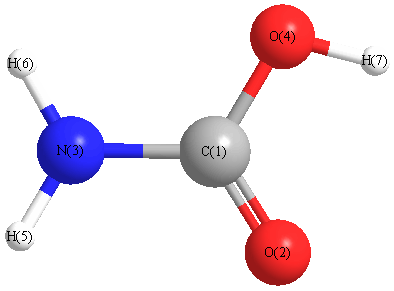 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.477 |
|
C1 |
N3 |
H6 |
121.437 |
| C1 |
O4 |
H7 |
105.743 |
|
O2 |
C1 |
N3 |
126.047 |
| O2 |
C1 |
O4 |
123.442 |
|
N3 |
C1 |
O4 |
110.511 |
| H5 |
N3 |
H6 |
120.086 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.577 |
|
|
|
| 2 |
O |
-0.395 |
|
|
|
| 3 |
N |
-0.802 |
|
|
|
| 4 |
O |
-0.546 |
|
|
|
| 5 |
H |
0.377 |
|
|
|
| 6 |
H |
0.375 |
|
|
|
| 7 |
H |
0.415 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.809 |
-2.266 |
0.000 |
2.407 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.910 |
-2.500 |
0.000 |
| y |
-2.500 |
-26.001 |
0.000 |
| z |
0.000 |
0.000 |
-23.814 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
9.998 |
-2.500 |
0.000 |
| y |
-2.500 |
-6.639 |
0.000 |
| z |
0.000 |
0.000 |
-3.359 |
|
| Polar |
|---|
| 3z2-r2 | -6.717 |
|---|
| x2-y2 | 11.091 |
|---|
| xy | -2.500 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.142 |
-0.314 |
0.000 |
| y |
-0.314 |
4.018 |
0.000 |
| z |
0.000 |
0.000 |
1.956 |
<r2> (average value of r
2) Å
2
| <r2> |
63.831 |
| (<r2>)1/2 |
7.989 |
Jump to
S1C1
Energy calculated at HSEh1PBE/6-311G*
| | hartrees |
|---|
| Energy at 0K | -244.958914 |
| Energy at 298.15K | -244.963585 |
| HF Energy | -244.958914 |
| Nuclear repulsion energy | 124.018792 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3836 |
3681 |
63.14 |
|
|
|
| 2 |
A |
3801 |
3647 |
51.77 |
|
|
|
| 3 |
A |
3668 |
3520 |
45.50 |
|
|
|
| 4 |
A |
1886 |
1810 |
537.73 |
|
|
|
| 5 |
A |
1661 |
1594 |
136.90 |
|
|
|
| 6 |
A |
1463 |
1404 |
133.99 |
|
|
|
| 7 |
A |
1265 |
1214 |
217.30 |
|
|
|
| 8 |
A |
1103 |
1058 |
48.64 |
|
|
|
| 9 |
A |
981 |
941 |
36.75 |
|
|
|
| 10 |
A |
800 |
768 |
43.10 |
|
|
|
| 11 |
A |
609 |
584 |
83.27 |
|
|
|
| 12 |
A |
595 |
571 |
40.56 |
|
|
|
| 13 |
A |
504 |
483 |
41.81 |
|
|
|
| 14 |
A |
483 |
464 |
27.16 |
|
|
|
| 15 |
A |
201 |
193 |
280.46 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11427.1 cm
-1
Scaled (by 0.9596) Zero Point Vibrational Energy (zpe) 10965.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.040 |
0.124 |
-0.001 |
| O2 |
-0.476 |
1.248 |
0.004 |
| N3 |
1.268 |
-0.223 |
-0.032 |
| O4 |
-0.823 |
-0.980 |
0.002 |
| H5 |
1.945 |
0.507 |
0.086 |
| H6 |
1.546 |
-1.177 |
0.098 |
| H7 |
-1.730 |
-0.658 |
0.003 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2058 | 1.3532 | 1.3532 | 2.0238 | 2.0540 | 1.8623 |
O2 | 1.2058 | | 2.2812 | 2.2549 | 2.5335 | 3.1591 | 2.2818 | N3 | 1.3532 | 2.2812 | | 2.2239 | 1.0023 | 1.0030 | 3.0297 | O4 | 1.3532 | 2.2549 | 2.2239 | | 3.1432 | 2.3793 | 0.9628 | H5 | 2.0238 | 2.5335 | 1.0023 | 3.1432 | | 1.7304 | 3.8565 | H6 | 2.0540 | 3.1591 | 1.0030 | 2.3793 | 1.7304 | | 3.3188 | H7 | 1.8623 | 2.2818 | 3.0297 | 0.9628 | 3.8565 | 3.3188 | |
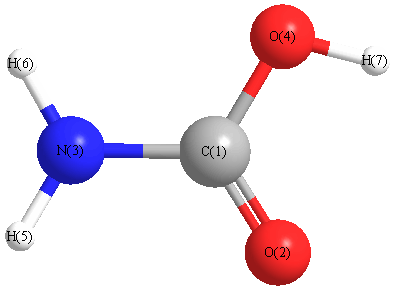 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.678 |
|
C1 |
N3 |
H6 |
120.602 |
| C1 |
O4 |
H7 |
105.808 |
|
O2 |
C1 |
N3 |
126.016 |
| O2 |
C1 |
O4 |
123.466 |
|
N3 |
C1 |
O4 |
110.511 |
| H5 |
N3 |
H6 |
119.294 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.569 |
|
|
|
| 2 |
O |
-0.393 |
|
|
|
| 3 |
N |
-0.794 |
|
|
|
| 4 |
O |
-0.543 |
|
|
|
| 5 |
H |
0.374 |
|
|
|
| 6 |
H |
0.372 |
|
|
|
| 7 |
H |
0.415 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.399 |
-1.889 |
0.418 |
2.388 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.646 |
1.249 |
0.829 |
| y |
1.249 |
-26.437 |
-0.200 |
| z |
0.829 |
-0.200 |
-23.779 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
10.462 |
1.249 |
0.829 |
| y |
1.249 |
-7.224 |
-0.200 |
| z |
0.829 |
-0.200 |
-3.238 |
|
| Polar |
|---|
| 3z2-r2 | -6.476 |
|---|
| x2-y2 | 11.791 |
|---|
| xy | 1.249 |
|---|
| xz | 0.829 |
|---|
| yz | -0.200 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.313 |
-0.214 |
0.004 |
| y |
-0.214 |
3.862 |
0.002 |
| z |
0.004 |
0.002 |
1.968 |
<r2> (average value of r
2) Å
2
| <r2> |
63.843 |
| (<r2>)1/2 |
7.990 |
