Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -245.009645 |
| Energy at 298.15K | |
| HF Energy | -244.760776 |
| Nuclear repulsion energy | 123.581560 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3815 |
3815 |
65.38 |
|
|
|
| 2 |
A' |
3808 |
3808 |
52.93 |
|
|
|
| 3 |
A' |
3672 |
3672 |
52.20 |
|
|
|
| 4 |
A' |
1860 |
1860 |
514.68 |
|
|
|
| 5 |
A' |
1670 |
1670 |
117.97 |
|
|
|
| 6 |
A' |
1454 |
1454 |
128.20 |
|
|
|
| 7 |
A' |
1262 |
1262 |
231.06 |
|
|
|
| 8 |
A' |
1098 |
1098 |
48.51 |
|
|
|
| 9 |
A' |
963 |
963 |
41.94 |
|
|
|
| 10 |
A' |
591 |
591 |
36.18 |
|
|
|
| 11 |
A' |
493 |
493 |
6.51 |
|
|
|
| 12 |
A" |
790 |
790 |
34.44 |
|
|
|
| 13 |
A" |
609 |
609 |
73.11 |
|
|
|
| 14 |
A" |
492 |
492 |
76.37 |
|
|
|
| 15 |
A" |
236i |
236i |
276.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11169.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11169.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.132 |
0.000 |
| O2 |
-0.075 |
1.340 |
0.000 |
| N3 |
1.143 |
-0.597 |
0.000 |
| O4 |
-1.085 |
-0.689 |
0.000 |
| H5 |
2.019 |
-0.113 |
0.000 |
| H6 |
1.121 |
-1.599 |
0.000 |
| H7 |
-1.855 |
-0.109 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2101 | 1.3559 | 1.3611 | 2.0340 | 2.0630 | 1.8702 |
O2 | 1.2101 | | 2.2887 | 2.2667 | 2.5491 | 3.1737 | 2.2943 | N3 | 1.3559 | 2.2887 | | 2.2299 | 1.0015 | 1.0020 | 3.0369 | O4 | 1.3611 | 2.2667 | 2.2299 | | 3.1575 | 2.3869 | 0.9639 | H5 | 2.0340 | 2.5491 | 1.0015 | 3.1575 | | 1.7366 | 3.8738 | H6 | 2.0630 | 3.1737 | 1.0020 | 2.3869 | 1.7366 | | 3.3285 | H7 | 1.8702 | 2.2943 | 3.0369 | 0.9639 | 3.8738 | 3.3285 | |
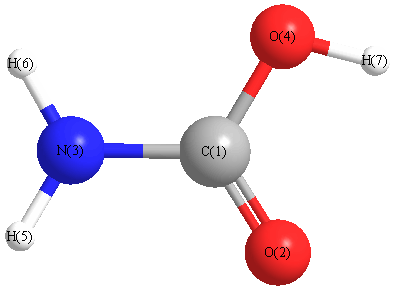 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
118.494 |
|
C1 |
N3 |
H6 |
121.342 |
| C1 |
O4 |
H7 |
105.831 |
|
O2 |
C1 |
N3 |
126.136 |
| O2 |
C1 |
O4 |
123.553 |
|
N3 |
C1 |
O4 |
110.311 |
| H5 |
N3 |
H6 |
120.164 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -245.009774 |
| Energy at 298.15K | -245.014596 |
| HF Energy | -244.760727 |
| Nuclear repulsion energy | 123.537985 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3815 |
3815 |
62.89 |
|
|
|
| 2 |
A |
3783 |
3783 |
45.76 |
|
|
|
| 3 |
A |
3654 |
3654 |
42.91 |
|
|
|
| 4 |
A |
1861 |
1861 |
494.78 |
|
|
|
| 5 |
A |
1677 |
1677 |
108.50 |
|
|
|
| 6 |
A |
1452 |
1452 |
127.57 |
|
|
|
| 7 |
A |
1266 |
1266 |
200.04 |
|
|
|
| 8 |
A |
1110 |
1110 |
78.64 |
|
|
|
| 9 |
A |
964 |
964 |
40.65 |
|
|
|
| 10 |
A |
790 |
790 |
46.42 |
|
|
|
| 11 |
A |
602 |
602 |
87.13 |
|
|
|
| 12 |
A |
590 |
590 |
44.13 |
|
|
|
| 13 |
A |
506 |
506 |
29.37 |
|
|
|
| 14 |
A |
473 |
473 |
29.98 |
|
|
|
| 15 |
A |
331 |
331 |
293.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11437.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11437.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.040 |
0.126 |
-0.002 |
| O2 |
-0.473 |
1.255 |
0.006 |
| N3 |
1.273 |
-0.229 |
-0.050 |
| O4 |
-0.829 |
-0.982 |
0.003 |
| H5 |
1.942 |
0.496 |
0.132 |
| H6 |
1.534 |
-1.177 |
0.152 |
| H7 |
-1.737 |
-0.662 |
0.004 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.2092 | 1.3613 | 1.3608 | 2.0209 | 2.0489 | 1.8715 |
O2 | 1.2092 | | 2.2926 | 2.2658 | 2.5351 | 3.1565 | 2.2962 | N3 | 1.3613 | 2.2926 | | 2.2337 | 1.0031 | 1.0036 | 3.0422 | O4 | 1.3608 | 2.2658 | 2.2337 | | 3.1435 | 2.3748 | 0.9638 | H5 | 2.0209 | 2.5351 | 1.0031 | 3.1435 | | 1.7225 | 3.8597 | H6 | 2.0489 | 3.1565 | 1.0036 | 2.3748 | 1.7225 | | 3.3146 | H7 | 1.8715 | 2.2962 | 3.0422 | 0.9638 | 3.8597 | 3.3146 | |
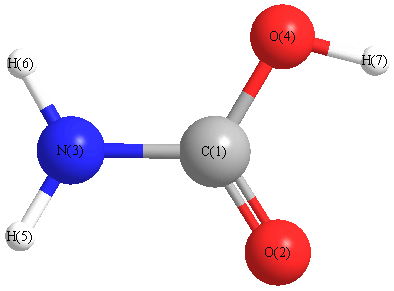 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
116.645 |
|
C1 |
N3 |
H6 |
119.304 |
| C1 |
O4 |
H7 |
105.973 |
|
O2 |
C1 |
N3 |
126.122 |
| O2 |
C1 |
O4 |
123.576 |
|
N3 |
C1 |
O4 |
110.283 |
| H5 |
N3 |
H6 |
118.267 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
