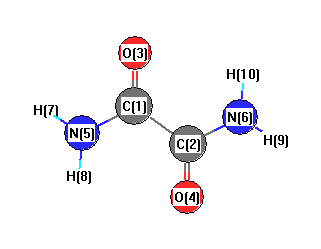
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3990 |
3609 |
0.00 |
|
|
|
| 2 |
Ag |
3854 |
3486 |
0.00 |
|
|
|
| 3 |
Ag |
1990 |
1800 |
0.00 |
|
|
|
| 4 |
Ag |
1801 |
1629 |
0.00 |
|
|
|
| 5 |
Ag |
1565 |
1416 |
0.00 |
|
|
|
| 6 |
Ag |
1224 |
1107 |
0.00 |
|
|
|
| 7 |
Ag |
842 |
762 |
0.00 |
|
|
|
| 8 |
Ag |
582 |
526 |
0.00 |
|
|
|
| 9 |
Ag |
441 |
399 |
0.00 |
|
|
|
| 10 |
Au |
717 |
648 |
16.82 |
|
|
|
| 11 |
Au |
530 |
479 |
251.71 |
|
|
|
| 12 |
Au |
378 |
342 |
415.49 |
|
|
|
| 13 |
Au |
83 |
75 |
3.56 |
|
|
|
| 14 |
Bg |
933 |
844 |
0.00 |
|
|
|
| 15 |
Bg |
686 |
620 |
0.00 |
|
|
|
| 16 |
Bg |
382 |
345 |
0.00 |
|
|
|
| 17 |
Bu |
3991 |
3609 |
169.84 |
|
|
|
| 18 |
Bu |
3854 |
3486 |
150.36 |
|
|
|
| 19 |
Bu |
1944 |
1758 |
977.55 |
|
|
|
| 20 |
Bu |
1789 |
1618 |
320.28 |
|
|
|
| 21 |
Bu |
1418 |
1282 |
284.59 |
|
|
|
| 22 |
Bu |
1205 |
1090 |
0.92 |
|
|
|
| 23 |
Bu |
636 |
575 |
42.14 |
|
|
|
| 24 |
Bu |
306 |
277 |
37.33 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17570.3 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 15890.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.579 |
0.766 |
|
0.714 |
| 2 |
C |
0.579 |
0.766 |
|
0.759 |
| 3 |
O |
-0.487 |
-0.622 |
|
-0.596 |
| 4 |
O |
-0.487 |
-0.622 |
|
-0.608 |
| 5 |
N |
-0.903 |
-1.046 |
|
-1.046 |
| 6 |
N |
-0.903 |
-1.046 |
|
-1.059 |
| 7 |
H |
0.391 |
0.474 |
|
0.480 |
| 8 |
H |
0.421 |
0.429 |
|
0.436 |
| 9 |
H |
0.391 |
0.474 |
|
0.480 |
| 10 |
H |
0.421 |
0.429 |
|
0.439 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.012 |
-0.016 |
0.000 |
0.020 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.737 |
-12.132 |
0.000 |
| y |
-12.132 |
-33.188 |
0.000 |
| z |
0.000 |
0.000 |
-35.199 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.457 |
-12.132 |
0.000 |
| y |
-12.132 |
-0.220 |
0.000 |
| z |
0.000 |
0.000 |
-3.237 |
|
| Polar |
|---|
| 3z2-r2 | -6.474 |
|---|
| x2-y2 | 2.451 |
|---|
| xy | -12.132 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.839 |
-0.088 |
0.000 |
| y |
-0.088 |
5.457 |
0.000 |
| z |
0.000 |
0.000 |
2.914 |
<r2> (average value of r
2) Å
2
| <r2> |
142.559 |
| (<r2>)1/2 |
11.940 |