return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H10S (Thiophene, tetrahydro-3-methyl-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-593.774219
Energy at 298.15K-593.786245
HF Energy-592.733038
Nuclear repulsion energy314.493024
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3175 3007 12.75      
2 A 3171 3003 25.30      
3 A 3164 2997 21.99      
4 A 3162 2994 13.14      
5 A 3142 2976 16.88      
6 A 3110 2946 18.94      
7 A 3103 2939 47.41      
8 A 3090 2927 26.90      
9 A 3081 2918 7.59      
10 A 3077 2914 20.11      
11 A 1550 1468 8.34      
12 A 1545 1463 1.15      
13 A 1530 1449 8.61      
14 A 1524 1444 4.41      
15 A 1517 1436 6.52      
16 A 1449 1373 13.67      
17 A 1400 1326 4.51      
18 A 1368 1296 2.42      
19 A 1352 1281 1.93      
20 A 1332 1262 11.88      
21 A 1302 1233 15.27      
22 A 1264 1197 3.32      
23 A 1211 1147 2.69      
24 A 1158 1096 0.76      
25 A 1111 1053 2.42      
26 A 1105 1046 0.66      
27 A 1068 1011 0.39      
28 A 1026 972 2.53      
29 A 994 941 0.19      
30 A 961 910 0.37      
31 A 933 884 5.18      
32 A 885 839 1.08      
33 A 831 787 1.48      
34 A 748 708 4.40      
35 A 722 683 1.64      
36 A 629 596 1.37      
37 A 504 477 0.16      
38 A 392 371 0.09      
39 A 360 341 0.12      
40 A 251 238 0.09      
41 A 194 184 0.17      
42 A 117 111 2.05      

Unscaled Zero Point Vibrational Energy (zpe) 31802.7 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 30120.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.14262 0.08783 0.07214

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 -1.456 -0.436 -0.261
C2 1.852 -0.315 -0.828
H3 1.157 -0.284 -1.671
H4 2.594 0.477 -0.964
H5 2.375 -1.274 -0.868
C6 -0.031 -1.156 0.634
H7 0.216 -2.124 0.192
H8 -0.297 -1.305 1.683
C9 1.120 -0.156 0.501
H10 1.828 -0.313 1.324
C11 -0.727 1.239 -0.320
H12 -1.480 1.970 -0.017
H13 -0.411 1.470 -1.339
C14 0.461 1.217 0.639
H15 0.104 1.348 1.666
H16 1.164 2.028 0.420

Atom - Atom Distances (Å)
  S1 C2 H3 H4 H5 C6 H7 H8 C9 H10 C11 H12 H13 C14 H15 H16
S13.35832.97304.21023.96811.83032.41932.42462.70043.64831.82772.41782.42652.68653.05463.6600
C23.35831.09251.09351.09322.52812.64433.45041.52592.15243.05394.12012.92702.53683.47012.7422
H32.97301.09251.77251.76262.73582.78283.79552.17593.06912.77443.84282.37622.84163.86113.1170
H44.21021.09351.77251.76753.48013.70934.30582.17272.53923.46754.44003.18632.76883.72482.5228
H53.96811.09321.76261.76752.83882.55103.69432.16782.45504.03005.10903.93873.48444.29573.7455
C61.83032.52812.73583.48012.83881.09261.09251.53032.15452.67083.50593.30692.42382.71193.4077
H72.41932.64432.78283.70932.55101.09261.77682.18762.67553.53114.43623.95743.38033.77394.2653
H82.42463.45043.79554.30583.69431.09251.77682.17352.37223.26683.87444.10492.83302.68323.8522
C92.70041.52592.17592.17272.16781.53032.18762.17351.09712.45623.39742.89381.52942.15662.1862
H103.64832.15243.06912.53922.45502.15452.67552.37221.09713.41214.23653.90972.16332.41822.5964
C111.82773.05392.77443.46754.03002.67083.53113.26682.45623.41211.09131.09191.52732.15612.1782
H122.41784.12013.84284.44005.10903.50594.43623.87443.39744.23651.09131.77212.18212.39322.6795
H132.42652.92702.37623.18633.93873.30693.95744.10492.89383.90971.09191.77212.17673.05182.4256
C142.68652.53682.84162.76883.48442.42383.38032.83301.52942.16331.52732.18212.17671.09511.0952
H153.05463.47013.86113.72484.29572.71193.77392.68322.15662.41822.15612.39323.05181.09511.7717
H163.66002.74223.11702.52283.74553.40774.26533.85222.18622.59642.17822.67952.42561.09521.7717

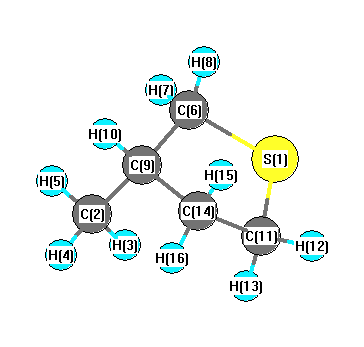
picture of Thiophene, tetrahydro-3-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C6 H7 109.106 S1 C6 H8 109.497
S1 C6 C9 106.603 S1 C11 H12 109.241
S1 C11 H13 109.845 S1 C11 C14 106.054
C2 C9 C6 111.632 C2 C9 H10 109.196
C2 C9 C14 112.256 H3 C2 H4 108.357
H3 C2 H5 107.502 H3 C2 C9 111.332
H4 C2 H5 107.855 H4 C2 C9 111.009
H5 C2 C9 110.642 C6 S1 C11 93.794
C6 C9 H10 109.066 C6 C9 C14 104.777
H7 C6 H8 108.807 H7 C6 C9 111.953
H8 C6 C9 110.825 C9 C14 C11 106.942
C9 C14 H15 109.404 C9 C14 H16 111.739
H10 C9 C14 109.808 C11 C14 H15 109.508
C11 C14 H16 111.253 H12 C11 H13 108.526
H12 C11 C14 111.805 H13 C11 C14 111.332
H15 C14 H16 107.976
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability