return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H12 (Ethylcyclobutane)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-235.151626
Energy at 298.15K-235.165285
HF Energy-234.205653
Nuclear repulsion energy249.378197
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3177 3009 62.57      
2 A 3168 3000 43.86      
3 A 3163 2996 30.42      
4 A 3156 2989 4.38      
5 A 3150 2983 16.20      
6 A 3115 2950 48.45      
7 A 3110 2945 12.72      
8 A 3096 2932 29.31      
9 A 3090 2927 64.04      
10 A 3081 2918 25.27      
11 A 3077 2914 18.72      
12 A 3064 2902 21.55      
13 A 1554 1472 4.38      
14 A 1544 1462 8.17      
15 A 1540 1459 9.02      
16 A 1527 1447 3.96      
17 A 1525 1444 0.87      
18 A 1514 1433 4.74      
19 A 1446 1370 8.20      
20 A 1420 1345 0.17      
21 A 1367 1295 2.82      
22 A 1328 1258 0.05      
23 A 1302 1233 4.65      
24 A 1298 1230 2.27      
25 A 1279 1211 0.18      
26 A 1270 1202 0.11      
27 A 1268 1201 0.03      
28 A 1212 1148 0.11      
29 A 1204 1141 1.01      
30 A 1152 1091 1.12      
31 A 1112 1053 0.15      
32 A 1055 1000 0.06      
33 A 1026 971 0.34      
34 A 1018 964 3.89      
35 A 980 928 0.53      
36 A 956 906 1.81      
37 A 947 897 1.86      
38 A 890 843 0.88      
39 A 841 797 0.47      
40 A 789 747 0.35      
41 A 771 730 2.09      
42 A 674 638 3.88      
43 A 428 405 0.16      
44 A 371 351 0.15      
45 A 252 239 0.07      
46 A 220 209 0.04      
47 A 174 165 0.03      
48 A 120 114 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 37408.1 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 35429.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.23792 0.07875 0.07022

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.390 0.259 -0.053
H2 2.737 0.093 0.971
H3 3.208 0.001 -0.730
H4 2.189 1.328 -0.167
C5 1.142 -0.571 -0.342
H6 1.376 -1.638 -0.238
H7 0.830 -0.423 -1.383
C8 -1.764 0.326 -0.569
H9 -2.807 0.641 -0.491
H10 -1.495 0.273 -1.626
C11 -0.715 1.112 0.254
H12 -1.161 1.562 1.145
H13 -0.125 1.874 -0.264
C14 -0.018 -0.235 0.586
H15 0.300 -0.360 1.628
C16 -1.339 -0.949 0.198
H17 -1.252 -1.882 -0.367
H18 -1.975 -1.122 1.071

Atom - Atom Distances (Å)
  C1 H2 H3 H4 C5 H6 H7 C8 H9 H10 C11 H12 H13 C14 H15 C16 H17 H18
C11.09351.09301.09271.52622.15872.16094.18575.22914.19073.23413.96742.99572.53952.75233.92744.23624.7136
H21.09351.76671.76592.16942.51143.07314.76235.75984.96803.67004.16973.59002.80112.56454.27724.64774.8669
H31.09301.76671.76462.17822.50612.50254.98466.05354.79474.19415.00413.85133.49173.76104.73664.85465.6001
H41.09271.76591.76462.17533.07552.52864.09765.05394.10032.94263.60592.38002.80743.10474.21464.71014.9872
C51.52622.16942.17822.17531.09711.09723.04964.13353.05182.57603.47372.75471.52272.15252.56702.72973.4660
H62.15872.51142.50613.07551.09711.75613.71844.77043.71733.48964.31203.81912.14252.50492.83522.64293.6345
H72.16093.07312.50252.52861.09721.75612.81953.89262.43832.72453.78122.72762.15163.05782.73462.73733.7912
C84.18574.76234.98464.09763.04963.71842.81951.09251.09221.54712.19712.27422.16683.09101.54722.27582.1978
H95.22915.75986.05355.05394.13354.77043.89261.09251.77342.27042.49732.96063.11553.89192.27082.96602.4981
H104.19074.96804.79474.10033.05183.71732.43831.09221.77342.20133.07442.50852.70733.76942.20072.50753.0738
C113.23413.67004.19412.94262.57603.48962.72451.54712.27042.20131.09361.09391.55272.25492.15383.10452.6917
H123.96744.16975.00413.60593.47374.31203.78122.19712.49733.07441.09361.77662.20252.46242.68953.76272.8060
H132.99573.59003.85132.38002.75473.81912.72762.27422.96062.50851.09391.77662.27632.95813.10693.92253.7654
C142.53952.80113.49172.80741.52272.14252.15162.16683.11552.70731.55272.20252.27631.09681.55052.26762.2024
H152.75232.56453.76103.10472.15252.50493.05783.09103.89193.76942.25492.46242.95811.09682.25332.95042.4628
C163.92744.27724.73664.21462.56702.83522.73461.54722.27082.20072.15382.68953.10691.55052.25331.09451.0936
H174.23624.64774.85464.71012.72972.64292.73732.27582.96602.50753.10453.76273.92252.26762.95041.09451.7795
H184.71364.86695.60014.98723.46603.63453.79122.19782.49813.07382.69172.80603.76542.20242.46281.09361.7795

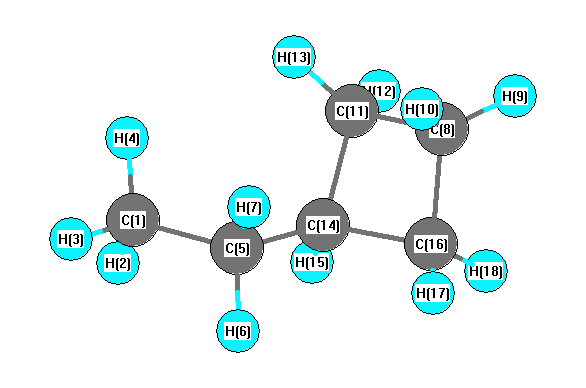
picture of Ethylcyclobutane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C5 H6 109.671 C1 C5 H7 109.835
C1 C5 C14 112.802 H2 C1 H3 107.806
H2 C1 H4 107.755 H2 C1 C5 110.729
H3 C1 H4 107.676 H3 C1 C5 111.467
H4 C1 C5 111.241 C5 C14 C11 113.778
C5 C14 H15 109.439 C5 C14 C16 113.289
H6 C5 H7 106.319 H6 C5 C14 108.649
H7 C5 C14 109.350 C8 C11 H12 111.459
C8 C11 H13 117.857 C8 C11 C14 88.693
C8 C16 C14 88.773 C8 C16 H17 117.952
C8 C16 H18 111.503 H9 C8 H10 108.526
H9 C8 C11 117.626 H9 C8 C16 117.660
H10 C8 C11 111.874 H10 C8 C16 111.822
C11 C8 C16 88.220 C11 C14 H15 115.581
C11 C14 C16 87.903 H12 C11 H13 108.617
H12 C11 C14 111.492 H13 C11 C14 117.601
C14 C16 H17 116.988 C14 C16 H18 111.641
H15 C14 C16 115.611 H17 C16 H18 108.834
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability