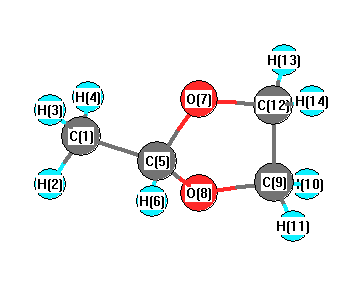
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3147 |
3030 |
22.57 |
|
|
|
| 2 |
A |
3143 |
3026 |
17.88 |
|
|
|
| 3 |
A |
3129 |
3013 |
36.73 |
|
|
|
| 4 |
A |
3082 |
2967 |
48.40 |
|
|
|
| 5 |
A |
3061 |
2947 |
10.00 |
|
|
|
| 6 |
A |
3042 |
2928 |
75.34 |
|
|
|
| 7 |
A |
3004 |
2892 |
63.67 |
|
|
|
| 8 |
A |
2940 |
2830 |
104.80 |
|
|
|
| 9 |
A |
1543 |
1485 |
0.48 |
|
|
|
| 10 |
A |
1524 |
1467 |
4.56 |
|
|
|
| 11 |
A |
1499 |
1443 |
2.55 |
|
|
|
| 12 |
A |
1498 |
1442 |
8.05 |
|
|
|
| 13 |
A |
1453 |
1399 |
78.32 |
|
|
|
| 14 |
A |
1403 |
1350 |
6.53 |
|
|
|
| 15 |
A |
1399 |
1347 |
6.15 |
|
|
|
| 16 |
A |
1386 |
1334 |
0.96 |
|
|
|
| 17 |
A |
1348 |
1298 |
0.48 |
|
|
|
| 18 |
A |
1264 |
1217 |
19.61 |
|
|
|
| 19 |
A |
1239 |
1192 |
1.23 |
|
|
|
| 20 |
A |
1187 |
1143 |
78.90 |
|
|
|
| 21 |
A |
1168 |
1125 |
44.73 |
|
|
|
| 22 |
A |
1144 |
1102 |
78.78 |
|
|
|
| 23 |
A |
1122 |
1080 |
49.57 |
|
|
|
| 24 |
A |
1085 |
1044 |
23.79 |
|
|
|
| 25 |
A |
1062 |
1022 |
56.80 |
|
|
|
| 26 |
A |
960 |
924 |
9.23 |
|
|
|
| 27 |
A |
888 |
855 |
52.79 |
|
|
|
| 28 |
A |
883 |
850 |
3.72 |
|
|
|
| 29 |
A |
858 |
826 |
9.72 |
|
|
|
| 30 |
A |
709 |
683 |
1.51 |
|
|
|
| 31 |
A |
693 |
667 |
3.00 |
|
|
|
| 32 |
A |
511 |
491 |
4.20 |
|
|
|
| 33 |
A |
331 |
318 |
5.98 |
|
|
|
| 34 |
A |
229 |
220 |
0.27 |
|
|
|
| 35 |
A |
201 |
193 |
5.25 |
|
|
|
| 36 |
A |
58 |
56 |
3.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26594.2 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 25602.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.659 |
|
|
|
| 2 |
H |
0.228 |
|
|
|
| 3 |
H |
0.231 |
|
|
|
| 4 |
H |
0.227 |
|
|
|
| 5 |
C |
0.152 |
|
|
|
| 6 |
H |
0.180 |
|
|
|
| 7 |
O |
-0.353 |
|
|
|
| 8 |
O |
-0.328 |
|
|
|
| 9 |
C |
-0.317 |
|
|
|
| 10 |
H |
0.228 |
|
|
|
| 11 |
H |
0.211 |
|
|
|
| 12 |
C |
-0.216 |
|
|
|
| 13 |
H |
0.212 |
|
|
|
| 14 |
H |
0.205 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.151 |
-0.242 |
0.494 |
1.276 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.149 |
-0.157 |
-0.805 |
| y |
-0.157 |
-40.806 |
-0.550 |
| z |
-0.805 |
-0.550 |
-35.739 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.123 |
-0.157 |
-0.805 |
| y |
-0.157 |
-7.361 |
-0.550 |
| z |
-0.805 |
-0.550 |
0.238 |
|
| Polar |
|---|
| 3z2-r2 | 0.477 |
|---|
| x2-y2 | 9.656 |
|---|
| xy | -0.157 |
|---|
| xz | -0.805 |
|---|
| yz | -0.550 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
149.562 |
| (<r2>)1/2 |
12.230 |