Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -193.077558 |
| Energy at 298.15K | -193.084837 |
| HF Energy | -193.077558 |
| Nuclear repulsion energy | 128.428782 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3085 |
2970 |
32.52 |
|
|
|
| 2 |
A1 |
3033 |
2920 |
2.96 |
|
|
|
| 3 |
A1 |
1559 |
1501 |
0.01 |
|
|
|
| 4 |
A1 |
1502 |
1446 |
4.85 |
|
|
|
| 5 |
A1 |
1396 |
1344 |
2.44 |
|
|
|
| 6 |
A1 |
1067 |
1028 |
8.38 |
|
|
|
| 7 |
A1 |
949 |
914 |
32.17 |
|
|
|
| 8 |
A1 |
819 |
788 |
3.92 |
|
|
|
| 9 |
A2 |
3070 |
2956 |
0.00 |
|
|
|
| 10 |
A2 |
1248 |
1201 |
0.00 |
|
|
|
| 11 |
A2 |
1177 |
1133 |
0.00 |
|
|
|
| 12 |
A2 |
849 |
817 |
0.00 |
|
|
|
| 13 |
B1 |
3141 |
3024 |
45.16 |
|
|
|
| 14 |
B1 |
3068 |
2954 |
93.60 |
|
|
|
| 15 |
B1 |
1207 |
1162 |
0.37 |
|
|
|
| 16 |
B1 |
1162 |
1118 |
2.71 |
|
|
|
| 17 |
B1 |
772 |
743 |
0.29 |
|
|
|
| 18 |
B1 |
83 |
80 |
2.64 |
|
|
|
| 19 |
B2 |
3023 |
2910 |
178.66 |
|
|
|
| 20 |
B2 |
1529 |
1472 |
1.50 |
|
|
|
| 21 |
B2 |
1325 |
1275 |
0.27 |
|
|
|
| 22 |
B2 |
1271 |
1224 |
13.23 |
|
|
|
| 23 |
B2 |
1061 |
1021 |
103.31 |
|
|
|
| 24 |
B2 |
963 |
927 |
4.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19178.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 18463.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.068 |
| C2 |
0.000 |
0.000 |
-1.074 |
| C3 |
0.000 |
1.031 |
0.064 |
| C4 |
0.000 |
-1.031 |
0.064 |
| H5 |
0.890 |
0.000 |
-1.705 |
| H6 |
-0.890 |
0.000 |
-1.705 |
| H7 |
0.893 |
1.664 |
0.134 |
| H8 |
-0.893 |
1.664 |
0.134 |
| H9 |
-0.893 |
-1.664 |
0.134 |
| H10 |
0.893 |
-1.664 |
0.134 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1416 | 1.4388 | 1.4388 | 2.9124 | 2.9124 | 2.1063 | 2.1063 | 2.1063 | 2.1063 |
C2 | 2.1416 | | 1.5358 | 1.5358 | 1.0911 | 1.0911 | 2.2411 | 2.2411 | 2.2411 | 2.2411 | C3 | 1.4388 | 1.5358 | | 2.0623 | 2.2331 | 2.2331 | 1.0961 | 1.0961 | 2.8396 | 2.8396 | C4 | 1.4388 | 1.5358 | 2.0623 | | 2.2331 | 2.2331 | 2.8396 | 2.8396 | 1.0961 | 1.0961 | H5 | 2.9124 | 1.0911 | 2.2331 | 2.2331 | | 1.7794 | 2.4800 | 3.0539 | 3.0539 | 2.4800 | H6 | 2.9124 | 1.0911 | 2.2331 | 2.2331 | 1.7794 | | 3.0539 | 2.4800 | 2.4800 | 3.0539 | H7 | 2.1063 | 2.2411 | 1.0961 | 2.8396 | 2.4800 | 3.0539 | | 1.7850 | 3.7758 | 3.3272 | H8 | 2.1063 | 2.2411 | 1.0961 | 2.8396 | 3.0539 | 2.4800 | 1.7850 | | 3.3272 | 3.7758 | H9 | 2.1063 | 2.2411 | 2.8396 | 1.0961 | 3.0539 | 2.4800 | 3.7758 | 3.3272 | | 1.7850 | H10 | 2.1063 | 2.2411 | 2.8396 | 1.0961 | 2.4800 | 3.0539 | 3.3272 | 3.7758 | 1.7850 | |
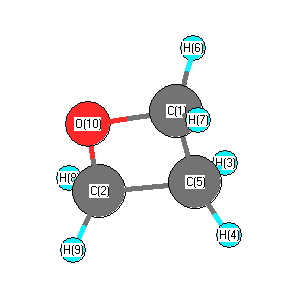 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.044 |
|
O1 |
C3 |
H7 |
111.674 |
| O1 |
C3 |
H8 |
111.674 |
|
O1 |
C4 |
C2 |
92.044 |
| O1 |
C4 |
H9 |
111.674 |
|
O1 |
C4 |
H10 |
111.674 |
| C2 |
C3 |
H7 |
115.746 |
|
C2 |
C3 |
H8 |
115.746 |
| C2 |
C4 |
H9 |
115.746 |
|
C2 |
C4 |
H10 |
115.746 |
| C3 |
O1 |
C4 |
91.559 |
|
C3 |
C2 |
C4 |
84.353 |
| C3 |
C2 |
H5 |
115.402 |
|
C3 |
C2 |
H6 |
115.402 |
| C4 |
C2 |
H5 |
115.402 |
|
C4 |
C2 |
H6 |
115.402 |
| H5 |
C2 |
H6 |
109.261 |
|
H7 |
C3 |
H8 |
109.031 |
| H9 |
C4 |
H10 |
109.031 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.347 |
|
|
|
| 2 |
C |
-0.576 |
|
|
|
| 3 |
C |
-0.169 |
|
|
|
| 4 |
C |
-0.169 |
|
|
|
| 5 |
H |
0.231 |
|
|
|
| 6 |
H |
0.231 |
|
|
|
| 7 |
H |
0.200 |
|
|
|
| 8 |
H |
0.200 |
|
|
|
| 9 |
H |
0.200 |
|
|
|
| 10 |
H |
0.200 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-1.977 |
1.977 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.983 |
0.000 |
0.000 |
| y |
0.000 |
-22.913 |
0.000 |
| z |
0.000 |
0.000 |
-27.775 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.361 |
0.000 |
0.000 |
| y |
0.000 |
2.966 |
0.000 |
| z |
0.000 |
0.000 |
-4.327 |
|
| Polar |
|---|
| 3z2-r2 | -8.655 |
|---|
| x2-y2 | -1.070 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.783 |
0.000 |
0.000 |
| y |
0.000 |
5.818 |
0.000 |
| z |
0.000 |
0.000 |
4.776 |
<r2> (average value of r
2) Å
2
| <r2> |
66.124 |
| (<r2>)1/2 |
8.132 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -193.077558 |
| Energy at 298.15K | -193.084837 |
| HF Energy | -193.077558 |
| Nuclear repulsion energy | 128.429828 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3141 |
3024 |
45.22 |
|
|
|
| 2 |
A' |
3085 |
2970 |
32.52 |
|
|
|
| 3 |
A' |
3068 |
2954 |
93.55 |
|
|
|
| 4 |
A' |
3033 |
2920 |
2.96 |
|
|
|
| 5 |
A' |
1559 |
1501 |
0.01 |
|
|
|
| 6 |
A' |
1502 |
1446 |
4.85 |
|
|
|
| 7 |
A' |
1396 |
1344 |
2.44 |
|
|
|
| 8 |
A' |
1207 |
1162 |
0.37 |
|
|
|
| 9 |
A' |
1162 |
1118 |
2.71 |
|
|
|
| 10 |
A' |
1067 |
1028 |
8.41 |
|
|
|
| 11 |
A' |
949 |
914 |
32.16 |
|
|
|
| 12 |
A' |
819 |
788 |
3.91 |
|
|
|
| 13 |
A' |
772 |
743 |
0.29 |
|
|
|
| 14 |
A' |
83 |
80 |
2.64 |
|
|
|
| 15 |
A" |
3070 |
2956 |
0.00 |
|
|
|
| 16 |
A" |
3023 |
2910 |
178.67 |
|
|
|
| 17 |
A" |
1529 |
1472 |
1.50 |
|
|
|
| 18 |
A" |
1325 |
1275 |
0.27 |
|
|
|
| 19 |
A" |
1271 |
1224 |
13.23 |
|
|
|
| 20 |
A" |
1248 |
1201 |
0.00 |
|
|
|
| 21 |
A" |
1177 |
1133 |
0.00 |
|
|
|
| 22 |
A" |
1061 |
1021 |
103.35 |
|
|
|
| 23 |
A" |
963 |
927 |
4.58 |
|
|
|
| 24 |
A" |
849 |
817 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19178.8 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 18463.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.001 |
-1.068 |
0.000 |
| C2 |
0.000 |
1.074 |
0.000 |
| C3 |
0.000 |
-0.064 |
1.031 |
| C4 |
0.000 |
-0.064 |
-1.031 |
| H5 |
0.890 |
1.706 |
0.000 |
| H6 |
-0.890 |
1.705 |
0.000 |
| H7 |
0.894 |
-0.135 |
1.663 |
| H8 |
-0.892 |
-0.133 |
1.665 |
| H9 |
0.894 |
-0.135 |
-1.663 |
| H10 |
-0.892 |
-0.133 |
-1.665 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1416 | 1.4388 | 1.4388 | 2.9133 | 2.9117 | 2.1063 | 2.1063 | 2.1063 | 2.1063 |
C2 | 2.1416 | | 1.5358 | 1.5358 | 1.0911 | 1.0911 | 2.2410 | 2.2412 | 2.2410 | 2.2412 | C3 | 1.4388 | 1.5358 | | 2.0623 | 2.2334 | 2.2330 | 1.0961 | 1.0961 | 2.8388 | 2.8402 | C4 | 1.4388 | 1.5358 | 2.0623 | | 2.2334 | 2.2330 | 2.8388 | 2.8402 | 1.0961 | 1.0961 | H5 | 2.9133 | 1.0911 | 2.2334 | 2.2334 | | 1.7794 | 2.4801 | 3.0538 | 2.4801 | 3.0538 | H6 | 2.9117 | 1.0911 | 2.2330 | 2.2330 | 1.7794 | | 3.0542 | 2.4799 | 3.0542 | 2.4799 | H7 | 2.1063 | 2.2410 | 1.0961 | 2.8388 | 2.4801 | 3.0542 | | 1.7850 | 3.3252 | 3.7757 | H8 | 2.1063 | 2.2412 | 1.0961 | 2.8402 | 3.0538 | 2.4799 | 1.7850 | | 3.7757 | 3.3290 | H9 | 2.1063 | 2.2410 | 2.8388 | 1.0961 | 2.4801 | 3.0542 | 3.3252 | 3.7757 | | 1.7850 | H10 | 2.1063 | 2.2412 | 2.8402 | 1.0961 | 3.0538 | 2.4799 | 3.7757 | 3.3290 | 1.7850 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
92.046 |
|
O1 |
C3 |
H7 |
111.673 |
| O1 |
C3 |
H8 |
111.677 |
|
O1 |
C4 |
C2 |
92.046 |
| O1 |
C4 |
H9 |
111.673 |
|
O1 |
C4 |
H10 |
111.677 |
| C2 |
C3 |
H7 |
115.735 |
|
C2 |
C3 |
H8 |
115.750 |
| C2 |
C4 |
H9 |
115.735 |
|
C2 |
C4 |
H10 |
115.750 |
| C3 |
O1 |
C4 |
91.560 |
|
C3 |
C2 |
C4 |
84.348 |
| C3 |
C2 |
H5 |
115.421 |
|
C3 |
C2 |
H6 |
115.392 |
| C4 |
C2 |
H5 |
115.421 |
|
C4 |
C2 |
H6 |
115.392 |
| H5 |
C2 |
H6 |
109.253 |
|
H7 |
C3 |
H8 |
109.034 |
| H9 |
C4 |
H10 |
109.034 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.347 |
|
|
|
| 2 |
C |
-0.576 |
|
|
|
| 3 |
C |
-0.169 |
|
|
|
| 4 |
C |
-0.169 |
|
|
|
| 5 |
H |
0.231 |
|
|
|
| 6 |
H |
0.231 |
|
|
|
| 7 |
H |
0.200 |
|
|
|
| 8 |
H |
0.200 |
|
|
|
| 9 |
H |
0.200 |
|
|
|
| 10 |
H |
0.200 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.003 |
1.977 |
0.000 |
1.977 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.983 |
-0.005 |
0.000 |
| y |
-0.005 |
-27.775 |
0.000 |
| z |
0.000 |
0.000 |
-22.914 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.361 |
-0.005 |
0.000 |
| y |
-0.005 |
-4.327 |
0.000 |
| z |
0.000 |
0.000 |
2.966 |
|
| Polar |
|---|
| 3z2-r2 | 5.931 |
|---|
| x2-y2 | 3.792 |
|---|
| xy | -0.005 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.783 |
0.000 |
0.000 |
| y |
0.000 |
4.777 |
0.000 |
| z |
0.000 |
0.000 |
5.818 |
<r2> (average value of r
2) Å
2
| <r2> |
66.123 |
| (<r2>)1/2 |
8.132 |
