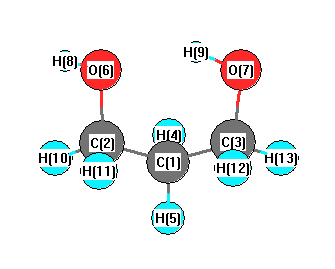
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3835 |
3692 |
17.55 |
|
|
|
| 2 |
A |
3739 |
3600 |
101.40 |
|
|
|
| 3 |
A |
3090 |
2975 |
42.29 |
|
|
|
| 4 |
A |
3084 |
2969 |
57.32 |
|
|
|
| 5 |
A |
3044 |
2930 |
52.30 |
|
|
|
| 6 |
A |
3027 |
2914 |
35.68 |
|
|
|
| 7 |
A |
2991 |
2879 |
70.11 |
|
|
|
| 8 |
A |
2951 |
2841 |
84.49 |
|
|
|
| 9 |
A |
1535 |
1478 |
3.12 |
|
|
|
| 10 |
A |
1519 |
1463 |
1.70 |
|
|
|
| 11 |
A |
1482 |
1427 |
96.35 |
|
|
|
| 12 |
A |
1470 |
1415 |
8.03 |
|
|
|
| 13 |
A |
1459 |
1405 |
5.49 |
|
|
|
| 14 |
A |
1403 |
1350 |
0.87 |
|
|
|
| 15 |
A |
1382 |
1331 |
0.80 |
|
|
|
| 16 |
A |
1321 |
1272 |
32.47 |
|
|
|
| 17 |
A |
1294 |
1246 |
2.91 |
|
|
|
| 18 |
A |
1236 |
1190 |
13.77 |
|
|
|
| 19 |
A |
1210 |
1165 |
70.11 |
|
|
|
| 20 |
A |
1141 |
1098 |
26.32 |
|
|
|
| 21 |
A |
1113 |
1071 |
81.80 |
|
|
|
| 22 |
A |
1098 |
1057 |
57.64 |
|
|
|
| 23 |
A |
971 |
935 |
1.64 |
|
|
|
| 24 |
A |
919 |
884 |
3.37 |
|
|
|
| 25 |
A |
913 |
879 |
14.23 |
|
|
|
| 26 |
A |
828 |
797 |
7.78 |
|
|
|
| 27 |
A |
564 |
543 |
170.39 |
|
|
|
| 28 |
A |
504 |
486 |
3.67 |
|
|
|
| 29 |
A |
391 |
376 |
6.03 |
|
|
|
| 30 |
A |
335 |
323 |
31.89 |
|
|
|
| 31 |
A |
290 |
279 |
78.45 |
|
|
|
| 32 |
A |
212 |
204 |
2.27 |
|
|
|
| 33 |
A |
110 |
106 |
0.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 25229.5 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 24288.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.503 |
|
|
|
| 2 |
C |
-0.196 |
|
|
|
| 3 |
C |
-0.184 |
|
|
|
| 4 |
H |
0.235 |
|
|
|
| 5 |
H |
0.215 |
|
|
|
| 6 |
O |
-0.605 |
|
|
|
| 7 |
O |
-0.575 |
|
|
|
| 8 |
H |
0.407 |
|
|
|
| 9 |
H |
0.405 |
|
|
|
| 10 |
H |
0.209 |
|
|
|
| 11 |
H |
0.200 |
|
|
|
| 12 |
H |
0.179 |
|
|
|
| 13 |
H |
0.214 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.291 |
1.328 |
1.281 |
3.773 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.626 |
-4.711 |
1.188 |
| y |
-4.711 |
-30.234 |
-1.387 |
| z |
1.188 |
-1.387 |
-31.865 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.423 |
-4.711 |
1.188 |
| y |
-4.711 |
1.011 |
-1.387 |
| z |
1.188 |
-1.387 |
-1.435 |
|
| Polar |
|---|
| 3z2-r2 | -2.869 |
|---|
| x2-y2 | -0.392 |
|---|
| xy | -4.711 |
|---|
| xz | 1.188 |
|---|
| yz | -1.387 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.759 |
-0.155 |
0.101 |
| y |
-0.155 |
6.359 |
0.014 |
| z |
0.101 |
0.014 |
5.592 |
<r2> (average value of r
2) Å
2
| <r2> |
0.000 |
| (<r2>)1/2 |
0.000 |