return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H8O2 (1,3-Propanediol)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-268.922568
Energy at 298.15K-268.932695
Nuclear repulsion energy195.575019
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3863 3659 25.00      
2 A 3809 3607 101.49      
3 A 3152 2985 28.81      
4 A 3148 2981 43.51      
5 A 3101 2937 60.33      
6 A 3086 2922 10.42      
7 A 3047 2886 59.68      
8 A 3022 2862 67.87      
9 A 1569 1486 2.81      
10 A 1551 1469 0.99      
11 A 1507 1427 42.73      
12 A 1499 1420 47.48      
13 A 1485 1406 17.23      
14 A 1426 1351 0.46      
15 A 1405 1330 1.66      
16 A 1341 1270 32.09      
17 A 1312 1243 3.51      
18 A 1252 1185 12.44      
19 A 1225 1160 73.70      
20 A 1161 1100 27.55      
21 A 1133 1073 14.81      
22 A 1125 1066 110.81      
23 A 989 937 2.03      
24 A 938 888 5.97      
25 A 928 879 9.27      
26 A 843 799 8.80      
27 A 574 544 174.94      
28 A 516 489 4.18      
29 A 402 381 6.00      
30 A 335 317 25.77      
31 A 286 271 93.43      
32 A 226 214 1.58      
33 A 114 108 1.67      

Unscaled Zero Point Vibrational Energy (zpe) 25683.5 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 24324.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.25770 0.13345 0.09735

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.014 1.041 -0.343
C2 1.296 0.461 0.150
C3 -1.231 0.408 0.326
H4 -0.098 0.894 -1.424
H5 -0.001 2.121 -0.154
O6 1.262 -0.952 -0.066
O7 -1.504 -0.892 -0.158
H8 2.050 -1.346 0.316
H9 -0.668 -1.368 -0.120
H10 2.141 0.909 -0.387
H11 1.414 0.678 1.220
H12 -1.081 0.403 1.418
H13 -2.124 1.002 0.120

Atom - Atom Distances (Å)
  C1 C2 C3 H4 H5 O6 O7 H8 H9 H10 H11 H12 H13
C11.51571.52601.09411.09632.38352.44763.22402.50622.15962.14922.15572.1600
C21.51572.53322.14672.12871.42973.12451.96442.69661.09701.09862.69463.4622
C31.52602.53322.14072.16272.86661.41353.72011.91563.48232.80521.10171.0921
H41.09412.14672.14071.76922.66502.60173.55752.67202.46713.05423.04692.5496
H51.09632.12872.16271.76923.32393.36704.05533.55222.47212.44462.56682.4151
O62.38351.42972.86662.66503.32392.76820.95991.97472.08302.08183.08683.9137
O72.44763.12451.41352.60173.36702.76823.61420.96314.07173.58882.08252.0120
H83.22401.96443.72013.55754.05530.95993.61422.75262.36322.30553.75204.7927
H92.50622.69661.91562.67203.55221.97470.96312.75263.62513.21182.38152.7918
H102.15961.09703.48232.46712.47212.08304.07172.36323.62511.77933.72774.2955
H112.14921.09862.80523.05422.44462.08183.58882.30553.21181.77932.51863.7194
H122.15572.69461.10173.04692.56683.08682.08253.75202.38153.72772.51861.7687
H132.16003.46221.09212.54962.41513.91372.01204.79272.79184.29553.71941.7687

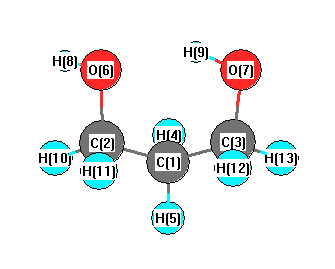
picture of 1,3-Propanediol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O6 108.006 C1 C2 H10 110.472
C1 C2 H11 109.562 C1 C3 O7 112.684
C1 C3 H12 109.174 C1 C3 H13 110.073
C2 C1 C3 112.777 C2 C1 H4 109.621
C2 C1 H5 108.095 C2 O6 H8 108.997
C3 C1 H4 108.446 C3 C1 H5 110.042
C3 O7 H9 105.857 H4 C1 H5 107.741
O6 C2 H10 110.363 O6 C2 H11 110.169
O7 C3 H12 111.171 O7 C3 H13 106.120
H10 C2 H11 108.268 H12 C3 H13 107.457
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability