Jump to
S1C2
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -132.492748 |
| Energy at 298.15K | -132.497014 |
| HF Energy | -132.492748 |
| Nuclear repulsion energy | 65.025021 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3724 |
3368 |
1.19 |
|
|
|
| 2 |
A' |
3412 |
3086 |
12.90 |
|
|
|
| 3 |
A' |
3308 |
2992 |
10.08 |
|
|
|
| 4 |
A' |
3255 |
2944 |
47.92 |
|
|
|
| 5 |
A' |
1635 |
1479 |
13.68 |
|
|
|
| 6 |
A' |
1565 |
1416 |
9.79 |
|
|
|
| 7 |
A' |
1439 |
1302 |
28.97 |
|
|
|
| 8 |
A' |
1264 |
1143 |
13.14 |
|
|
|
| 9 |
A' |
1176 |
1063 |
21.71 |
|
|
|
| 10 |
A' |
1060 |
959 |
3.04 |
|
|
|
| 11 |
A' |
516 |
467 |
12.43 |
|
|
|
| 12 |
A" |
1068 |
966 |
0.12 |
|
|
|
| 13 |
A" |
811 |
734 |
14.61 |
|
|
|
| 14 |
A" |
720 |
651 |
124.73 |
|
|
|
| 15 |
A" |
564 |
510 |
11.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12758.9 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 11539.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.418 |
0.000 |
| C2 |
1.157 |
-0.363 |
0.000 |
| N3 |
-1.193 |
-0.147 |
0.000 |
| H4 |
0.110 |
1.495 |
0.000 |
| H5 |
2.133 |
0.085 |
0.000 |
| H6 |
1.077 |
-1.434 |
0.000 |
| H7 |
-1.911 |
0.554 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3952 | 1.3198 | 1.0826 | 2.1593 | 2.1419 | 1.9159 |
C2 | 1.3952 | | 2.3596 | 2.1320 | 1.0741 | 1.0740 | 3.2018 | N3 | 1.3198 | 2.3596 | | 2.0956 | 3.3345 | 2.6096 | 1.0036 | H4 | 1.0826 | 2.1320 | 2.0956 | | 2.4666 | 3.0840 | 2.2288 | H5 | 2.1593 | 1.0741 | 3.3345 | 2.4666 | | 1.8493 | 4.0716 | H6 | 2.1419 | 1.0740 | 2.6096 | 3.0840 | 1.8493 | | 3.5891 | H7 | 1.9159 | 3.2018 | 1.0036 | 2.2288 | 4.0716 | 3.5891 | |
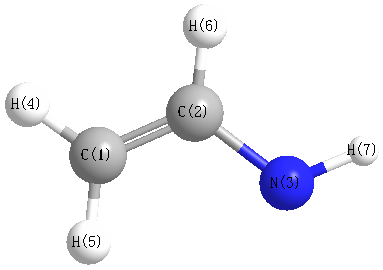 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.413 |
|
C1 |
C2 |
H6 |
119.757 |
| C1 |
N3 |
H7 |
110.356 |
|
C2 |
C1 |
N3 |
120.680 |
| C2 |
C1 |
H4 |
118.182 |
|
N3 |
C1 |
H4 |
121.139 |
| H5 |
C2 |
H6 |
118.830 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.064 |
|
|
|
| 2 |
C |
-0.424 |
|
|
|
| 3 |
N |
-0.478 |
|
|
|
| 4 |
H |
0.198 |
|
|
|
| 5 |
H |
0.228 |
|
|
|
| 6 |
H |
0.242 |
|
|
|
| 7 |
H |
0.298 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.797 |
1.772 |
0.000 |
1.942 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.784 |
-2.963 |
0.000 |
| y |
-2.963 |
-17.822 |
0.000 |
| z |
0.000 |
0.000 |
-20.396 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.325 |
-2.963 |
0.000 |
| y |
-2.963 |
0.768 |
0.000 |
| z |
0.000 |
0.000 |
-3.093 |
|
| Polar |
|---|
| 3z2-r2 | -6.186 |
|---|
| x2-y2 | 1.038 |
|---|
| xy | -2.963 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.689 |
-0.307 |
0.000 |
| y |
-0.307 |
3.860 |
0.000 |
| z |
0.000 |
0.000 |
2.195 |
<r2> (average value of r
2) Å
2
| <r2> |
45.408 |
| (<r2>)1/2 |
6.739 |
Jump to
S1C1
Energy calculated at HF/6-311G*
| | hartrees |
|---|
| Energy at 0K | -132.491850 |
| Energy at 298.15K | -132.496121 |
| HF Energy | -132.491850 |
| Nuclear repulsion energy | 65.053616 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3688 |
3335 |
4.65 |
|
|
|
| 2 |
A' |
3396 |
3071 |
19.39 |
|
|
|
| 3 |
A' |
3310 |
2993 |
20.97 |
|
|
|
| 4 |
A' |
3292 |
2978 |
15.32 |
|
|
|
| 5 |
A' |
1625 |
1469 |
5.42 |
|
|
|
| 6 |
A' |
1566 |
1416 |
3.32 |
|
|
|
| 7 |
A' |
1435 |
1298 |
47.25 |
|
|
|
| 8 |
A' |
1270 |
1149 |
9.56 |
|
|
|
| 9 |
A' |
1194 |
1080 |
29.45 |
|
|
|
| 10 |
A' |
1047 |
947 |
1.04 |
|
|
|
| 11 |
A' |
523 |
473 |
10.71 |
|
|
|
| 12 |
A" |
1096 |
991 |
63.72 |
|
|
|
| 13 |
A" |
820 |
741 |
80.18 |
|
|
|
| 14 |
A" |
720 |
651 |
7.23 |
|
|
|
| 15 |
A" |
553 |
500 |
0.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12766.5 cm
-1
Scaled (by 0.9044) Zero Point Vibrational Energy (zpe) 11546.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.454 |
0.000 |
| C2 |
1.129 |
-0.368 |
0.000 |
| N3 |
-1.255 |
0.045 |
0.000 |
| H4 |
0.144 |
1.523 |
0.000 |
| H5 |
2.119 |
0.048 |
0.000 |
| H6 |
1.038 |
-1.441 |
0.000 |
| H7 |
-1.290 |
-0.961 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3968 | 1.3202 | 1.0787 | 2.1580 | 2.1603 | 1.9147 |
C2 | 1.3968 | | 2.4200 | 2.1326 | 1.0740 | 1.0765 | 2.4910 | N3 | 1.3202 | 2.4200 | | 2.0351 | 3.3746 | 2.7322 | 1.0062 | H4 | 1.0787 | 2.1326 | 2.0351 | | 2.4658 | 3.0957 | 2.8680 | H5 | 2.1580 | 1.0740 | 3.3746 | 2.4658 | | 1.8400 | 3.5557 | H6 | 2.1603 | 1.0765 | 2.7322 | 3.0957 | 1.8400 | | 2.3770 | H7 | 1.9147 | 2.4910 | 1.0062 | 2.8680 | 3.5557 | 2.3770 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.157 |
|
C1 |
C2 |
H6 |
121.180 |
| C1 |
N3 |
H7 |
110.048 |
|
C2 |
C1 |
N3 |
125.895 |
| C2 |
C1 |
H4 |
118.400 |
|
N3 |
C1 |
H4 |
115.705 |
| H5 |
C2 |
H6 |
117.663 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.030 |
|
|
|
| 2 |
C |
-0.468 |
|
|
|
| 3 |
N |
-0.484 |
|
|
|
| 4 |
H |
0.227 |
|
|
|
| 5 |
H |
0.233 |
|
|
|
| 6 |
H |
0.232 |
|
|
|
| 7 |
H |
0.292 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.020 |
-1.132 |
0.000 |
2.315 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.125 |
2.274 |
0.000 |
| y |
2.274 |
-16.174 |
0.000 |
| z |
0.000 |
0.000 |
-20.393 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.842 |
2.274 |
0.000 |
| y |
2.274 |
4.585 |
0.000 |
| z |
0.000 |
0.000 |
-1.743 |
|
| Polar |
|---|
| 3z2-r2 | -3.486 |
|---|
| x2-y2 | -4.951 |
|---|
| xy | 2.274 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.378 |
-0.060 |
0.000 |
| y |
-0.060 |
4.020 |
0.000 |
| z |
0.000 |
0.000 |
2.213 |
<r2> (average value of r
2) Å
2
| <r2> |
45.330 |
| (<r2>)1/2 |
6.733 |
