Jump to
S1C2
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.258009 |
| Energy at 298.15K | -133.262184 |
| HF Energy | -133.258009 |
| Nuclear repulsion energy | 65.053665 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3466 |
3466 |
0.15 |
|
|
|
| 2 |
A' |
3294 |
3294 |
2.25 |
|
|
|
| 3 |
A' |
3179 |
3179 |
3.00 |
|
|
|
| 4 |
A' |
3087 |
3087 |
41.35 |
|
|
|
| 5 |
A' |
1528 |
1528 |
10.71 |
|
|
|
| 6 |
A' |
1494 |
1494 |
2.85 |
|
|
|
| 7 |
A' |
1401 |
1401 |
43.27 |
|
|
|
| 8 |
A' |
1274 |
1274 |
9.66 |
|
|
|
| 9 |
A' |
1105 |
1105 |
26.54 |
|
|
|
| 10 |
A' |
1007 |
1007 |
6.31 |
|
|
|
| 11 |
A' |
499 |
499 |
13.64 |
|
|
|
| 12 |
A" |
1080 |
1080 |
0.21 |
|
|
|
| 13 |
A" |
802 |
802 |
7.86 |
|
|
|
| 14 |
A" |
700 |
700 |
127.46 |
|
|
|
| 15 |
A" |
501 |
501 |
4.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12207.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12207.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| C2 |
1.146 |
-0.394 |
0.000 |
| N3 |
-1.186 |
-0.117 |
0.000 |
| H4 |
0.155 |
1.503 |
0.000 |
| H5 |
2.143 |
0.025 |
0.000 |
| H6 |
1.023 |
-1.469 |
0.000 |
| H7 |
-1.898 |
0.614 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4054 | 1.3014 | 1.0948 | 2.1790 | 2.1477 | 1.9081 |
C2 | 1.4054 | | 2.3485 | 2.1404 | 1.0814 | 1.0823 | 3.2070 | N3 | 1.3014 | 2.3485 | | 2.1027 | 3.3320 | 2.5900 | 1.0207 | H4 | 1.0948 | 2.1404 | 2.1027 | | 2.4772 | 3.0964 | 2.2371 | H5 | 2.1790 | 1.0814 | 3.3320 | 2.4772 | | 1.8677 | 4.0839 | H6 | 2.1477 | 1.0823 | 2.5900 | 3.0964 | 1.8677 | | 3.5881 | H7 | 1.9081 | 3.2070 | 1.0207 | 2.2371 | 4.0839 | 3.5881 | |
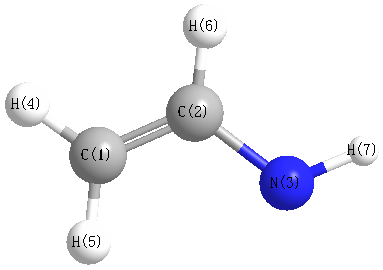 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.840 |
|
C1 |
C2 |
H6 |
118.813 |
| C1 |
N3 |
H7 |
109.925 |
|
C2 |
C1 |
N3 |
120.325 |
| C2 |
C1 |
H4 |
117.218 |
|
N3 |
C1 |
H4 |
122.457 |
| H5 |
C2 |
H6 |
119.348 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.064 |
|
|
|
| 2 |
C |
-0.428 |
|
|
|
| 3 |
N |
-0.484 |
|
|
|
| 4 |
H |
0.203 |
|
|
|
| 5 |
H |
0.237 |
|
|
|
| 6 |
H |
0.249 |
|
|
|
| 7 |
H |
0.287 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.173 |
1.652 |
0.000 |
2.026 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-16.908 |
-2.903 |
0.000 |
| y |
-2.903 |
-17.605 |
0.000 |
| z |
0.000 |
0.000 |
-20.464 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.127 |
-2.903 |
0.000 |
| y |
-2.903 |
1.081 |
0.000 |
| z |
0.000 |
0.000 |
-3.207 |
|
| Polar |
|---|
| 3z2-r2 | -6.414 |
|---|
| x2-y2 | 0.697 |
|---|
| xy | -2.903 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.488 |
-0.428 |
0.000 |
| y |
-0.428 |
4.134 |
0.000 |
| z |
0.000 |
0.000 |
2.221 |
<r2> (average value of r
2) Å
2
| <r2> |
45.316 |
| (<r2>)1/2 |
6.732 |
Jump to
S1C1
Energy calculated at M06-2X/6-311G*
| | hartrees |
|---|
| Energy at 0K | -133.256971 |
| Energy at 298.15K | -133.261141 |
| HF Energy | -133.256971 |
| Nuclear repulsion energy | 65.034578 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3425 |
3425 |
6.77 |
|
|
|
| 2 |
A' |
3277 |
3277 |
5.54 |
|
|
|
| 3 |
A' |
3170 |
3170 |
2.89 |
|
|
|
| 4 |
A' |
3139 |
3139 |
20.04 |
|
|
|
| 5 |
A' |
1514 |
1514 |
7.68 |
|
|
|
| 6 |
A' |
1480 |
1480 |
3.25 |
|
|
|
| 7 |
A' |
1411 |
1411 |
9.34 |
|
|
|
| 8 |
A' |
1270 |
1270 |
54.49 |
|
|
|
| 9 |
A' |
1126 |
1126 |
21.11 |
|
|
|
| 10 |
A' |
995 |
995 |
0.32 |
|
|
|
| 11 |
A' |
507 |
507 |
8.50 |
|
|
|
| 12 |
A" |
1105 |
1105 |
60.29 |
|
|
|
| 13 |
A" |
815 |
815 |
74.81 |
|
|
|
| 14 |
A" |
702 |
702 |
1.09 |
|
|
|
| 15 |
A" |
476 |
476 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12204.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12204.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.449 |
0.000 |
| C2 |
1.123 |
-0.399 |
0.000 |
| N3 |
-1.249 |
0.084 |
0.000 |
| H4 |
0.182 |
1.525 |
0.000 |
| H5 |
2.129 |
-0.003 |
0.000 |
| H6 |
1.001 |
-1.476 |
0.000 |
| H7 |
-1.306 |
-0.939 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.4069 | 1.3012 | 1.0909 | 2.1770 | 2.1702 | 1.9060 |
C2 | 1.4069 | | 2.4203 | 2.1413 | 1.0818 | 1.0845 | 2.4877 | N3 | 1.3012 | 2.4203 | | 2.0305 | 3.3796 | 2.7382 | 1.0252 | H4 | 1.0909 | 2.1413 | 2.0305 | | 2.4753 | 3.1110 | 2.8784 | H5 | 2.1770 | 1.0818 | 3.3796 | 2.4753 | | 1.8560 | 3.5605 | H6 | 2.1702 | 1.0845 | 2.7382 | 3.1110 | 1.8560 | | 2.3682 | H7 | 1.9060 | 2.4877 | 1.0252 | 2.8784 | 3.5605 | 2.3682 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.475 |
|
C1 |
C2 |
H6 |
120.620 |
| C1 |
N3 |
H7 |
109.458 |
|
C2 |
C1 |
N3 |
126.644 |
| C2 |
C1 |
H4 |
117.465 |
|
N3 |
C1 |
H4 |
115.890 |
| H5 |
C2 |
H6 |
117.905 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.042 |
|
|
|
| 2 |
C |
-0.468 |
|
|
|
| 3 |
N |
-0.477 |
|
|
|
| 4 |
H |
0.231 |
|
|
|
| 5 |
H |
0.240 |
|
|
|
| 6 |
H |
0.241 |
|
|
|
| 7 |
H |
0.275 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.281 |
-1.171 |
0.000 |
2.564 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.874 |
2.252 |
0.000 |
| y |
2.252 |
-16.282 |
0.000 |
| z |
0.000 |
0.000 |
-20.463 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.501 |
2.252 |
0.000 |
| y |
2.252 |
4.386 |
0.000 |
| z |
0.000 |
0.000 |
-1.885 |
|
| Polar |
|---|
| 3z2-r2 | -3.769 |
|---|
| x2-y2 | -4.592 |
|---|
| xy | 2.252 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.230 |
-0.150 |
0.000 |
| y |
-0.150 |
4.269 |
0.000 |
| z |
0.000 |
0.000 |
2.227 |
<r2> (average value of r
2) Å
2
| <r2> |
45.354 |
| (<r2>)1/2 |
6.735 |
