Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -168.490159 |
| Energy at 298.15K | |
| HF Energy | -168.296036 |
| Nuclear repulsion energy | 60.337439 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3520 |
3520 |
238.59 |
27.43 |
0.27 |
0.43 |
| 2 |
Σ |
2295 |
2295 |
369.22 |
14.38 |
0.08 |
0.14 |
| 3 |
Σ |
1296 |
1296 |
84.34 |
26.00 |
0.29 |
0.45 |
| 4 |
Π |
563 |
563 |
1.23 |
0.21 |
0.75 |
0.86 |
| 4 |
Π |
563 |
563 |
1.23 |
0.21 |
0.75 |
0.86 |
| 5 |
Π |
203i |
203i |
95.52 |
5.37 |
0.75 |
0.86 |
| 5 |
Π |
203i |
203i |
95.52 |
5.37 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 3915.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 3915.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.184 |
| N2 |
0.000 |
0.000 |
-0.017 |
| C3 |
0.000 |
0.000 |
-1.185 |
| H4 |
0.000 |
0.000 |
-2.245 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.2001 | 2.3681 | 3.4287 |
N2 | 1.2001 | | 1.1680 | 2.2285 | C3 | 2.3681 | 1.1680 | | 1.0605 | H4 | 3.4287 | 2.2285 | 1.0605 | |
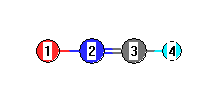 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -168.490349 |
| Energy at 298.15K | -168.490972 |
| HF Energy | -168.294483 |
| Nuclear repulsion energy | 60.311214 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3484 |
3484 |
184.43 |
|
|
|
| 2 |
A' |
2270 |
2270 |
384.25 |
|
|
|
| 3 |
A' |
1299 |
1299 |
66.16 |
|
|
|
| 4 |
A' |
559 |
559 |
3.18 |
|
|
|
| 5 |
A' |
298 |
298 |
157.62 |
|
|
|
| 6 |
A" |
562 |
562 |
1.47 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4236.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4236.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.864 |
-0.805 |
0.000 |
| N2 |
0.000 |
0.025 |
0.000 |
| C3 |
0.915 |
0.762 |
0.000 |
| H4 |
1.422 |
1.695 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.1978 | 2.3698 | 3.3876 |
N2 | 1.1978 | | 1.1743 | 2.1937 | C3 | 2.3698 | 1.1743 | | 1.0628 | H4 | 3.3876 | 2.1937 | 1.0628 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
174.992 |
|
N2 |
C3 |
H4 |
157.362 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
