Jump to
S1C2
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -151.899695 |
| Energy at 298.15K | |
| HF Energy | -151.899695 |
| Nuclear repulsion energy | 52.608892 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3474 |
3344 |
104.27 |
|
|
|
| 2 |
Σ |
2131 |
2051 |
232.71 |
|
|
|
| 3 |
Σ |
1317 |
1268 |
23.79 |
|
|
|
| 4 |
Π |
580 |
558 |
0.08 |
|
|
|
| 4 |
Π |
535 |
515 |
11.39 |
|
|
|
| 5 |
Π |
422 |
406 |
26.88 |
|
|
|
| 5 |
Π |
365i |
351i |
150.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4046.7 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 3895.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.021 |
| C2 |
0.000 |
0.000 |
-1.239 |
| O3 |
0.000 |
0.000 |
1.201 |
| H4 |
0.000 |
0.000 |
-2.302 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2594 | 1.1807 | 2.3230 |
C2 | 1.2594 | | 2.4401 | 1.0636 | O3 | 1.1807 | 2.4401 | | 3.5037 | H4 | 2.3230 | 1.0636 | 3.5037 | |
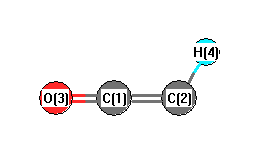 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.210 |
|
|
|
| 2 |
C |
-0.256 |
|
|
|
| 3 |
O |
-0.233 |
|
|
|
| 4 |
H |
0.278 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.078 |
2.078 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-17.786 |
0.000 |
0.000 |
| y |
0.000 |
-16.052 |
0.000 |
| z |
0.000 |
0.000 |
-14.570 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.475 |
0.000 |
0.000 |
| y |
0.000 |
0.126 |
0.000 |
| z |
0.000 |
0.000 |
2.349 |
|
| Polar |
|---|
| 3z2-r2 | 4.697 |
|---|
| x2-y2 | -1.734 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.707 |
0.000 |
0.000 |
| y |
0.000 |
1.601 |
0.000 |
| z |
0.000 |
0.000 |
5.527 |
<r2> (average value of r
2) Å
2
| <r2> |
36.135 |
| (<r2>)1/2 |
6.011 |
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -151.901907 |
| Energy at 298.15K | |
| HF Energy | -151.901907 |
| Nuclear repulsion energy | 52.468806 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3351 |
3226 |
34.95 |
|
|
|
| 2 |
A' |
2118 |
2039 |
313.73 |
|
|
|
| 3 |
A' |
1269 |
1222 |
7.09 |
|
|
|
| 4 |
A' |
569 |
548 |
11.74 |
|
|
|
| 5 |
A' |
494 |
476 |
237.66 |
|
|
|
| 6 |
A" |
509 |
490 |
4.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4154.6 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 3999.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.047 |
0.000 |
| C2 |
1.045 |
-0.711 |
0.000 |
| O3 |
-1.048 |
0.570 |
0.000 |
| H4 |
2.110 |
-0.573 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2908 | 1.1708 | 2.1992 |
C2 | 1.2908 | | 2.4533 | 1.0738 | O3 | 1.1708 | 2.4533 | | 3.3580 | H4 | 2.1992 | 1.0738 | 3.3580 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
136.693 |
|
C2 |
C1 |
O3 |
170.583 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.257 |
|
|
|
| 2 |
C |
-0.320 |
|
|
|
| 3 |
O |
-0.214 |
|
|
|
| 4 |
H |
0.277 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.685 |
0.408 |
0.000 |
1.733 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-14.823 |
0.832 |
0.000 |
| y |
0.832 |
-19.131 |
0.000 |
| z |
0.000 |
0.000 |
-16.165 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.825 |
0.832 |
0.000 |
| y |
0.832 |
-3.637 |
0.000 |
| z |
0.000 |
0.000 |
0.812 |
|
| Polar |
|---|
| 3z2-r2 | 1.624 |
|---|
| x2-y2 | 4.308 |
|---|
| xy | 0.832 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.571 |
-1.717 |
0.000 |
| y |
-1.717 |
3.102 |
0.000 |
| z |
0.000 |
0.000 |
2.089 |
<r2> (average value of r
2) Å
2
| <r2> |
36.188 |
| (<r2>)1/2 |
6.016 |
