Jump to
S1C2
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -151.840614 |
| Energy at 298.15K | |
| HF Energy | -151.680028 |
| Nuclear repulsion energy | 52.507872 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3492 |
3492 |
115.31 |
38.67 |
0.28 |
0.44 |
| 2 |
Σ |
2146 |
2146 |
275.52 |
1.18 |
0.09 |
0.17 |
| 3 |
Σ |
1303 |
1303 |
24.54 |
26.11 |
0.23 |
0.37 |
| 4 |
Π |
579 |
579 |
0.02 |
1.34 |
0.75 |
0.86 |
| 4 |
Π |
539 |
539 |
10.00 |
1.82 |
0.75 |
0.86 |
| 5 |
Π |
415 |
415 |
28.66 |
0.46 |
0.75 |
0.86 |
| 5 |
Π |
421i |
421i |
149.08 |
7.77 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4026.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4026.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
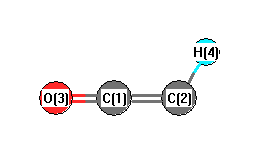
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.020 |
| C2 |
0.000 |
0.000 |
-1.242 |
| O3 |
0.000 |
0.000 |
1.204 |
| H4 |
0.000 |
0.000 |
-2.303 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2619 | 1.1837 | 2.3232 |
C2 | 1.2619 | | 2.4455 | 1.0613 | O3 | 1.1837 | 2.4455 | | 3.5069 | H4 | 2.3232 | 1.0613 | 3.5069 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULL/6-311G*
| | hartrees |
|---|
| Energy at 0K | -151.843664 |
| Energy at 298.15K | |
| HF Energy | -151.680968 |
| Nuclear repulsion energy | 52.310555 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3358 |
3358 |
31.96 |
75.88 |
0.36 |
0.53 |
| 2 |
A' |
2118 |
2118 |
342.28 |
0.39 |
0.20 |
0.34 |
| 3 |
A' |
1242 |
1242 |
6.78 |
27.38 |
0.30 |
0.46 |
| 4 |
A' |
570 |
570 |
57.73 |
7.72 |
0.47 |
0.64 |
| 5 |
A' |
534 |
534 |
200.90 |
5.96 |
0.04 |
0.07 |
| 6 |
A" |
511 |
511 |
3.51 |
0.55 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 4166.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4166.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULL/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.050 |
0.000 |
| C2 |
1.070 |
-0.685 |
0.000 |
| O3 |
-1.068 |
0.536 |
0.000 |
| H4 |
2.123 |
-0.480 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2983 | 1.1734 | 2.1880 |
C2 | 1.2983 | | 2.4623 | 1.0728 | O3 | 1.1734 | 2.4623 | | 3.3487 | H4 | 2.1880 | 1.0728 | 3.3487 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
134.466 |
|
C2 |
C1 |
O3 |
169.995 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
