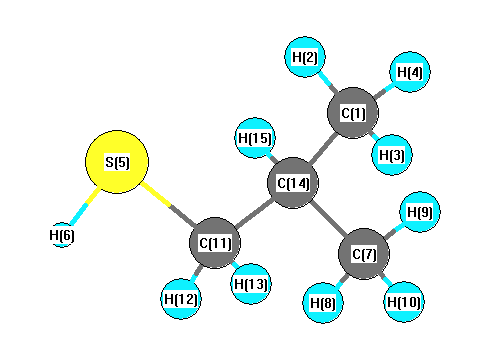
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3138 |
2995 |
32.83 |
|
|
|
| 2 |
A |
3131 |
2989 |
44.94 |
|
|
|
| 3 |
A |
3129 |
2986 |
46.16 |
|
|
|
| 4 |
A |
3122 |
2979 |
26.40 |
|
|
|
| 5 |
A |
3117 |
2975 |
8.83 |
|
|
|
| 6 |
A |
3072 |
2932 |
37.09 |
|
|
|
| 7 |
A |
3053 |
2914 |
23.88 |
|
|
|
| 8 |
A |
3048 |
2909 |
26.01 |
|
|
|
| 9 |
A |
3047 |
2908 |
2.74 |
|
|
|
| 10 |
A |
2673 |
2551 |
12.91 |
|
|
|
| 11 |
A |
1530 |
1460 |
18.31 |
|
|
|
| 12 |
A |
1525 |
1456 |
6.53 |
|
|
|
| 13 |
A |
1515 |
1446 |
5.02 |
|
|
|
| 14 |
A |
1506 |
1437 |
0.31 |
|
|
|
| 15 |
A |
1494 |
1426 |
4.50 |
|
|
|
| 16 |
A |
1438 |
1373 |
11.32 |
|
|
|
| 17 |
A |
1421 |
1356 |
9.77 |
|
|
|
| 18 |
A |
1389 |
1326 |
2.60 |
|
|
|
| 19 |
A |
1373 |
1311 |
3.46 |
|
|
|
| 20 |
A |
1302 |
1243 |
26.68 |
|
|
|
| 21 |
A |
1256 |
1199 |
5.62 |
|
|
|
| 22 |
A |
1210 |
1155 |
5.68 |
|
|
|
| 23 |
A |
1163 |
1110 |
3.76 |
|
|
|
| 24 |
A |
1106 |
1056 |
6.30 |
|
|
|
| 25 |
A |
992 |
947 |
2.06 |
|
|
|
| 26 |
A |
981 |
936 |
0.67 |
|
|
|
| 27 |
A |
942 |
899 |
1.86 |
|
|
|
| 28 |
A |
919 |
877 |
2.66 |
|
|
|
| 29 |
A |
889 |
848 |
3.55 |
|
|
|
| 30 |
A |
816 |
779 |
3.47 |
|
|
|
| 31 |
A |
742 |
708 |
1.73 |
|
|
|
| 32 |
A |
426 |
407 |
0.12 |
|
|
|
| 33 |
A |
395 |
377 |
1.38 |
|
|
|
| 34 |
A |
341 |
325 |
0.16 |
|
|
|
| 35 |
A |
251 |
240 |
0.07 |
|
|
|
| 36 |
A |
226 |
215 |
0.27 |
|
|
|
| 37 |
A |
205 |
196 |
1.94 |
|
|
|
| 38 |
A |
162 |
154 |
16.74 |
|
|
|
| 39 |
A |
81 |
78 |
12.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29062.2 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 27737.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.641 |
|
|
|
| 2 |
H |
0.252 |
|
|
|
| 3 |
H |
0.218 |
|
|
|
| 4 |
H |
0.225 |
|
|
|
| 5 |
S |
-0.118 |
|
|
|
| 6 |
H |
0.156 |
|
|
|
| 7 |
C |
-0.639 |
|
|
|
| 8 |
H |
0.225 |
|
|
|
| 9 |
H |
0.230 |
|
|
|
| 10 |
H |
0.220 |
|
|
|
| 11 |
C |
-0.576 |
|
|
|
| 12 |
H |
0.257 |
|
|
|
| 13 |
H |
0.251 |
|
|
|
| 14 |
C |
-0.296 |
|
|
|
| 15 |
H |
0.236 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.098 |
-1.199 |
0.652 |
1.751 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.273 |
2.684 |
-1.935 |
| y |
2.684 |
-40.592 |
-1.219 |
| z |
-1.935 |
-1.219 |
-42.000 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.023 |
2.684 |
-1.935 |
| y |
2.684 |
1.045 |
-1.219 |
| z |
-1.935 |
-1.219 |
-1.068 |
|
| Polar |
|---|
| 3z2-r2 | -2.135 |
|---|
| x2-y2 | -0.681 |
|---|
| xy | 2.684 |
|---|
| xz | -1.935 |
|---|
| yz | -1.219 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
200.790 |
| (<r2>)1/2 |
14.170 |