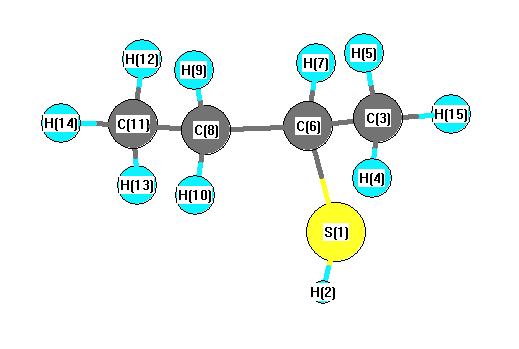
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3149 |
3006 |
23.49 |
|
|
|
| 2 |
A |
3145 |
3001 |
26.07 |
|
|
|
| 3 |
A |
3126 |
2983 |
47.29 |
|
|
|
| 4 |
A |
3117 |
2975 |
34.59 |
|
|
|
| 5 |
A |
3079 |
2939 |
22.23 |
|
|
|
| 6 |
A |
3063 |
2923 |
7.79 |
|
|
|
| 7 |
A |
3057 |
2918 |
23.74 |
|
|
|
| 8 |
A |
3050 |
2911 |
38.81 |
|
|
|
| 9 |
A |
3036 |
2898 |
23.69 |
|
|
|
| 10 |
A |
2653 |
2532 |
10.78 |
|
|
|
| 11 |
A |
1529 |
1459 |
7.57 |
|
|
|
| 12 |
A |
1518 |
1448 |
14.52 |
|
|
|
| 13 |
A |
1516 |
1447 |
2.89 |
|
|
|
| 14 |
A |
1512 |
1443 |
10.17 |
|
|
|
| 15 |
A |
1496 |
1428 |
1.90 |
|
|
|
| 16 |
A |
1430 |
1364 |
13.97 |
|
|
|
| 17 |
A |
1426 |
1361 |
1.66 |
|
|
|
| 18 |
A |
1396 |
1332 |
1.33 |
|
|
|
| 19 |
A |
1348 |
1286 |
12.14 |
|
|
|
| 20 |
A |
1320 |
1260 |
0.28 |
|
|
|
| 21 |
A |
1290 |
1231 |
3.43 |
|
|
|
| 22 |
A |
1195 |
1141 |
1.46 |
|
|
|
| 23 |
A |
1141 |
1089 |
4.08 |
|
|
|
| 24 |
A |
1134 |
1082 |
10.96 |
|
|
|
| 25 |
A |
1056 |
1008 |
0.25 |
|
|
|
| 26 |
A |
1020 |
973 |
8.41 |
|
|
|
| 27 |
A |
984 |
939 |
4.36 |
|
|
|
| 28 |
A |
887 |
846 |
6.65 |
|
|
|
| 29 |
A |
866 |
826 |
4.59 |
|
|
|
| 30 |
A |
796 |
760 |
7.80 |
|
|
|
| 31 |
A |
621 |
593 |
4.55 |
|
|
|
| 32 |
A |
460 |
439 |
0.42 |
|
|
|
| 33 |
A |
380 |
363 |
1.01 |
|
|
|
| 34 |
A |
335 |
320 |
0.92 |
|
|
|
| 35 |
A |
262 |
250 |
0.36 |
|
|
|
| 36 |
A |
235 |
225 |
1.54 |
|
|
|
| 37 |
A |
220 |
210 |
4.90 |
|
|
|
| 38 |
A |
210 |
201 |
14.25 |
|
|
|
| 39 |
A |
113 |
108 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29085.2 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 27758.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.113 |
|
|
|
| 2 |
H |
0.149 |
|
|
|
| 3 |
C |
-0.625 |
|
|
|
| 4 |
H |
0.221 |
|
|
|
| 5 |
H |
0.224 |
|
|
|
| 6 |
C |
-0.441 |
|
|
|
| 7 |
H |
0.258 |
|
|
|
| 8 |
C |
-0.410 |
|
|
|
| 9 |
H |
0.221 |
|
|
|
| 10 |
H |
0.219 |
|
|
|
| 11 |
C |
-0.652 |
|
|
|
| 12 |
H |
0.217 |
|
|
|
| 13 |
H |
0.255 |
|
|
|
| 14 |
H |
0.224 |
|
|
|
| 15 |
H |
0.253 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.657 |
1.682 |
0.550 |
1.887 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.858 |
0.775 |
1.447 |
| y |
0.775 |
-43.189 |
-1.459 |
| z |
1.447 |
-1.459 |
-38.914 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.807 |
0.775 |
1.447 |
| y |
0.775 |
-2.303 |
-1.459 |
| z |
1.447 |
-1.459 |
4.110 |
|
| Polar |
|---|
| 3z2-r2 | 8.219 |
|---|
| x2-y2 | 0.331 |
|---|
| xy | 0.775 |
|---|
| xz | 1.447 |
|---|
| yz | -1.459 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
187.820 |
| (<r2>)1/2 |
13.705 |