return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H12S (Propane, 2-(ethylthio)-)

using model chemistry: MP2=FULL/6-311G*

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A'
Energy calculated at MP2=FULL/6-311G*
 hartrees
Energy at 0K-594.942194
Energy at 298.15K-594.956024
HF Energy-593.894348
Nuclear repulsion energy315.467185
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/6-311G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3183 3015 26.59      
2 A 3174 3006 18.58      
3 A 3162 2995 50.82      
4 A 3101 2937 21.39      
5 A 3092 2928 7.30      
6 A 3086 2923 23.59      
7 A 3078 2915 22.52      
8 A 1547 1465 8.21      
9 A 1543 1461 12.04      
10 A 1541 1459 13.30      
11 A 1525 1445 1.31      
12 A 1456 1379 7.75      
13 A 1450 1373 4.60      
14 A 1345 1274 22.08      
15 A 1339 1269 11.05      
16 A 1215 1151 4.82      
17 A 1127 1067 23.27      
18 A 1081 1024 0.98      
19 A 1027 973 3.20      
20 A 936 886 0.58      
21 A 731 692 0.49      
22 A 651 616 1.19      
23 A 474 449 1.39      
24 A 382 362 0.18      
25 A 302 286 0.79      
26 A 269 255 0.12      
27 A 172 163 0.29      
28 A 3186 3017 14.16      
29 A 3181 3013 19.01      
30 A 3161 2993 4.26      
31 A 3154 2987 1.91      
32 A 3075 2912 16.62      
33 A 1535 1453 7.59      
34 A 1530 1449 6.01      
35 A 1520 1439 0.47      
36 A 1437 1361 12.69      
37 A 1354 1282 0.06      
38 A 1303 1234 0.30      
39 A 1165 1104 0.90      
40 A 1087 1029 0.85      
41 A 982 930 0.01      
42 A 946 896 2.27      
43 A 819 776 5.84      
44 A 334 316 0.88      
45 A 266 252 0.02      
46 A 247 234 0.04      
47 A 74 70 1.43      

Unscaled Zero Point Vibrational Energy (zpe) 36170.4 cm-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 34257.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/6-311G*
ABC
0.15704 0.05258 0.05129

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/6-311G*

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.706 0.730 0.000
C2 1.440 -0.119 0.000
C3 -1.448 -1.041 1.265
C4 -1.448 -1.041 -1.265
S5 0.000 0.984 0.000
C6 -1.395 -0.190 0.000
H7 3.590 0.088 0.000
H8 -2.260 0.483 0.000
H9 -1.426 -0.416 2.159
H10 -1.426 -0.416 -2.159
H11 2.750 1.369 0.884
H12 2.750 1.369 -0.884
H13 1.422 -0.757 -0.887
H14 1.422 -0.757 0.887
H15 -2.367 -1.635 1.280
H16 -2.367 -1.635 -1.280
H17 -0.607 -1.736 1.310
H18 -0.607 -1.736 -1.310

Atom - Atom Distances (Å)
  C1 C2 C3 C4 S5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16 H17 H18
C11.52494.68954.68952.71794.20271.09264.97214.80064.80061.09171.09172.15522.15525.74165.74164.33264.3326
C21.52493.28443.28441.81392.83512.16043.74823.59993.59992.17112.17111.09211.09214.29244.29242.91852.9185
C34.68953.28442.52932.79221.52485.31562.13981.09143.48054.85545.29603.59832.90901.09442.76991.09252.7969
C44.68953.28442.52932.79221.52485.31562.13983.48051.09145.29604.85542.90903.59832.76991.09442.79691.0925
S52.71791.81392.79222.79221.82343.70012.31492.94212.94212.91402.91402.41642.41643.75523.75523.07963.0796
C64.20272.83511.52481.52481.82344.99241.09632.17132.17134.51564.51563.00693.00692.16122.16122.17412.1741
H71.09262.16045.31565.31563.70014.99245.86325.48385.48381.76881.76882.48982.48986.33176.33174.75984.7598
H84.97213.74822.13982.13982.31491.09635.86322.48342.48345.16385.16383.98493.98492.47702.47703.06153.0615
H94.80063.59991.09143.48052.94212.17135.48382.48344.31874.71675.46664.18373.13761.77363.76841.77003.8016
H104.80063.59993.48051.09142.94212.17135.48382.48344.31875.46664.71673.13764.18373.76841.77363.80161.7700
H111.09172.17114.85545.29602.91404.51561.76885.16384.71675.46661.76813.06872.50635.94676.31584.59225.0718
H121.09172.17115.29604.85542.91404.51561.76885.16385.46664.71671.76812.50633.06876.31585.94675.07184.5922
H132.15521.09213.59832.90902.41643.00692.48983.98494.18373.13763.06872.50631.77344.45223.90933.14682.2923
H142.15521.09212.90903.59832.41643.00692.48983.98493.13764.18372.50633.06871.77343.90934.45222.29233.1468
H155.74164.29241.09442.76993.75522.16126.33172.47701.77363.76845.94676.31584.45223.90932.55981.76333.1334
H165.74164.29242.76991.09443.75522.16126.33172.47703.76841.77366.31585.94673.90934.45222.55983.13341.7633
H174.33262.91851.09252.79693.07962.17414.75983.06151.77003.80164.59225.07183.14682.29231.76333.13342.6209
H184.33262.91852.79691.09253.07962.17414.75983.06153.80161.77005.07184.59222.29233.14683.13341.76332.6209

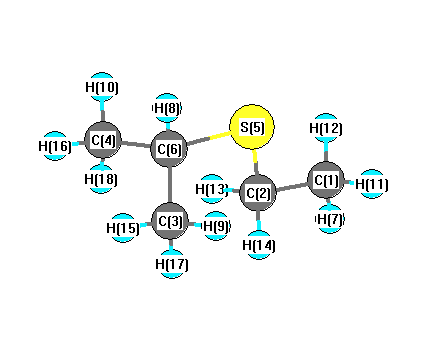
picture of Propane, 2-(ethylthio)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 S5 108.677 C1 C2 H13 109.774
C1 C2 H14 109.774 C2 C1 H7 110.156
C2 C1 H11 111.065 C2 C1 H12 111.065
C2 S5 C6 102.424 C3 C6 C4 112.069
C3 C6 S5 112.707 C3 C6 H8 108.342
C4 C6 S5 112.707 C4 C6 H8 108.342
S5 C2 H13 110.019 S5 C2 H14 110.019
S5 C6 H8 102.013 C6 C3 H9 111.109
C6 C3 H15 110.121 C6 C3 H17 111.259
C6 C4 H10 111.109 C6 C4 H16 110.121
C6 C4 H18 111.259 H7 C1 H11 108.146
H7 C1 H12 108.146 H9 C3 H15 108.478
H9 C3 H17 108.285 H10 C4 H16 108.478
H10 C4 H18 108.285 H11 C1 H12 108.156
H13 C2 H14 108.571 H15 C3 H17 107.468
H16 C4 H18 107.468
Electronic energy levels

Electronic state

Charges, Dipole, Quadrupole and Polarizability