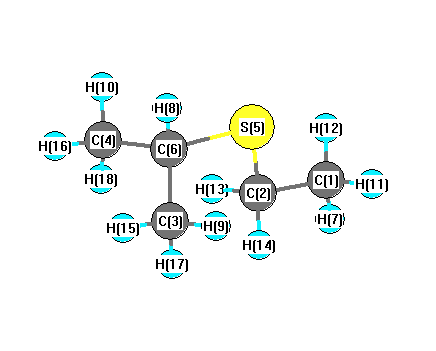
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3149 |
3005 |
32.50 |
|
|
|
| 2 |
A |
3135 |
2992 |
27.96 |
|
|
|
| 3 |
A |
3123 |
2981 |
68.65 |
|
|
|
| 4 |
A |
3074 |
2934 |
21.03 |
|
|
|
| 5 |
A |
3071 |
2931 |
11.20 |
|
|
|
| 6 |
A |
3062 |
2923 |
32.66 |
|
|
|
| 7 |
A |
3056 |
2917 |
32.86 |
|
|
|
| 8 |
A |
1528 |
1459 |
6.79 |
|
|
|
| 9 |
A |
1525 |
1455 |
11.49 |
|
|
|
| 10 |
A |
1522 |
1452 |
12.54 |
|
|
|
| 11 |
A |
1506 |
1437 |
1.58 |
|
|
|
| 12 |
A |
1434 |
1369 |
6.41 |
|
|
|
| 13 |
A |
1428 |
1363 |
3.92 |
|
|
|
| 14 |
A |
1318 |
1258 |
26.99 |
|
|
|
| 15 |
A |
1314 |
1254 |
11.45 |
|
|
|
| 16 |
A |
1197 |
1142 |
7.25 |
|
|
|
| 17 |
A |
1112 |
1062 |
25.23 |
|
|
|
| 18 |
A |
1067 |
1018 |
1.70 |
|
|
|
| 19 |
A |
1013 |
966 |
4.08 |
|
|
|
| 20 |
A |
916 |
874 |
1.10 |
|
|
|
| 21 |
A |
709 |
677 |
0.71 |
|
|
|
| 22 |
A |
628 |
599 |
2.08 |
|
|
|
| 23 |
A |
467 |
446 |
1.50 |
|
|
|
| 24 |
A |
372 |
355 |
0.11 |
|
|
|
| 25 |
A |
292 |
278 |
0.90 |
|
|
|
| 26 |
A |
255 |
244 |
0.21 |
|
|
|
| 27 |
A |
162 |
154 |
0.25 |
|
|
|
| 28 |
A |
3148 |
3004 |
7.49 |
|
|
|
| 29 |
A |
3146 |
3002 |
45.30 |
|
|
|
| 30 |
A |
3120 |
2977 |
4.42 |
|
|
|
| 31 |
A |
3115 |
2973 |
0.62 |
|
|
|
| 32 |
A |
3051 |
2912 |
20.27 |
|
|
|
| 33 |
A |
1515 |
1446 |
6.40 |
|
|
|
| 34 |
A |
1511 |
1442 |
7.08 |
|
|
|
| 35 |
A |
1502 |
1434 |
0.36 |
|
|
|
| 36 |
A |
1417 |
1352 |
10.80 |
|
|
|
| 37 |
A |
1348 |
1286 |
0.21 |
|
|
|
| 38 |
A |
1288 |
1229 |
0.34 |
|
|
|
| 39 |
A |
1151 |
1098 |
1.28 |
|
|
|
| 40 |
A |
1061 |
1013 |
0.56 |
|
|
|
| 41 |
A |
967 |
923 |
0.05 |
|
|
|
| 42 |
A |
938 |
896 |
2.20 |
|
|
|
| 43 |
A |
807 |
770 |
6.29 |
|
|
|
| 44 |
A |
325 |
310 |
1.14 |
|
|
|
| 45 |
A |
250 |
238 |
0.02 |
|
|
|
| 46 |
A |
234 |
223 |
0.07 |
|
|
|
| 47 |
A |
73 |
70 |
1.21 |
|
|
|
| 48 |
A |
46 |
44 |
1.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 35721.9 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 34093.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.672 |
|
|
|
| 2 |
C |
-0.592 |
|
|
|
| 3 |
C |
-0.626 |
|
|
|
| 4 |
C |
-0.626 |
|
|
|
| 5 |
S |
0.124 |
|
|
|
| 6 |
C |
-0.451 |
|
|
|
| 7 |
H |
0.229 |
|
|
|
| 8 |
H |
0.257 |
|
|
|
| 9 |
H |
0.247 |
|
|
|
| 10 |
H |
0.247 |
|
|
|
| 11 |
H |
0.238 |
|
|
|
| 12 |
H |
0.238 |
|
|
|
| 13 |
H |
0.249 |
|
|
|
| 14 |
H |
0.249 |
|
|
|
| 15 |
H |
0.223 |
|
|
|
| 16 |
H |
0.223 |
|
|
|
| 17 |
H |
0.221 |
|
|
|
| 18 |
H |
0.221 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.026 |
-1.771 |
0.000 |
1.771 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-42.965 |
0.096 |
0.000 |
| y |
0.096 |
-49.796 |
0.000 |
| z |
0.000 |
0.000 |
-48.522 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.194 |
0.096 |
0.000 |
| y |
0.096 |
-4.052 |
0.000 |
| z |
0.000 |
0.000 |
-2.142 |
|
| Polar |
|---|
| 3z2-r2 | -4.284 |
|---|
| x2-y2 | 6.831 |
|---|
| xy | 0.096 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
265.383 |
| (<r2>)1/2 |
16.291 |