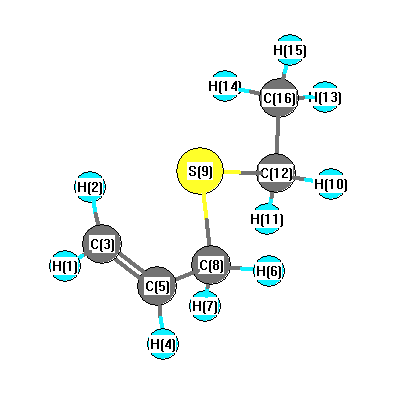
Vibrational Frequencies calculated at mPW1PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3252 |
3104 |
21.10 |
|
|
|
| 2 |
A |
3170 |
3025 |
4.40 |
|
|
|
| 3 |
A |
3154 |
3010 |
21.10 |
|
|
|
| 4 |
A |
3136 |
2993 |
28.37 |
|
|
|
| 5 |
A |
3064 |
2924 |
27.92 |
|
|
|
| 6 |
A |
3060 |
2921 |
33.07 |
|
|
|
| 7 |
A |
3040 |
2901 |
30.77 |
|
|
|
| 8 |
A |
1740 |
1660 |
23.09 |
|
|
|
| 9 |
A |
1525 |
1456 |
3.87 |
|
|
|
| 10 |
A |
1506 |
1437 |
2.07 |
|
|
|
| 11 |
A |
1482 |
1415 |
12.16 |
|
|
|
| 12 |
A |
1463 |
1396 |
6.38 |
|
|
|
| 13 |
A |
1428 |
1363 |
5.64 |
|
|
|
| 14 |
A |
1349 |
1288 |
6.48 |
|
|
|
| 15 |
A |
1337 |
1276 |
5.81 |
|
|
|
| 16 |
A |
1302 |
1243 |
37.00 |
|
|
|
| 17 |
A |
1102 |
1052 |
3.86 |
|
|
|
| 18 |
A |
1081 |
1032 |
2.32 |
|
|
|
| 19 |
A |
1014 |
968 |
4.70 |
|
|
|
| 20 |
A |
954 |
911 |
3.16 |
|
|
|
| 21 |
A |
770 |
735 |
3.76 |
|
|
|
| 22 |
A |
708 |
675 |
1.26 |
|
|
|
| 23 |
A |
508 |
485 |
0.26 |
|
|
|
| 24 |
A |
337 |
321 |
0.76 |
|
|
|
| 25 |
A |
241 |
230 |
0.19 |
|
|
|
| 26 |
A |
122 |
117 |
0.25 |
|
|
|
| 27 |
A |
3146 |
3002 |
33.46 |
|
|
|
| 28 |
A |
3105 |
2963 |
7.73 |
|
|
|
| 29 |
A |
3075 |
2935 |
18.55 |
|
|
|
| 30 |
A |
1514 |
1445 |
11.29 |
|
|
|
| 31 |
A |
1286 |
1227 |
0.28 |
|
|
|
| 32 |
A |
1200 |
1145 |
2.84 |
|
|
|
| 33 |
A |
1066 |
1018 |
0.94 |
|
|
|
| 34 |
A |
1026 |
979 |
16.06 |
|
|
|
| 35 |
A |
947 |
904 |
44.71 |
|
|
|
| 36 |
A |
928 |
886 |
5.93 |
|
|
|
| 37 |
A |
802 |
765 |
7.08 |
|
|
|
| 38 |
A |
554 |
528 |
16.65 |
|
|
|
| 39 |
A |
251 |
240 |
0.07 |
|
|
|
| 40 |
A |
145 |
139 |
0.80 |
|
|
|
| 41 |
A |
65 |
62 |
0.15 |
|
|
|
| 42 |
A |
34 |
32 |
1.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30493.9 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 29103.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.220 |
|
|
|
| 2 |
H |
0.240 |
|
|
|
| 3 |
C |
-0.459 |
|
|
|
| 4 |
H |
0.208 |
|
|
|
| 5 |
C |
-0.164 |
|
|
|
| 6 |
H |
0.255 |
|
|
|
| 7 |
H |
0.255 |
|
|
|
| 8 |
C |
-0.639 |
|
|
|
| 9 |
S |
0.162 |
|
|
|
| 10 |
H |
0.243 |
|
|
|
| 11 |
H |
0.243 |
|
|
|
| 12 |
C |
-0.595 |
|
|
|
| 13 |
H |
0.231 |
|
|
|
| 14 |
H |
0.238 |
|
|
|
| 15 |
H |
0.238 |
|
|
|
| 16 |
C |
-0.677 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.404 |
-1.223 |
0.000 |
1.862 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.763 |
1.597 |
0.000 |
| y |
1.597 |
-43.314 |
0.000 |
| z |
0.000 |
0.000 |
-48.772 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.280 |
1.597 |
0.000 |
| y |
1.597 |
1.953 |
0.000 |
| z |
0.000 |
0.000 |
-6.233 |
|
| Polar |
|---|
| 3z2-r2 | -12.467 |
|---|
| x2-y2 | 1.551 |
|---|
| xy | 1.597 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
294.360 |
| (<r2>)1/2 |
17.157 |