Jump to
S1C2
Vibrational Frequencies calculated at B3PW91/6-311G*
Geometric Data calculated at B3PW91/6-311G*
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3PW91/6-311G*
| | hartrees |
|---|
| Energy at 0K | -269.299961 |
| Energy at 298.15K | -269.306572 |
| HF Energy | -269.299961 |
| Nuclear repulsion energy | 222.970334 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3288 |
3166 |
2.30 |
|
|
|
| 2 |
A' |
3260 |
3139 |
3.01 |
|
|
|
| 3 |
A' |
3248 |
3127 |
6.25 |
|
|
|
| 4 |
A' |
3138 |
3021 |
13.74 |
|
|
|
| 5 |
A' |
3038 |
2925 |
33.80 |
|
|
|
| 6 |
A' |
1655 |
1593 |
15.26 |
|
|
|
| 7 |
A' |
1560 |
1502 |
17.23 |
|
|
|
| 8 |
A' |
1503 |
1447 |
10.41 |
|
|
|
| 9 |
A' |
1430 |
1377 |
6.15 |
|
|
|
| 10 |
A' |
1418 |
1365 |
0.12 |
|
|
|
| 11 |
A' |
1262 |
1215 |
13.31 |
|
|
|
| 12 |
A' |
1256 |
1209 |
5.60 |
|
|
|
| 13 |
A' |
1185 |
1141 |
20.17 |
|
|
|
| 14 |
A' |
1124 |
1082 |
5.02 |
|
|
|
| 15 |
A' |
1043 |
1004 |
23.90 |
|
|
|
| 16 |
A' |
996 |
959 |
2.30 |
|
|
|
| 17 |
A' |
947 |
912 |
17.51 |
|
|
|
| 18 |
A' |
905 |
872 |
3.14 |
|
|
|
| 19 |
A' |
664 |
639 |
0.70 |
|
|
|
| 20 |
A' |
336 |
324 |
2.18 |
|
|
|
| 21 |
A" |
3092 |
2976 |
19.16 |
|
|
|
| 22 |
A" |
1486 |
1431 |
9.56 |
|
|
|
| 23 |
A" |
1068 |
1028 |
2.27 |
|
|
|
| 24 |
A" |
857 |
825 |
0.13 |
|
|
|
| 25 |
A" |
802 |
772 |
26.59 |
|
|
|
| 26 |
A" |
731 |
704 |
56.70 |
|
|
|
| 27 |
A" |
644 |
620 |
2.02 |
|
|
|
| 28 |
A" |
617 |
594 |
7.27 |
|
|
|
| 29 |
A" |
242 |
233 |
6.29 |
|
|
|
| 30 |
A" |
117 |
113 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21456.0 cm
-1
Scaled (by 0.9627) Zero Point Vibrational Energy (zpe) 20655.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/6-311G*
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.034 |
-0.243 |
0.000 |
| C2 |
0.000 |
0.645 |
0.000 |
| C3 |
0.497 |
-1.491 |
0.000 |
| C4 |
-1.181 |
-0.031 |
0.000 |
| C5 |
-0.856 |
-1.425 |
0.000 |
| C6 |
0.355 |
2.084 |
0.000 |
| H7 |
1.207 |
-2.303 |
0.000 |
| H8 |
-2.167 |
0.411 |
0.000 |
| H9 |
-1.542 |
-2.260 |
0.000 |
| H10 |
-0.551 |
2.693 |
0.000 |
| H11 |
0.945 |
2.352 |
0.882 |
| H12 |
0.945 |
2.352 |
-0.882 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| O1 | | 1.3625 | 1.3586 | 2.2248 | 2.2292 | 2.4238 | 2.0675 | 3.2670 | 3.2713 | 3.3359 | 2.7418 | 2.7418 |
C2 | 1.3625 | | 2.1925 | 1.3601 | 2.2393 | 1.4823 | 3.1851 | 2.1795 | 3.2880 | 2.1208 | 2.1414 | 2.1414 | C3 | 1.3586 | 2.1925 | | 2.2242 | 1.3550 | 3.5775 | 1.0785 | 3.2732 | 2.1790 | 4.3127 | 3.9678 | 3.9678 | C4 | 2.2248 | 1.3601 | 2.2242 | | 1.4312 | 2.6131 | 3.2960 | 1.0806 | 2.2579 | 2.7952 | 3.3124 | 3.3124 | C5 | 2.2292 | 2.2393 | 1.3550 | 1.4312 | | 3.7115 | 2.2422 | 2.2554 | 1.0803 | 4.1285 | 4.2758 | 4.2758 | C6 | 2.4238 | 1.4823 | 3.5775 | 2.6131 | 3.7115 | | 4.4689 | 3.0262 | 4.7394 | 1.0911 | 1.0949 | 1.0949 | H7 | 2.0675 | 3.1851 | 1.0785 | 3.2960 | 2.2422 | 4.4689 | | 4.3298 | 2.7488 | 5.2958 | 4.7447 | 4.7447 | H8 | 3.2670 | 2.1795 | 3.2732 | 1.0806 | 2.2554 | 3.0262 | 4.3298 | | 2.7426 | 2.7961 | 3.7722 | 3.7722 | H9 | 3.2713 | 3.2880 | 2.1790 | 2.2579 | 1.0803 | 4.7394 | 2.7488 | 2.7426 | | 5.0503 | 5.3127 | 5.3127 | H10 | 3.3359 | 2.1208 | 4.3127 | 2.7952 | 4.1285 | 1.0911 | 5.2958 | 2.7961 | 5.0503 | | 1.7699 | 1.7699 | H11 | 2.7418 | 2.1414 | 3.9678 | 3.3124 | 4.2758 | 1.0949 | 4.7447 | 3.7722 | 5.3127 | 1.7699 | | 1.7649 | H12 | 2.7418 | 2.1414 | 3.9678 | 3.3124 | 4.2758 | 1.0949 | 4.7447 | 3.7722 | 5.3127 | 1.7699 | 1.7649 | |
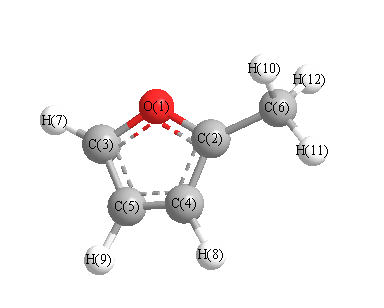 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
109.597 |
|
O1 |
C2 |
C6 |
116.794 |
| O1 |
C3 |
C5 |
110.472 |
|
O1 |
C3 |
H7 |
115.579 |
| C2 |
O1 |
C3 |
107.362 |
|
C2 |
C4 |
C5 |
106.663 |
| C2 |
C4 |
H8 |
126.118 |
|
C2 |
C6 |
H10 |
110.068 |
| C2 |
C6 |
H11 |
111.494 |
|
C2 |
C6 |
H12 |
111.494 |
| C3 |
C5 |
C6 |
73.761 |
|
C3 |
C5 |
H9 |
126.583 |
| C4 |
C2 |
C6 |
133.609 |
|
C4 |
C5 |
H9 |
127.511 |
| C5 |
C3 |
H7 |
133.948 |
|
C5 |
C4 |
H8 |
127.219 |
| H10 |
C6 |
H11 |
108.121 |
|
H10 |
C6 |
H12 |
108.121 |
| H11 |
C6 |
H12 |
107.404 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/6-311G*
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.254 |
|
|
|
| 2 |
C |
0.245 |
|
|
|
| 3 |
C |
-0.049 |
|
|
|
| 4 |
C |
-0.305 |
|
|
|
| 5 |
C |
-0.268 |
|
|
|
| 6 |
C |
-0.715 |
|
|
|
| 7 |
H |
0.214 |
|
|
|
| 8 |
H |
0.203 |
|
|
|
| 9 |
H |
0.206 |
|
|
|
| 10 |
H |
0.239 |
|
|
|
| 11 |
H |
0.242 |
|
|
|
| 12 |
H |
0.242 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.448 |
0.334 |
0.000 |
0.559 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.942 |
-0.545 |
0.000 |
| y |
-0.545 |
-29.907 |
0.000 |
| z |
0.000 |
0.000 |
-37.997 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.010 |
-0.545 |
0.000 |
| y |
-0.545 |
6.062 |
0.000 |
| z |
0.000 |
0.000 |
-6.073 |
|
| Polar |
|---|
| 3z2-r2 | -12.145 |
|---|
| x2-y2 | -4.035 |
|---|
| xy | -0.545 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.457 |
0.386 |
0.000 |
| y |
0.386 |
10.222 |
0.000 |
| z |
0.000 |
0.000 |
4.689 |
<r2> (average value of r
2) Å
2
| <r2> |
140.357 |
| (<r2>)1/2 |
11.847 |
