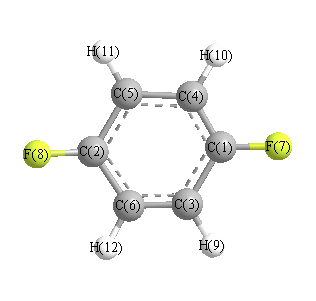
Vibrational Frequencies calculated at MP2=FULL/6-311G*
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3250 |
3078 |
0.00 |
262.87 |
0.18 |
0.30 |
| 2 |
Ag |
1677 |
1588 |
0.00 |
6.47 |
0.58 |
0.73 |
| 3 |
Ag |
1302 |
1233 |
0.00 |
18.67 |
0.12 |
0.22 |
| 4 |
Ag |
1174 |
1112 |
0.00 |
5.98 |
0.64 |
0.78 |
| 5 |
Ag |
869 |
823 |
0.00 |
30.95 |
0.10 |
0.17 |
| 6 |
Ag |
454 |
430 |
0.00 |
4.71 |
0.47 |
0.64 |
| 7 |
Au |
888 |
841 |
0.00 |
0.00 |
0.00 |
0.00 |
| 8 |
Au |
420 |
398 |
0.00 |
0.00 |
0.00 |
0.00 |
| 9 |
B1g |
799 |
757 |
0.00 |
0.66 |
0.75 |
0.86 |
| 10 |
B1u |
3236 |
3065 |
3.88 |
0.00 |
0.48 |
0.65 |
| 11 |
B1u |
1562 |
1479 |
268.42 |
0.00 |
0.49 |
0.66 |
| 12 |
B1u |
1247 |
1181 |
155.02 |
0.00 |
0.50 |
0.67 |
| 13 |
B1u |
1031 |
977 |
4.05 |
0.00 |
0.00 |
0.00 |
| 14 |
B1u |
753 |
713 |
53.81 |
0.00 |
0.51 |
0.67 |
| 15 |
B2g |
849 |
805 |
0.00 |
0.59 |
0.75 |
0.86 |
| 16 |
B2g |
589 |
558 |
0.00 |
1.42 |
0.75 |
0.86 |
| 17 |
B2g |
364 |
345 |
0.00 |
2.80 |
0.75 |
0.86 |
| 18 |
B2u |
3250 |
3078 |
0.37 |
0.00 |
0.18 |
0.31 |
| 19 |
B2u |
1475 |
1397 |
0.98 |
0.00 |
0.51 |
0.67 |
| 20 |
B2u |
1434 |
1358 |
0.12 |
0.00 |
0.51 |
0.67 |
| 21 |
B2u |
1115 |
1056 |
15.26 |
0.00 |
0.51 |
0.67 |
| 22 |
B2u |
348 |
329 |
4.89 |
0.00 |
0.00 |
0.00 |
| 23 |
B3g |
3239 |
3067 |
0.00 |
118.07 |
0.75 |
0.86 |
| 24 |
B3g |
1665 |
1577 |
0.00 |
5.16 |
0.75 |
0.86 |
| 25 |
B3g |
1307 |
1238 |
0.00 |
1.53 |
0.75 |
0.86 |
| 26 |
B3g |
644 |
610 |
0.00 |
7.49 |
0.75 |
0.86 |
| 27 |
B3g |
442 |
418 |
0.00 |
0.09 |
0.75 |
0.86 |
| 28 |
B3u |
834 |
790 |
102.98 |
0.00 |
0.00 |
0.00 |
| 29 |
B3u |
509 |
482 |
18.46 |
0.00 |
0.00 |
0.00 |
| 30 |
B3u |
159 |
151 |
1.41 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 18442.7 cm
-1
Scaled (by 0.9471) Zero Point Vibrational Energy (zpe) 17467.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.